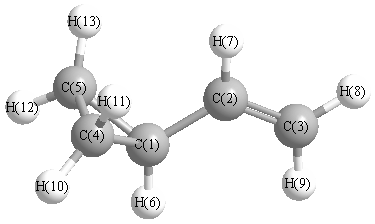
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3277 |
3110 |
19.42 |
|
|
|
| 2 |
A' |
3271 |
3105 |
19.20 |
|
|
|
| 3 |
A' |
3193 |
3031 |
8.07 |
|
|
|
| 4 |
A' |
3192 |
3029 |
7.51 |
|
|
|
| 5 |
A' |
3182 |
3021 |
14.40 |
|
|
|
| 6 |
A' |
3165 |
3004 |
9.77 |
|
|
|
| 7 |
A' |
1730 |
1642 |
21.67 |
|
|
|
| 8 |
A' |
1553 |
1474 |
0.83 |
|
|
|
| 9 |
A' |
1505 |
1428 |
0.21 |
|
|
|
| 10 |
A' |
1389 |
1318 |
1.40 |
|
|
|
| 11 |
A' |
1343 |
1275 |
3.21 |
|
|
|
| 12 |
A' |
1262 |
1197 |
3.01 |
|
|
|
| 13 |
A' |
1243 |
1180 |
1.65 |
|
|
|
| 14 |
A' |
1088 |
1033 |
1.88 |
|
|
|
| 15 |
A' |
1028 |
976 |
4.39 |
|
|
|
| 16 |
A' |
983 |
933 |
35.10 |
|
|
|
| 17 |
A' |
843 |
800 |
1.02 |
|
|
|
| 18 |
A' |
785 |
745 |
0.27 |
|
|
|
| 19 |
A' |
448 |
426 |
1.13 |
|
|
|
| 20 |
A' |
272 |
258 |
1.21 |
|
|
|
| 21 |
A" |
3257 |
3092 |
0.12 |
|
|
|
| 22 |
A" |
3178 |
3017 |
20.38 |
|
|
|
| 23 |
A" |
1514 |
1437 |
0.88 |
|
|
|
| 24 |
A" |
1227 |
1165 |
0.58 |
|
|
|
| 25 |
A" |
1156 |
1097 |
1.88 |
|
|
|
| 26 |
A" |
1097 |
1041 |
5.14 |
|
|
|
| 27 |
A" |
1033 |
980 |
8.55 |
|
|
|
| 28 |
A" |
937 |
890 |
4.19 |
|
|
|
| 29 |
A" |
915 |
869 |
43.06 |
|
|
|
| 30 |
A" |
844 |
801 |
7.61 |
|
|
|
| 31 |
A" |
683 |
649 |
3.83 |
|
|
|
| 32 |
A" |
314 |
299 |
1.00 |
|
|
|
| 33 |
A" |
123 |
117 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25514.7 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 24218.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.150 |
|
|
|
| 2 |
C |
-0.044 |
|
|
|
| 3 |
C |
-0.373 |
|
|
|
| 4 |
C |
-0.332 |
|
|
|
| 5 |
C |
-0.332 |
|
|
|
| 6 |
H |
0.154 |
|
|
|
| 7 |
H |
0.132 |
|
|
|
| 8 |
H |
0.146 |
|
|
|
| 9 |
H |
0.142 |
|
|
|
| 10 |
H |
0.165 |
|
|
|
| 11 |
H |
0.164 |
|
|
|
| 12 |
H |
0.165 |
|
|
|
| 13 |
H |
0.164 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.041 |
0.488 |
0.000 |
0.490 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.404 |
-0.947 |
0.000 |
| y |
-0.947 |
-31.237 |
0.000 |
| z |
0.000 |
0.000 |
-33.334 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.882 |
-0.947 |
0.000 |
| y |
-0.947 |
0.132 |
0.000 |
| z |
0.000 |
0.000 |
-3.014 |
|
| Polar |
|---|
| 3z2-r2 | -6.027 |
|---|
| x2-y2 | 1.833 |
|---|
| xy | -0.947 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.893 |
0.819 |
0.000 |
| y |
0.819 |
10.153 |
0.000 |
| z |
0.000 |
0.000 |
5.288 |
<r2> (average value of r
2) Å
2
| <r2> |
134.504 |
| (<r2>)1/2 |
11.598 |