return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (Cyclopentene, 1-methyl-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-234.345619
Energy at 298.15K-234.356563
HF Energy-234.089453
Nuclear repulsion energy233.202920
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3219 3056 19.95      
2 A 3155 2994 22.96      
3 A 3146 2986 44.53      
4 A 3114 2956 25.84      
5 A 3106 2948 55.96      
6 A 3097 2939 31.15      
7 A 3087 2930 10.49      
8 A 3061 2906 39.19      
9 A 3049 2894 51.13      
10 A 3045 2891 36.98      
11 A 1738 1650 1.65      
12 A 1559 1479 0.36      
13 A 1542 1464 4.23      
14 A 1539 1461 1.62      
15 A 1533 1455 0.40      
16 A 1529 1452 7.74      
17 A 1460 1386 0.77      
18 A 1394 1323 1.00      
19 A 1364 1294 0.31      
20 A 1358 1289 3.12      
21 A 1313 1247 1.04      
22 A 1275 1210 1.43      
23 A 1260 1196 1.14      
24 A 1197 1136 1.14      
25 A 1177 1117 1.01      
26 A 1119 1062 0.52      
27 A 1062 1008 2.47      
28 A 1050 997 4.68      
29 A 1045 992 3.84      
30 A 959 911 1.34      
31 A 935 888 0.34      
32 A 920 873 0.60      
33 A 890 845 0.36      
34 A 846 803 3.26      
35 A 816 775 9.97      
36 A 660 626 1.48      
37 A 583 553 0.10      
38 A 441 419 2.55      
39 A 320 304 0.49      
40 A 245 233 1.86      
41 A 182 173 0.54      
42 A 146 139 0.08      

Unscaled Zero Point Vibrational Energy (zpe) 32267.1 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 30628.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.23233 0.10103 0.07493

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.739 -0.036 -0.034
H2 0.347 -2.106 -0.239
C3 -0.035 -1.131 -0.110
H4 2.597 -0.636 0.831
H5 2.618 0.973 0.110
H6 2.636 -0.456 -0.927
C7 2.225 -0.038 -0.006
H8 0.039 1.561 1.155
H9 0.197 1.980 -0.572
C10 -0.076 1.187 0.130
H11 -1.606 0.725 -1.272
H12 -2.314 1.280 0.252
C13 -1.508 0.683 -0.186
H14 -1.648 -0.963 1.244
H15 -2.173 -1.426 -0.394
C16 -1.460 -0.819 0.175

Atom - Atom Distances (Å)
  C1 H2 C3 H4 H5 H6 C7 H8 H9 C10 H11 H12 C13 H14 H15 C16
C12.11601.34322.13552.13762.13781.48552.11162.15671.47932.75983.33742.36432.86233.24682.3445
H22.11601.05502.89253.84152.90412.80213.93524.10233.34003.59154.33403.34922.73632.61472.2569
C31.34321.05502.83943.39362.87402.51242.97593.15402.33062.69533.33692.33712.11282.17631.4864
H42.13552.89252.83941.76341.76731.09393.38843.81813.31104.89395.30384.43024.27824.98804.1147
H52.13763.84153.39361.76341.76571.09152.84442.70982.70274.45204.94394.14684.82025.38174.4556
H62.13782.90412.87401.76731.76571.09153.89243.46583.34234.41745.37654.36094.82974.93474.2578
C71.48552.80212.51241.09391.09151.09152.94722.91692.61024.10664.73353.80584.17334.62753.7714
H82.11163.93522.97593.38842.84443.89242.94721.78441.09773.04972.53642.22803.03734.02712.9795
H92.15674.10233.15403.81812.70983.46582.91691.78441.09392.30592.73402.17703.91964.15343.3382
C101.47933.34002.33063.31102.70273.34232.61021.09771.09392.12652.24341.55062.88683.39102.4380
H112.75983.59152.69534.89394.45204.41744.10663.04972.30592.12651.76971.09183.03062.39172.1219
H123.33744.33403.33695.30384.94395.37654.73352.53642.73402.24341.76971.09472.54082.78532.2672
C132.36433.34922.33714.43024.14684.36093.80582.22802.17701.55061.09181.09472.18432.22061.5453
H142.86232.73632.11284.27824.82024.82974.17333.03733.91962.88683.03062.54082.18431.78161.0949
H153.24682.61472.17634.98805.38174.93474.62754.02714.15343.39102.39172.78532.22061.78161.0952
C162.34452.25691.48644.11474.45564.25783.77142.97953.33822.43802.12192.26721.54531.09491.0952

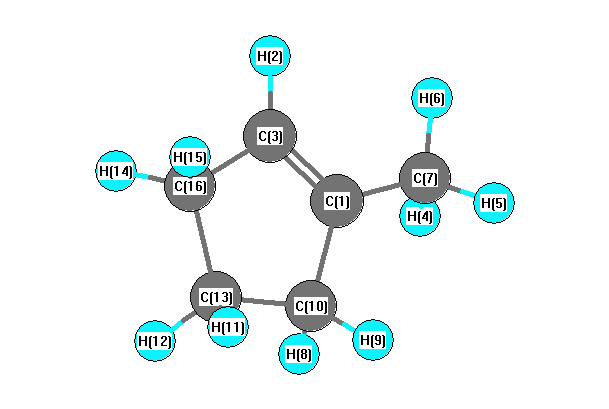
picture of Cyclopentene, 1-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 H2 123.404 C1 C3 C16 111.802
C1 C7 H4 110.850 C1 C7 H5 111.167
C1 C7 H6 111.191 C1 C10 H8 109.151
C1 C10 H9 113.031 C1 C10 C13 102.556
H2 C3 C16 124.389 C3 C1 C7 125.217
C3 C1 C10 111.233 C3 C16 C13 100.852
C3 C16 H14 108.923 C3 C16 H15 114.065
H4 C7 H5 107.578 H4 C7 H6 107.936
H5 C7 H6 107.965 C7 C1 C10 123.380
H8 C10 H9 109.011 H8 C10 C13 113.450
H9 C10 C13 109.606 C10 C13 H11 105.859
C10 C13 H12 114.915 C10 C13 C16 103.904
H11 C13 H12 108.062 H11 C13 C16 105.863
H12 C13 C16 117.341 C13 C16 H14 110.486
C13 C16 H15 113.392 H14 C16 H15 108.875
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.126      
2 H 0.141      
3 C -0.184      
4 H 0.160      
5 H 0.160      
6 H 0.162      
7 C -0.517      
8 H 0.151      
9 H 0.147      
10 C -0.330      
11 H 0.154      
12 H 0.146      
13 C -0.300      
14 H 0.151      
15 H 0.149      
16 C -0.315      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 10.268 0.839 0.083
y 0.839 9.484 -0.108
z 0.083 -0.108 6.709


<r2> (average value of r2) Å2
<r2> 170.498
(<r2>)1/2 13.057