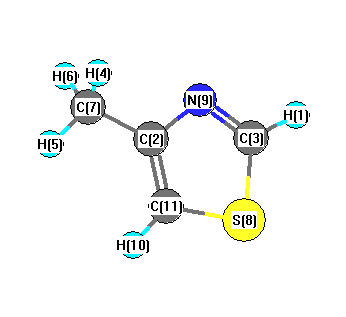
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3287 |
3145 |
1.42 |
|
|
|
| 2 |
A |
3256 |
3115 |
1.31 |
|
|
|
| 3 |
A |
3158 |
3021 |
13.60 |
|
|
|
| 4 |
A |
3130 |
2995 |
12.18 |
|
|
|
| 5 |
A |
3064 |
2932 |
21.73 |
|
|
|
| 6 |
A |
1593 |
1524 |
32.42 |
|
|
|
| 7 |
A |
1513 |
1448 |
49.61 |
|
|
|
| 8 |
A |
1503 |
1438 |
3.58 |
|
|
|
| 9 |
A |
1501 |
1436 |
6.81 |
|
|
|
| 10 |
A |
1435 |
1373 |
3.22 |
|
|
|
| 11 |
A |
1374 |
1314 |
13.85 |
|
|
|
| 12 |
A |
1269 |
1214 |
0.98 |
|
|
|
| 13 |
A |
1177 |
1126 |
3.66 |
|
|
|
| 14 |
A |
1074 |
1027 |
5.33 |
|
|
|
| 15 |
A |
1025 |
980 |
9.26 |
|
|
|
| 16 |
A |
958 |
916 |
15.85 |
|
|
|
| 17 |
A |
898 |
859 |
29.05 |
|
|
|
| 18 |
A |
842 |
805 |
12.61 |
|
|
|
| 19 |
A |
818 |
783 |
42.29 |
|
|
|
| 20 |
A |
734 |
702 |
9.14 |
|
|
|
| 21 |
A |
680 |
650 |
1.55 |
|
|
|
| 22 |
A |
654 |
625 |
0.33 |
|
|
|
| 23 |
A |
561 |
537 |
1.93 |
|
|
|
| 24 |
A |
491 |
470 |
1.97 |
|
|
|
| 25 |
A |
333 |
319 |
2.52 |
|
|
|
| 26 |
A |
233 |
223 |
3.99 |
|
|
|
| 27 |
A |
125 |
119 |
0.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18342.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 17547.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.211 |
|
|
0.164 |
| 2 |
C |
0.299 |
|
|
0.581 |
| 3 |
C |
-0.169 |
|
|
0.052 |
| 4 |
H |
0.196 |
|
|
0.200 |
| 5 |
H |
0.176 |
|
|
0.161 |
| 6 |
H |
0.196 |
|
|
0.200 |
| 7 |
C |
-0.567 |
|
|
-0.691 |
| 8 |
S |
0.271 |
|
|
0.016 |
| 9 |
N |
-0.385 |
|
|
-0.482 |
| 10 |
H |
0.207 |
|
|
0.248 |
| 11 |
C |
-0.435 |
|
|
-0.446 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.104 |
-1.170 |
0.000 |
1.175 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.118 |
-1.117 |
0.000 |
1.123 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.605 |
3.897 |
-0.001 |
| y |
3.897 |
-40.555 |
0.000 |
| z |
-0.001 |
0.000 |
-43.822 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.584 |
3.897 |
-0.001 |
| y |
3.897 |
0.658 |
0.000 |
| z |
-0.001 |
0.000 |
-4.242 |
|
| Polar |
|---|
| 3z2-r2 | -8.484 |
|---|
| x2-y2 | 1.951 |
|---|
| xy | 3.897 |
|---|
| xz | -0.001 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
179.101 |
| (<r2>)1/2 |
13.383 |