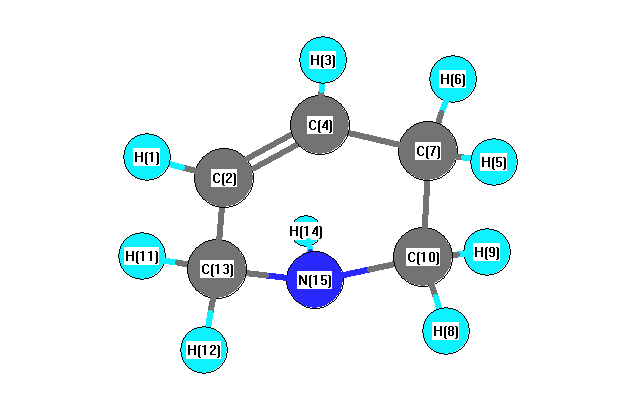
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3501 |
3349 |
0.77 |
|
|
|
| 2 |
A |
3185 |
3047 |
41.59 |
|
|
|
| 3 |
A |
3162 |
3025 |
10.99 |
|
|
|
| 4 |
A |
3106 |
2971 |
39.88 |
|
|
|
| 5 |
A |
3073 |
2940 |
29.58 |
|
|
|
| 6 |
A |
3065 |
2932 |
64.27 |
|
|
|
| 7 |
A |
3051 |
2919 |
16.53 |
|
|
|
| 8 |
A |
3016 |
2885 |
45.67 |
|
|
|
| 9 |
A |
3012 |
2882 |
41.24 |
|
|
|
| 10 |
A |
1744 |
1668 |
1.97 |
|
|
|
| 11 |
A |
1514 |
1449 |
6.32 |
|
|
|
| 12 |
A |
1512 |
1447 |
1.64 |
|
|
|
| 13 |
A |
1500 |
1435 |
8.31 |
|
|
|
| 14 |
A |
1496 |
1431 |
1.12 |
|
|
|
| 15 |
A |
1431 |
1369 |
0.38 |
|
|
|
| 16 |
A |
1408 |
1347 |
7.47 |
|
|
|
| 17 |
A |
1384 |
1324 |
4.48 |
|
|
|
| 18 |
A |
1364 |
1305 |
0.50 |
|
|
|
| 19 |
A |
1342 |
1284 |
0.52 |
|
|
|
| 20 |
A |
1271 |
1216 |
5.15 |
|
|
|
| 21 |
A |
1226 |
1173 |
11.96 |
|
|
|
| 22 |
A |
1206 |
1154 |
4.49 |
|
|
|
| 23 |
A |
1146 |
1096 |
16.44 |
|
|
|
| 24 |
A |
1118 |
1070 |
3.97 |
|
|
|
| 25 |
A |
1060 |
1014 |
3.82 |
|
|
|
| 26 |
A |
1016 |
972 |
7.37 |
|
|
|
| 27 |
A |
999 |
956 |
0.12 |
|
|
|
| 28 |
A |
988 |
945 |
3.54 |
|
|
|
| 29 |
A |
928 |
888 |
7.72 |
|
|
|
| 30 |
A |
906 |
867 |
8.04 |
|
|
|
| 31 |
A |
869 |
831 |
20.00 |
|
|
|
| 32 |
A |
822 |
786 |
94.64 |
|
|
|
| 33 |
A |
763 |
730 |
26.50 |
|
|
|
| 34 |
A |
653 |
625 |
23.25 |
|
|
|
| 35 |
A |
527 |
504 |
1.20 |
|
|
|
| 36 |
A |
485 |
464 |
0.50 |
|
|
|
| 37 |
A |
400 |
382 |
1.13 |
|
|
|
| 38 |
A |
295 |
282 |
8.13 |
|
|
|
| 39 |
A |
167 |
160 |
1.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29853.8 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 28561.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.150 |
|
|
0.110 |
| 2 |
C |
-0.150 |
|
|
-0.258 |
| 3 |
H |
0.152 |
|
|
0.107 |
| 4 |
C |
-0.131 |
|
|
-0.144 |
| 5 |
H |
0.167 |
|
|
0.013 |
| 6 |
H |
0.166 |
|
|
0.043 |
| 7 |
C |
-0.365 |
|
|
-0.009 |
| 8 |
H |
0.173 |
|
|
0.083 |
| 9 |
H |
0.166 |
|
|
0.032 |
| 10 |
C |
-0.206 |
|
|
0.098 |
| 11 |
H |
0.171 |
|
|
0.021 |
| 12 |
H |
0.178 |
|
|
0.048 |
| 13 |
C |
-0.233 |
|
|
0.254 |
| 14 |
H |
0.312 |
|
|
0.382 |
| 15 |
N |
-0.549 |
|
|
-0.780 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.518 |
0.795 |
-0.519 |
1.081 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.499 |
0.749 |
-0.508 |
1.034 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.148 |
-2.049 |
0.210 |
| y |
-2.049 |
-39.519 |
0.908 |
| z |
0.210 |
0.908 |
-36.189 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.707 |
-2.049 |
0.210 |
| y |
-2.049 |
-3.851 |
0.908 |
| z |
0.210 |
0.908 |
1.144 |
|
| Polar |
|---|
| 3z2-r2 | 2.289 |
|---|
| x2-y2 | 4.372 |
|---|
| xy | -2.049 |
|---|
| xz | 0.210 |
|---|
| yz | 0.908 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
146.727 |
| (<r2>)1/2 |
12.113 |