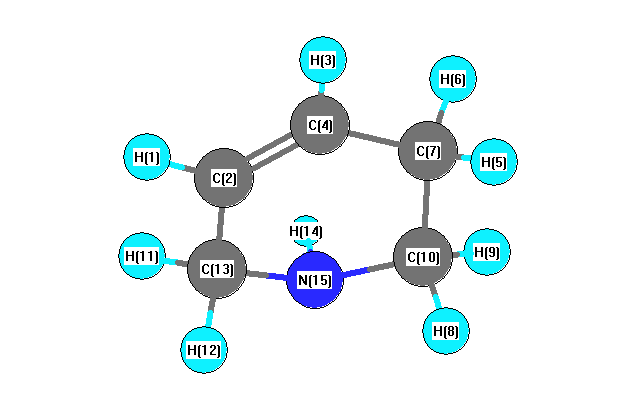
Vibrational Frequencies calculated at B3LYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3469 |
3331 |
1.89 |
|
|
|
| 2 |
A |
3172 |
3046 |
44.46 |
|
|
|
| 3 |
A |
3149 |
3024 |
12.18 |
|
|
|
| 4 |
A |
3091 |
2969 |
43.55 |
|
|
|
| 5 |
A |
3059 |
2938 |
33.41 |
|
|
|
| 6 |
A |
3052 |
2930 |
71.01 |
|
|
|
| 7 |
A |
3037 |
2916 |
15.38 |
|
|
|
| 8 |
A |
3002 |
2883 |
85.95 |
|
|
|
| 9 |
A |
3001 |
2882 |
4.10 |
|
|
|
| 10 |
A |
1735 |
1666 |
2.00 |
|
|
|
| 11 |
A |
1520 |
1459 |
2.71 |
|
|
|
| 12 |
A |
1514 |
1454 |
2.71 |
|
|
|
| 13 |
A |
1507 |
1447 |
5.76 |
|
|
|
| 14 |
A |
1502 |
1442 |
1.63 |
|
|
|
| 15 |
A |
1433 |
1376 |
0.99 |
|
|
|
| 16 |
A |
1409 |
1353 |
9.09 |
|
|
|
| 17 |
A |
1385 |
1330 |
4.50 |
|
|
|
| 18 |
A |
1369 |
1315 |
0.58 |
|
|
|
| 19 |
A |
1342 |
1289 |
0.69 |
|
|
|
| 20 |
A |
1271 |
1220 |
4.68 |
|
|
|
| 21 |
A |
1222 |
1173 |
9.06 |
|
|
|
| 22 |
A |
1208 |
1160 |
3.96 |
|
|
|
| 23 |
A |
1130 |
1085 |
20.30 |
|
|
|
| 24 |
A |
1121 |
1076 |
4.13 |
|
|
|
| 25 |
A |
1050 |
1009 |
2.63 |
|
|
|
| 26 |
A |
1015 |
974 |
8.77 |
|
|
|
| 27 |
A |
1000 |
961 |
0.12 |
|
|
|
| 28 |
A |
990 |
951 |
5.04 |
|
|
|
| 29 |
A |
918 |
881 |
6.58 |
|
|
|
| 30 |
A |
897 |
861 |
2.90 |
|
|
|
| 31 |
A |
865 |
831 |
34.72 |
|
|
|
| 32 |
A |
814 |
782 |
78.58 |
|
|
|
| 33 |
A |
765 |
734 |
24.30 |
|
|
|
| 34 |
A |
655 |
629 |
19.61 |
|
|
|
| 35 |
A |
528 |
507 |
1.03 |
|
|
|
| 36 |
A |
487 |
468 |
0.51 |
|
|
|
| 37 |
A |
399 |
384 |
1.01 |
|
|
|
| 38 |
A |
295 |
283 |
7.74 |
|
|
|
| 39 |
A |
167 |
161 |
1.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29771.5 cm
-1
Scaled (by 0.9603) Zero Point Vibrational Energy (zpe) 28589.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.120 |
0.091 |
|
0.100 |
| 2 |
C |
-0.119 |
-0.282 |
|
-0.244 |
| 3 |
H |
0.121 |
0.095 |
|
0.100 |
| 4 |
C |
-0.102 |
-0.131 |
|
-0.144 |
| 5 |
H |
0.139 |
-0.024 |
|
-0.005 |
| 6 |
H |
0.139 |
0.002 |
|
0.024 |
| 7 |
C |
-0.307 |
0.047 |
|
0.038 |
| 8 |
H |
0.146 |
0.004 |
|
0.065 |
| 9 |
H |
0.138 |
-0.028 |
|
0.015 |
| 10 |
C |
-0.151 |
0.303 |
|
0.135 |
| 11 |
H |
0.143 |
-0.043 |
|
0.012 |
| 12 |
H |
0.152 |
-0.024 |
|
0.038 |
| 13 |
C |
-0.180 |
0.471 |
|
0.270 |
| 14 |
H |
0.291 |
0.347 |
|
0.367 |
| 15 |
N |
-0.531 |
-0.828 |
|
-0.770 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.500 |
0.795 |
-0.514 |
1.070 |
| CHELPG |
0.464 |
0.761 |
-0.502 |
1.024 |
| AIM |
|
|
|
|
| ESP |
0.474 |
0.752 |
-0.504 |
1.021 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.550 |
-1.972 |
0.219 |
| y |
-1.972 |
-39.864 |
0.900 |
| z |
0.219 |
0.900 |
-36.410 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.587 |
-1.972 |
0.219 |
| y |
-1.972 |
-3.883 |
0.900 |
| z |
0.219 |
0.900 |
1.297 |
|
| Polar |
|---|
| 3z2-r2 | 2.593 |
|---|
| x2-y2 | 4.313 |
|---|
| xy | -1.972 |
|---|
| xz | 0.219 |
|---|
| yz | 0.900 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.264 |
-0.144 |
-0.224 |
| y |
-0.144 |
8.130 |
0.092 |
| z |
-0.224 |
0.092 |
6.341 |
<r2> (average value of r
2) Å
2
| <r2> |
147.679 |
| (<r2>)1/2 |
12.152 |