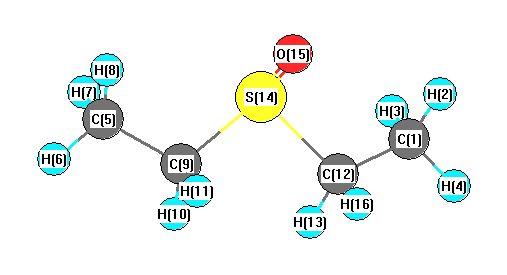
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3159 |
3022 |
14.82 |
|
|
|
| 2 |
A |
3152 |
3016 |
17.49 |
|
|
|
| 3 |
A |
3145 |
3009 |
22.56 |
|
|
|
| 4 |
A |
3139 |
3004 |
17.70 |
|
|
|
| 5 |
A |
3134 |
2998 |
4.36 |
|
|
|
| 6 |
A |
3131 |
2995 |
1.39 |
|
|
|
| 7 |
A |
3088 |
2954 |
5.30 |
|
|
|
| 8 |
A |
3083 |
2950 |
15.28 |
|
|
|
| 9 |
A |
3069 |
2936 |
20.86 |
|
|
|
| 10 |
A |
3064 |
2931 |
19.40 |
|
|
|
| 11 |
A |
1532 |
1465 |
4.98 |
|
|
|
| 12 |
A |
1527 |
1461 |
9.78 |
|
|
|
| 13 |
A |
1524 |
1458 |
2.35 |
|
|
|
| 14 |
A |
1522 |
1456 |
14.57 |
|
|
|
| 15 |
A |
1494 |
1429 |
9.21 |
|
|
|
| 16 |
A |
1484 |
1420 |
5.06 |
|
|
|
| 17 |
A |
1434 |
1372 |
7.71 |
|
|
|
| 18 |
A |
1426 |
1364 |
7.03 |
|
|
|
| 19 |
A |
1326 |
1269 |
0.39 |
|
|
|
| 20 |
A |
1299 |
1243 |
3.71 |
|
|
|
| 21 |
A |
1273 |
1218 |
1.61 |
|
|
|
| 22 |
A |
1263 |
1208 |
0.82 |
|
|
|
| 23 |
A |
1099 |
1051 |
76.42 |
|
|
|
| 24 |
A |
1096 |
1049 |
35.43 |
|
|
|
| 25 |
A |
1078 |
1031 |
11.23 |
|
|
|
| 26 |
A |
1068 |
1022 |
8.72 |
|
|
|
| 27 |
A |
1028 |
984 |
2.52 |
|
|
|
| 28 |
A |
998 |
954 |
1.06 |
|
|
|
| 29 |
A |
985 |
943 |
8.35 |
|
|
|
| 30 |
A |
800 |
765 |
11.53 |
|
|
|
| 31 |
A |
766 |
733 |
9.74 |
|
|
|
| 32 |
A |
678 |
649 |
21.71 |
|
|
|
| 33 |
A |
597 |
571 |
6.99 |
|
|
|
| 34 |
A |
470 |
450 |
4.67 |
|
|
|
| 35 |
A |
362 |
346 |
1.36 |
|
|
|
| 36 |
A |
303 |
290 |
3.02 |
|
|
|
| 37 |
A |
271 |
259 |
1.26 |
|
|
|
| 38 |
A |
212 |
203 |
2.74 |
|
|
|
| 39 |
A |
204 |
195 |
3.71 |
|
|
|
| 40 |
A |
194 |
186 |
5.33 |
|
|
|
| 41 |
A |
117 |
112 |
2.16 |
|
|
|
| 42 |
A |
47 |
45 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30319.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 29006.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.526 |
|
|
-0.184 |
| 2 |
H |
0.184 |
|
|
0.052 |
| 3 |
H |
0.173 |
|
|
0.076 |
| 4 |
H |
0.220 |
|
|
0.089 |
| 5 |
C |
-0.536 |
|
|
-0.312 |
| 6 |
H |
0.191 |
|
|
0.090 |
| 7 |
H |
0.192 |
|
|
0.093 |
| 8 |
H |
0.181 |
|
|
0.122 |
| 9 |
C |
-0.519 |
|
|
-0.030 |
| 10 |
H |
0.217 |
|
|
0.127 |
| 11 |
H |
0.213 |
|
|
0.064 |
| 12 |
C |
-0.525 |
|
|
-0.020 |
| 13 |
H |
0.193 |
|
|
0.030 |
| 14 |
S |
0.775 |
|
|
0.117 |
| 15 |
O |
-0.648 |
|
|
-0.423 |
| 16 |
H |
0.214 |
|
|
0.110 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.056 |
3.380 |
-0.778 |
4.032 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-2.075 |
3.392 |
-0.783 |
4.053 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.997 |
4.288 |
-2.452 |
| y |
4.288 |
-49.355 |
0.927 |
| z |
-2.452 |
0.927 |
-44.641 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.001 |
4.288 |
-2.452 |
| y |
4.288 |
-5.536 |
0.927 |
| z |
-2.452 |
0.927 |
1.535 |
|
| Polar |
|---|
| 3z2-r2 | 3.070 |
|---|
| x2-y2 | 6.359 |
|---|
| xy | 4.288 |
|---|
| xz | -2.452 |
|---|
| yz | 0.927 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
224.655 |
| (<r2>)1/2 |
14.989 |