return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10OS (Diethyl sulfoxide)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-631.373930
Energy at 298.15K-631.385282
HF Energy-631.092778
Nuclear repulsion energy329.114324
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3180 3019 13.73      
2 A 3174 3012 15.28      
3 A 3167 3006 32.70      
4 A 3163 3002 12.91      
5 A 3156 2996 5.14      
6 A 3151 2991 1.56      
7 A 3110 2952 6.01      
8 A 3104 2947 13.82      
9 A 3092 2935 18.41      
10 A 3088 2931 15.50      
11 A 1558 1479 4.04      
12 A 1553 1474 8.79      
13 A 1550 1471 1.95      
14 A 1548 1469 11.58      
15 A 1520 1443 7.84      
16 A 1509 1432 3.91      
17 A 1459 1385 5.29      
18 A 1452 1378 5.93      
19 A 1349 1280 0.18      
20 A 1323 1256 4.50      
21 A 1295 1230 1.57      
22 A 1283 1218 0.67      
23 A 1114 1057 14.47      
24 A 1097 1041 34.72      
25 A 1083 1028 9.77      
26 A 1069 1015 57.82      
27 A 1046 993 6.35      
28 A 1006 955 1.68      
29 A 995 944 8.20      
30 A 812 771 9.62      
31 A 779 739 7.93      
32 A 687 652 20.27      
33 A 603 573 7.36      
34 A 473 449 5.05      
35 A 365 346 0.97      
36 A 308 292 3.11      
37 A 279 265 1.35      
38 A 221 210 3.45      
39 A 212 201 4.55      
40 A 199 189 4.42      
41 A 122 116 2.17      
42 A 45 42 0.07      

Unscaled Zero Point Vibrational Energy (zpe) 30646.7 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 29089.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.11271 0.07987 0.05804

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.577 -1.469 0.551
H2 -2.258 -2.271 0.257
H3 -0.703 -1.920 1.027
H4 -2.082 -0.825 1.271
C5 2.192 -0.824 -0.203
H6 2.558 -0.250 -1.056
H7 3.043 -1.175 0.380
H8 1.630 -1.682 -0.570
C9 1.302 0.079 0.655
H10 1.877 0.934 1.006
H11 0.908 -0.462 1.522
C12 -1.150 -0.672 -0.696
H13 -0.650 -1.348 -1.388
S14 -0.033 0.728 -0.383
O15 -0.797 1.866 0.376
H16 -2.028 -0.254 -1.187

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 H11 C12 H13 S14 O15 H16
C11.09281.09241.09073.89694.60064.63213.40383.27064.23232.85171.54062.15232.84303.42962.1684
H21.09281.77111.77564.70185.38545.41454.01864.28485.28553.86012.16672.47833.78884.38932.4914
H31.09241.77111.77823.33024.21403.87322.83722.85523.84702.22812.17382.48223.07333.84223.0715
H41.09071.77561.77824.52105.22295.21384.23173.55704.34063.02302.18203.06503.05703.11332.5239
C53.89694.70183.33024.52101.09121.08991.08911.53042.15672.17993.38093.12332.71834.06214.3706
H64.60065.38544.21405.22291.09121.77551.77362.14772.47353.06753.74903.40712.85024.21714.5885
H74.63215.41453.87325.21381.08991.77551.77662.16272.48982.52334.35764.09813.69654.89795.3872
H83.40384.01862.83724.23171.08911.77361.77662.16993.06402.52682.96022.44532.93374.40113.9754
C93.27064.28482.85523.55701.53042.14772.16272.16991.08851.09482.89843.16571.81152.77083.8207
H104.23235.28553.84704.34062.15672.47352.48983.06401.08851.77583.82564.16242.37062.90084.6338
H112.85173.86012.22813.02302.17993.06752.52332.52681.09481.77583.03253.41742.43493.10464.0005
C121.54062.16672.17382.18203.38093.74904.35762.96022.89843.82563.03251.08901.81752.77691.0899
H132.15232.47832.48223.06503.12333.40714.09812.44533.16574.16243.41741.08902.38743.66891.7712
S142.84303.78883.07333.05702.71832.85023.69652.93371.81152.37062.43491.81752.38741.56662.3647
O153.42964.38933.84223.11334.06214.21714.89794.40112.77082.90083.10462.77693.66891.56662.9072
H162.16842.49143.07152.52394.37064.58855.38723.97543.82074.63384.00051.08991.77122.36472.9072

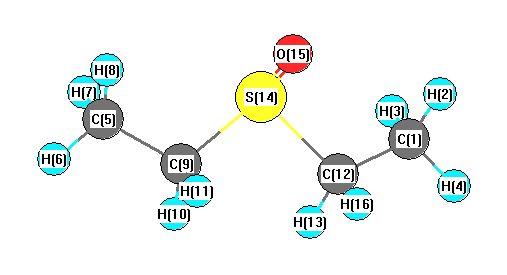
picture of Diethyl sulfoxide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C12 H13 108.651 C1 C12 S14 115.449
C1 C12 H16 109.859 H2 C1 H3 108.289
H2 C1 H4 108.820 H2 C1 C12 109.561
H3 C1 H4 109.086 H3 C1 C12 110.140
H4 C1 C12 110.893 C5 C9 H10 109.721
C5 C9 H11 111.189 C5 C9 S14 108.569
H6 C5 H7 108.982 H6 C5 H8 108.870
H6 C5 C9 108.864 H7 C5 H8 109.239
H7 C5 C9 110.115 H8 C5 C9 110.738
C9 S14 C12 106.006 C9 S14 O15 110.006
H10 C9 H11 108.846 H10 C9 S14 107.002
H11 C9 S14 111.418 C12 S14 O15 110.066
H13 C12 S14 107.808 H13 C12 H16 108.754
S14 C12 H16 106.129
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.481      
2 H 0.169      
3 H 0.156      
4 H 0.211      
5 C -0.491      
6 H 0.175      
7 H 0.178      
8 H 0.166      
9 C -0.494      
10 H 0.202      
11 H 0.202      
12 C -0.498      
13 H 0.179      
14 S 0.845      
15 O -0.719      
16 H 0.199      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 10.454 -0.008 -0.297
y -0.008 9.449 -0.305
z -0.297 -0.305 7.980


<r2> (average value of r2) Å2
<r2> 224.102
(<r2>)1/2 14.970