Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -193.639552 |
| Energy at 298.15K | -193.648626 |
| Nuclear repulsion energy | 130.873702 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3933 |
3633 |
28.74 |
|
|
|
| 2 |
A' |
3234 |
2987 |
33.05 |
|
|
|
| 3 |
A' |
3168 |
2926 |
22.32 |
|
|
|
| 4 |
A' |
3150 |
2910 |
28.45 |
|
|
|
| 5 |
A' |
3105 |
2868 |
52.70 |
|
|
|
| 6 |
A' |
1626 |
1502 |
3.66 |
|
|
|
| 7 |
A' |
1602 |
1480 |
6.09 |
|
|
|
| 8 |
A' |
1587 |
1466 |
0.93 |
|
|
|
| 9 |
A' |
1554 |
1436 |
4.56 |
|
|
|
| 10 |
A' |
1512 |
1396 |
1.39 |
|
|
|
| 11 |
A' |
1431 |
1322 |
11.57 |
|
|
|
| 12 |
A' |
1330 |
1229 |
72.10 |
|
|
|
| 13 |
A' |
1169 |
1079 |
25.88 |
|
|
|
| 14 |
A' |
1143 |
1056 |
64.99 |
|
|
|
| 15 |
A' |
1105 |
1021 |
4.92 |
|
|
|
| 16 |
A' |
939 |
867 |
5.56 |
|
|
|
| 17 |
A' |
478 |
442 |
11.21 |
|
|
|
| 18 |
A' |
289 |
267 |
4.96 |
|
|
|
| 19 |
A" |
3228 |
2982 |
70.91 |
|
|
|
| 20 |
A" |
3206 |
2961 |
1.47 |
|
|
|
| 21 |
A" |
3139 |
2900 |
56.33 |
|
|
|
| 22 |
A" |
1593 |
1471 |
7.08 |
|
|
|
| 23 |
A" |
1391 |
1285 |
0.26 |
|
|
|
| 24 |
A" |
1340 |
1238 |
0.15 |
|
|
|
| 25 |
A" |
1257 |
1162 |
2.24 |
|
|
|
| 26 |
A" |
950 |
877 |
3.05 |
|
|
|
| 27 |
A" |
801 |
740 |
0.37 |
|
|
|
| 28 |
A" |
297 |
274 |
136.51 |
|
|
|
| 29 |
A" |
244 |
226 |
4.36 |
|
|
|
| 30 |
A" |
136 |
125 |
5.92 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24968.3 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 23063.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.442 |
1.228 |
0.000 |
| C2 |
0.000 |
0.736 |
0.000 |
| C3 |
0.096 |
-0.775 |
0.000 |
| O4 |
1.468 |
-1.123 |
0.000 |
| H5 |
-1.484 |
2.316 |
0.000 |
| H6 |
-1.980 |
0.875 |
0.881 |
| H7 |
-1.980 |
0.875 |
-0.881 |
| H8 |
0.531 |
1.113 |
0.875 |
| H9 |
0.531 |
1.113 |
-0.875 |
| H10 |
-0.410 |
-1.181 |
0.883 |
| H11 |
-0.410 |
-1.181 |
-0.883 |
| H12 |
1.542 |
-2.081 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5239 | 2.5253 | 3.7410 | 1.0892 | 1.0908 | 1.0908 | 2.1617 | 2.1617 | 2.7651 | 2.7651 | 4.4552 |
C2 | 1.5239 | | 1.5144 | 2.3687 | 2.1678 | 2.1720 | 2.1720 | 1.0907 | 1.0907 | 2.1500 | 2.1500 | 3.2110 | C3 | 2.5253 | 1.5144 | | 1.4158 | 3.4716 | 2.7947 | 2.7947 | 2.1259 | 2.1259 | 1.0956 | 1.0956 | 1.9481 | O4 | 3.7410 | 2.3687 | 1.4158 | | 4.5324 | 4.0817 | 4.0817 | 2.5771 | 2.5771 | 2.0763 | 2.0763 | 0.9606 | H5 | 1.0892 | 2.1678 | 3.4716 | 4.5324 | | 1.7600 | 1.7600 | 2.5051 | 2.5051 | 3.7631 | 3.7631 | 5.3373 | H6 | 1.0908 | 2.1720 | 2.7947 | 4.0817 | 1.7600 | | 1.7624 | 2.5222 | 3.0736 | 2.5871 | 3.1315 | 4.6817 | H7 | 1.0908 | 2.1720 | 2.7947 | 4.0817 | 1.7600 | 1.7624 | | 3.0736 | 2.5222 | 3.1315 | 2.5871 | 4.6817 | H8 | 2.1617 | 1.0907 | 2.1259 | 2.5771 | 2.5051 | 2.5222 | 3.0736 | | 1.7506 | 2.4787 | 3.0392 | 3.4618 | H9 | 2.1617 | 1.0907 | 2.1259 | 2.5771 | 2.5051 | 3.0736 | 2.5222 | 1.7506 | | 3.0392 | 2.4787 | 3.4618 | H10 | 2.7651 | 2.1500 | 1.0956 | 2.0763 | 3.7631 | 2.5871 | 3.1315 | 2.4787 | 3.0392 | | 1.7665 | 2.3237 | H11 | 2.7651 | 2.1500 | 1.0956 | 2.0763 | 3.7631 | 3.1315 | 2.5871 | 3.0392 | 2.4787 | 1.7665 | | 2.3237 | H12 | 4.4552 | 3.2110 | 1.9481 | 0.9606 | 5.3373 | 4.6817 | 4.6817 | 3.4618 | 3.4618 | 2.3237 | 2.3237 | |
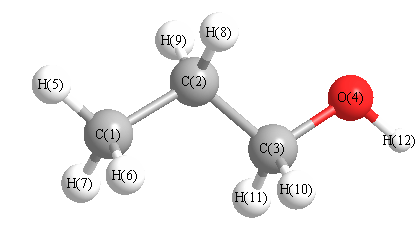 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.432 |
|
C1 |
C2 |
H8 |
110.445 |
| C1 |
C2 |
H9 |
110.445 |
|
C2 |
C1 |
H5 |
111.026 |
| C2 |
C1 |
H6 |
111.259 |
|
C2 |
C1 |
H7 |
111.259 |
| C2 |
C3 |
O4 |
107.830 |
|
C2 |
C3 |
H10 |
109.889 |
| C2 |
C3 |
H11 |
109.889 |
|
C3 |
C2 |
H8 |
108.286 |
| C3 |
C2 |
H9 |
108.286 |
|
C3 |
O4 |
H12 |
108.609 |
| O4 |
C3 |
H10 |
110.891 |
|
O4 |
C3 |
H11 |
110.891 |
| H5 |
C1 |
H6 |
107.677 |
|
H5 |
C1 |
H7 |
107.677 |
| H6 |
C1 |
H7 |
107.770 |
|
H8 |
C2 |
H9 |
106.746 |
| H10 |
C3 |
H11 |
107.459 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -193.639991 |
| Energy at 298.15K | |
| HF Energy | -193.110169 |
| Nuclear repulsion energy | 133.218095 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3936 |
3635 |
31.21 |
|
|
|
| 2 |
A |
3256 |
3008 |
23.78 |
|
|
|
| 3 |
A |
3225 |
2979 |
49.97 |
|
|
|
| 4 |
A |
3199 |
2955 |
22.44 |
|
|
|
| 5 |
A |
3159 |
2918 |
37.29 |
|
|
|
| 6 |
A |
3154 |
2913 |
57.89 |
|
|
|
| 7 |
A |
3143 |
2903 |
28.17 |
|
|
|
| 8 |
A |
3108 |
2871 |
56.75 |
|
|
|
| 9 |
A |
1623 |
1499 |
2.54 |
|
|
|
| 10 |
A |
1601 |
1479 |
8.45 |
|
|
|
| 11 |
A |
1588 |
1467 |
6.72 |
|
|
|
| 12 |
A |
1577 |
1456 |
1.48 |
|
|
|
| 13 |
A |
1546 |
1428 |
5.60 |
|
|
|
| 14 |
A |
1511 |
1396 |
3.85 |
|
|
|
| 15 |
A |
1463 |
1352 |
1.44 |
|
|
|
| 16 |
A |
1397 |
1291 |
27.39 |
|
|
|
| 17 |
A |
1340 |
1237 |
0.66 |
|
|
|
| 18 |
A |
1321 |
1220 |
55.15 |
|
|
|
| 19 |
A |
1232 |
1138 |
9.70 |
|
|
|
| 20 |
A |
1193 |
1102 |
12.04 |
|
|
|
| 21 |
A |
1138 |
1051 |
44.28 |
|
|
|
| 22 |
A |
1051 |
970 |
31.99 |
|
|
|
| 23 |
A |
978 |
903 |
2.81 |
|
|
|
| 24 |
A |
917 |
847 |
1.77 |
|
|
|
| 25 |
A |
816 |
754 |
0.55 |
|
|
|
| 26 |
A |
503 |
464 |
8.91 |
|
|
|
| 27 |
A |
346 |
319 |
14.71 |
|
|
|
| 28 |
A |
280 |
259 |
125.68 |
|
|
|
| 29 |
A |
243 |
224 |
3.74 |
|
|
|
| 30 |
A |
152 |
140 |
7.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24996.5 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 23089.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.528 |
-0.516 |
0.128 |
| C2 |
-0.631 |
0.644 |
-0.287 |
| C3 |
0.766 |
0.544 |
0.290 |
| O4 |
1.367 |
-0.635 |
-0.216 |
| H5 |
-2.523 |
-0.415 |
-0.305 |
| H6 |
-1.639 |
-0.553 |
1.213 |
| H7 |
-1.106 |
-1.463 |
-0.199 |
| H8 |
-0.549 |
0.682 |
-1.374 |
| H9 |
-1.068 |
1.592 |
0.033 |
| H10 |
0.715 |
0.513 |
1.384 |
| H11 |
1.351 |
1.426 |
0.009 |
| H12 |
2.238 |
-0.730 |
0.177 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5236 | 2.5324 | 2.9175 | 1.0898 | 1.0912 | 1.0875 | 2.1564 | 2.1600 | 2.7691 | 3.4742 | 3.7723 |
C2 | 1.5236 | | 1.5152 | 2.3732 | 2.1677 | 2.1674 | 2.1611 | 1.0911 | 1.0924 | 2.1500 | 2.1508 | 3.2144 | C3 | 2.5324 | 1.5152 | | 1.4170 | 3.4773 | 2.7998 | 2.7877 | 2.1259 | 2.1282 | 1.0956 | 1.0946 | 1.9495 | O4 | 2.9175 | 2.3732 | 1.4170 | | 3.8966 | 3.3290 | 2.6071 | 2.5971 | 3.3094 | 2.0751 | 2.0738 | 0.9605 | H5 | 1.0898 | 2.1677 | 3.4773 | 3.8966 | | 1.7623 | 1.7656 | 2.4983 | 2.5022 | 3.7682 | 4.2998 | 4.7954 | H6 | 1.0912 | 2.1674 | 2.7998 | 3.3290 | 1.7623 | | 1.7629 | 3.0671 | 2.5142 | 2.5897 | 3.7816 | 4.0168 | H7 | 1.0875 | 2.1611 | 2.7877 | 2.6071 | 1.7656 | 1.7629 | | 2.5078 | 3.0644 | 3.1191 | 3.7975 | 3.4437 | H8 | 2.1564 | 1.0911 | 2.1259 | 2.5971 | 2.4983 | 3.0671 | 2.5078 | | 1.7550 | 3.0390 | 2.4648 | 3.4876 | H9 | 2.1600 | 1.0924 | 2.1282 | 3.3094 | 2.5022 | 2.5142 | 3.0644 | 1.7550 | | 2.4836 | 2.4240 | 4.0423 | H10 | 2.7691 | 2.1500 | 1.0956 | 2.0751 | 3.7682 | 2.5897 | 3.1191 | 3.0390 | 2.4836 | | 1.7684 | 2.3070 | H11 | 3.4742 | 2.1508 | 1.0946 | 2.0738 | 4.2998 | 3.7816 | 3.7975 | 2.4648 | 2.4240 | 1.7684 | | 2.3371 | H12 | 3.7723 | 3.2144 | 1.9495 | 0.9605 | 4.7954 | 4.0168 | 3.4437 | 3.4876 | 4.0423 | 2.3070 | 2.3371 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.888 |
|
C1 |
C2 |
H8 |
110.025 |
| C1 |
C2 |
H9 |
110.231 |
|
C2 |
C1 |
H5 |
111.003 |
| C2 |
C1 |
H6 |
110.891 |
|
C2 |
C1 |
H7 |
110.609 |
| C2 |
C3 |
O4 |
108.019 |
|
C2 |
C3 |
H10 |
109.825 |
| C2 |
C3 |
H11 |
109.951 |
|
C3 |
C2 |
H8 |
108.208 |
| C3 |
C2 |
H9 |
108.310 |
|
C3 |
O4 |
H12 |
108.639 |
| O4 |
C3 |
H10 |
110.706 |
|
O4 |
C3 |
H11 |
110.656 |
| H5 |
C1 |
H6 |
107.815 |
|
H5 |
C1 |
H7 |
108.378 |
| H6 |
C1 |
H7 |
108.031 |
|
H8 |
C2 |
H9 |
106.978 |
| H10 |
C3 |
H11 |
107.687 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
