Jump to
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -194.112449 |
| Energy at 298.15K | -194.121403 |
| HF Energy | -193.924700 |
| Nuclear repulsion energy | 130.322753 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3779 |
3587 |
14.36 |
|
|
|
| 2 |
A' |
3159 |
2998 |
33.27 |
|
|
|
| 3 |
A' |
3090 |
2933 |
28.22 |
|
|
|
| 4 |
A' |
3078 |
2922 |
24.40 |
|
|
|
| 5 |
A' |
3015 |
2862 |
59.10 |
|
|
|
| 6 |
A' |
1580 |
1500 |
3.95 |
|
|
|
| 7 |
A' |
1559 |
1480 |
4.55 |
|
|
|
| 8 |
A' |
1544 |
1466 |
0.79 |
|
|
|
| 9 |
A' |
1498 |
1422 |
6.06 |
|
|
|
| 10 |
A' |
1461 |
1387 |
0.87 |
|
|
|
| 11 |
A' |
1383 |
1312 |
7.24 |
|
|
|
| 12 |
A' |
1288 |
1223 |
60.73 |
|
|
|
| 13 |
A' |
1122 |
1065 |
3.12 |
|
|
|
| 14 |
A' |
1091 |
1035 |
78.37 |
|
|
|
| 15 |
A' |
1063 |
1009 |
17.24 |
|
|
|
| 16 |
A' |
908 |
862 |
5.58 |
|
|
|
| 17 |
A' |
462 |
439 |
10.73 |
|
|
|
| 18 |
A' |
277 |
263 |
4.39 |
|
|
|
| 19 |
A" |
3152 |
2992 |
69.47 |
|
|
|
| 20 |
A" |
3128 |
2969 |
3.20 |
|
|
|
| 21 |
A" |
3046 |
2891 |
62.60 |
|
|
|
| 22 |
A" |
1552 |
1473 |
6.51 |
|
|
|
| 23 |
A" |
1349 |
1281 |
0.30 |
|
|
|
| 24 |
A" |
1296 |
1230 |
0.03 |
|
|
|
| 25 |
A" |
1215 |
1153 |
1.58 |
|
|
|
| 26 |
A" |
922 |
875 |
2.93 |
|
|
|
| 27 |
A" |
782 |
743 |
0.55 |
|
|
|
| 28 |
A" |
288 |
273 |
125.42 |
|
|
|
| 29 |
A" |
237 |
225 |
3.25 |
|
|
|
| 30 |
A" |
129 |
122 |
5.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24226.3 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 22995.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.446 |
1.232 |
0.000 |
| C2 |
0.000 |
0.738 |
0.000 |
| C3 |
0.095 |
-0.778 |
0.000 |
| O4 |
1.475 |
-1.127 |
0.000 |
| H5 |
-1.486 |
2.324 |
0.000 |
| H6 |
-1.987 |
0.880 |
0.884 |
| H7 |
-1.987 |
0.880 |
-0.884 |
| H8 |
0.533 |
1.116 |
0.879 |
| H9 |
0.533 |
1.116 |
-0.879 |
| H10 |
-0.418 |
-1.182 |
0.887 |
| H11 |
-0.418 |
-1.182 |
-0.887 |
| H12 |
1.536 |
-2.093 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5278 | 2.5323 | 3.7540 | 1.0929 | 1.0947 | 1.0947 | 2.1678 | 2.1678 | 2.7695 | 2.7695 | 4.4660 |
C2 | 1.5278 | | 1.5187 | 2.3773 | 2.1737 | 2.1790 | 2.1790 | 1.0948 | 1.0948 | 2.1554 | 2.1554 | 3.2205 | C3 | 2.5323 | 1.5187 | | 1.4234 | 3.4817 | 2.8036 | 2.8036 | 2.1331 | 2.1331 | 1.1008 | 1.1008 | 1.9509 | O4 | 3.7540 | 2.3773 | 1.4234 | | 4.5471 | 4.0972 | 4.0972 | 2.5863 | 2.5863 | 2.0905 | 2.0905 | 0.9681 | H5 | 1.0929 | 2.1737 | 3.4817 | 4.5471 | | 1.7661 | 1.7661 | 2.5115 | 2.5115 | 3.7710 | 3.7710 | 5.3520 | H6 | 1.0947 | 2.1790 | 2.8036 | 4.0972 | 1.7661 | | 1.7678 | 2.5304 | 3.0837 | 2.5906 | 3.1378 | 4.6930 | H7 | 1.0947 | 2.1790 | 2.8036 | 4.0972 | 1.7661 | 1.7678 | | 3.0837 | 2.5304 | 3.1378 | 2.5906 | 4.6930 | H8 | 2.1678 | 1.0948 | 2.1331 | 2.5863 | 2.5115 | 2.5304 | 3.0837 | | 1.7572 | 2.4866 | 3.0494 | 3.4749 | H9 | 2.1678 | 1.0948 | 2.1331 | 2.5863 | 2.5115 | 3.0837 | 2.5304 | 1.7572 | | 3.0494 | 2.4866 | 3.4749 | H10 | 2.7695 | 2.1554 | 1.1008 | 2.0905 | 3.7710 | 2.5906 | 3.1378 | 2.4866 | 3.0494 | | 1.7731 | 2.3307 | H11 | 2.7695 | 2.1554 | 1.1008 | 2.0905 | 3.7710 | 3.1378 | 2.5906 | 3.0494 | 2.4866 | 1.7731 | | 2.3307 | H12 | 4.4660 | 3.2205 | 1.9509 | 0.9681 | 5.3520 | 4.6930 | 4.6930 | 3.4749 | 3.4749 | 2.3307 | 2.3307 | |
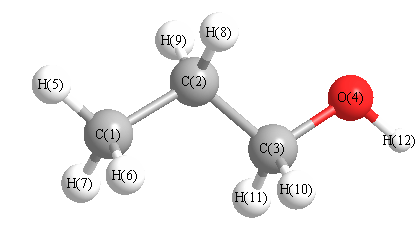 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.451 |
|
C1 |
C2 |
H8 |
110.406 |
| C1 |
C2 |
H9 |
110.406 |
|
C2 |
C1 |
H5 |
110.996 |
| C2 |
C1 |
H6 |
111.311 |
|
C2 |
C1 |
H7 |
111.311 |
| C2 |
C3 |
O4 |
107.763 |
|
C2 |
C3 |
H10 |
109.712 |
| C2 |
C3 |
H11 |
109.712 |
|
C3 |
C2 |
H8 |
108.319 |
| C3 |
C2 |
H9 |
108.319 |
|
C3 |
O4 |
H12 |
107.804 |
| O4 |
C3 |
H10 |
111.183 |
|
O4 |
C3 |
H11 |
111.183 |
| H5 |
C1 |
H6 |
107.672 |
|
H5 |
C1 |
H7 |
107.672 |
| H6 |
C1 |
H7 |
107.701 |
|
H8 |
C2 |
H9 |
106.741 |
| H10 |
C3 |
H11 |
107.287 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.468 |
|
|
|
| 2 |
C |
-0.275 |
|
|
|
| 3 |
C |
-0.024 |
|
|
|
| 4 |
O |
-0.643 |
|
|
|
| 5 |
H |
0.157 |
|
|
|
| 6 |
H |
0.149 |
|
|
|
| 7 |
H |
0.149 |
|
|
|
| 8 |
H |
0.156 |
|
|
|
| 9 |
H |
0.156 |
|
|
|
| 10 |
H |
0.122 |
|
|
|
| 11 |
H |
0.122 |
|
|
|
| 12 |
H |
0.402 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.144 |
-1.024 |
0.000 |
1.536 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.348 |
-1.053 |
0.000 |
| y |
-1.053 |
-21.625 |
0.000 |
| z |
0.000 |
0.000 |
-26.569 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.251 |
-1.053 |
0.000 |
| y |
-1.053 |
6.333 |
0.000 |
| z |
0.000 |
0.000 |
-1.083 |
|
| Polar |
|---|
| 3z2-r2 | -2.165 |
|---|
| x2-y2 | -7.722 |
|---|
| xy | -1.053 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.544 |
-0.499 |
0.000 |
| y |
-0.499 |
5.742 |
0.000 |
| z |
0.000 |
0.000 |
4.835 |
<r2> (average value of r
2) Å
2
| <r2> |
107.102 |
| (<r2>)1/2 |
10.349 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -194.112874 |
| Energy at 298.15K | |
| HF Energy | -193.924888 |
| Nuclear repulsion energy | 132.656484 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3781 |
3589 |
16.50 |
115.00 |
0.31 |
0.47 |
| 2 |
A |
3179 |
3018 |
24.03 |
38.82 |
0.73 |
0.85 |
| 3 |
A |
3149 |
2989 |
48.76 |
54.50 |
0.61 |
0.76 |
| 4 |
A |
3119 |
2961 |
23.93 |
79.11 |
0.75 |
0.85 |
| 5 |
A |
3081 |
2925 |
13.73 |
152.01 |
0.02 |
0.03 |
| 6 |
A |
3079 |
2923 |
60.11 |
49.68 |
0.25 |
0.40 |
| 7 |
A |
3051 |
2896 |
55.19 |
81.70 |
0.75 |
0.86 |
| 8 |
A |
3018 |
2865 |
62.92 |
91.97 |
0.13 |
0.24 |
| 9 |
A |
1574 |
1494 |
2.82 |
7.07 |
0.67 |
0.81 |
| 10 |
A |
1561 |
1482 |
6.80 |
8.88 |
0.75 |
0.86 |
| 11 |
A |
1546 |
1468 |
6.63 |
21.88 |
0.75 |
0.86 |
| 12 |
A |
1530 |
1453 |
1.86 |
18.84 |
0.75 |
0.86 |
| 13 |
A |
1491 |
1416 |
7.12 |
2.67 |
0.51 |
0.68 |
| 14 |
A |
1458 |
1384 |
2.74 |
3.66 |
0.74 |
0.85 |
| 15 |
A |
1416 |
1344 |
2.47 |
1.21 |
0.75 |
0.85 |
| 16 |
A |
1356 |
1287 |
21.02 |
17.11 |
0.73 |
0.85 |
| 17 |
A |
1295 |
1229 |
1.15 |
8.86 |
0.75 |
0.86 |
| 18 |
A |
1277 |
1212 |
44.79 |
7.58 |
0.71 |
0.83 |
| 19 |
A |
1190 |
1129 |
6.50 |
1.72 |
0.45 |
0.62 |
| 20 |
A |
1144 |
1086 |
6.55 |
3.63 |
0.57 |
0.72 |
| 21 |
A |
1092 |
1037 |
45.51 |
3.14 |
0.67 |
0.80 |
| 22 |
A |
1008 |
957 |
42.62 |
4.77 |
0.73 |
0.85 |
| 23 |
A |
949 |
900 |
2.92 |
0.57 |
0.74 |
0.85 |
| 24 |
A |
885 |
841 |
1.61 |
9.90 |
0.18 |
0.31 |
| 25 |
A |
791 |
751 |
0.71 |
0.64 |
0.52 |
0.68 |
| 26 |
A |
486 |
462 |
8.24 |
0.26 |
0.50 |
0.67 |
| 27 |
A |
335 |
318 |
12.64 |
0.59 |
0.59 |
0.74 |
| 28 |
A |
268 |
255 |
114.85 |
4.59 |
0.74 |
0.85 |
| 29 |
A |
233 |
221 |
2.53 |
0.05 |
0.63 |
0.77 |
| 30 |
A |
149 |
141 |
8.06 |
0.11 |
0.75 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 24246.6 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 23014.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.533 |
-0.518 |
0.130 |
| C2 |
-0.634 |
0.645 |
-0.289 |
| C3 |
0.769 |
0.548 |
0.291 |
| O4 |
1.372 |
-0.640 |
-0.218 |
| H5 |
-2.529 |
-0.425 |
-0.314 |
| H6 |
-1.653 |
-0.548 |
1.218 |
| H7 |
-1.102 |
-1.469 |
-0.188 |
| H8 |
-0.551 |
0.681 |
-1.380 |
| H9 |
-1.071 |
1.598 |
0.031 |
| H10 |
0.715 |
0.521 |
1.390 |
| H11 |
1.353 |
1.436 |
0.008 |
| H12 |
2.246 |
-0.728 |
0.190 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5283 | 2.5416 | 2.9289 | 1.0936 | 1.0951 | 1.0915 | 2.1637 | 2.1682 | 2.7790 | 3.4871 | 3.7855 |
C2 | 1.5283 | | 1.5203 | 2.3833 | 2.1763 | 2.1749 | 2.1671 | 1.0953 | 1.0964 | 2.1568 | 2.1585 | 3.2258 | C3 | 2.5416 | 1.5203 | | 1.4262 | 3.4905 | 2.8147 | 2.7917 | 2.1333 | 2.1344 | 1.1007 | 1.0997 | 1.9545 | O4 | 2.9289 | 2.3833 | 1.4262 | | 3.9081 | 3.3498 | 2.6094 | 2.6071 | 3.3230 | 2.0892 | 2.0880 | 0.9680 | H5 | 1.0936 | 2.1763 | 3.4905 | 3.9081 | | 1.7685 | 1.7722 | 2.5044 | 2.5173 | 3.7841 | 4.3164 | 4.8106 | H6 | 1.0951 | 2.1749 | 2.8147 | 3.3498 | 1.7685 | | 1.7686 | 3.0779 | 2.5206 | 2.6037 | 3.7988 | 4.0360 | H7 | 1.0915 | 2.1671 | 2.7917 | 2.6094 | 1.7722 | 1.7686 | | 2.5193 | 3.0749 | 3.1225 | 3.8077 | 3.4495 | H8 | 2.1637 | 1.0953 | 2.1333 | 2.6071 | 2.5044 | 3.0779 | 2.5193 | | 1.7611 | 3.0499 | 2.4740 | 3.5034 | H9 | 2.1682 | 1.0964 | 2.1344 | 3.3230 | 2.5173 | 2.5206 | 3.0749 | 1.7611 | | 2.4897 | 2.4295 | 4.0545 | H10 | 2.7790 | 2.1568 | 1.1007 | 2.0892 | 3.7841 | 2.6037 | 3.1225 | 3.0499 | 2.4897 | | 1.7754 | 2.3115 | H11 | 3.4871 | 2.1585 | 1.0997 | 2.0880 | 4.3164 | 3.7988 | 3.8077 | 2.4740 | 2.4295 | 1.7754 | | 2.3475 | H12 | 3.7855 | 3.2258 | 1.9545 | 0.9680 | 4.8106 | 4.0360 | 3.4495 | 3.5034 | 4.0545 | 2.3115 | 2.3475 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.957 |
|
C1 |
C2 |
H8 |
110.018 |
| C1 |
C2 |
H9 |
110.306 |
|
C2 |
C1 |
H5 |
111.123 |
| C2 |
C1 |
H6 |
110.914 |
|
C2 |
C1 |
H7 |
110.519 |
| C2 |
C3 |
O4 |
107.930 |
|
C2 |
C3 |
H10 |
109.709 |
| C2 |
C3 |
H11 |
109.907 |
|
C3 |
C2 |
H8 |
108.198 |
| C3 |
C2 |
H9 |
108.219 |
|
C3 |
O4 |
H12 |
107.905 |
| O4 |
C3 |
H10 |
110.878 |
|
O4 |
C3 |
H11 |
110.844 |
| H5 |
C1 |
H6 |
107.807 |
|
H5 |
C1 |
H7 |
108.397 |
| H6 |
C1 |
H7 |
107.964 |
|
H8 |
C2 |
H9 |
106.932 |
| H10 |
C3 |
H11 |
107.576 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.457 |
|
|
|
| 2 |
C |
-0.277 |
|
|
|
| 3 |
C |
-0.031 |
|
|
|
| 4 |
O |
-0.640 |
|
|
|
| 5 |
H |
0.146 |
|
|
|
| 6 |
H |
0.140 |
|
|
|
| 7 |
H |
0.176 |
|
|
|
| 8 |
H |
0.155 |
|
|
|
| 9 |
H |
0.138 |
|
|
|
| 10 |
H |
0.122 |
|
|
|
| 11 |
H |
0.126 |
|
|
|
| 12 |
H |
0.403 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.789 |
0.893 |
1.004 |
1.558 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.592 |
-0.223 |
2.018 |
| y |
-0.223 |
-27.035 |
-0.565 |
| z |
2.018 |
-0.565 |
-26.719 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.285 |
-0.223 |
2.018 |
| y |
-0.223 |
-2.379 |
-0.565 |
| z |
2.018 |
-0.565 |
-1.906 |
|
| Polar |
|---|
| 3z2-r2 | -3.811 |
|---|
| x2-y2 | 4.443 |
|---|
| xy | -0.223 |
|---|
| xz | 2.018 |
|---|
| yz | -0.565 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.900 |
-0.034 |
0.122 |
| y |
-0.034 |
5.231 |
0.034 |
| z |
0.122 |
0.034 |
4.963 |
<r2> (average value of r
2) Å
2
| <r2> |
94.875 |
| (<r2>)1/2 |
9.740 |
