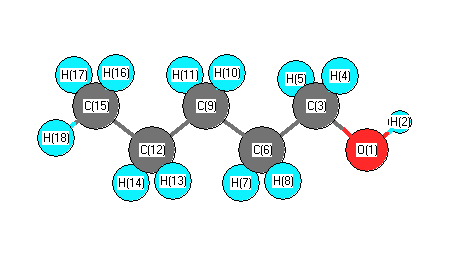
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3794 |
3630 |
14.34 |
|
|
|
| 2 |
A' |
3129 |
2994 |
38.87 |
|
|
|
| 3 |
A' |
3055 |
2922 |
55.63 |
|
|
|
| 4 |
A' |
3050 |
2918 |
33.26 |
|
|
|
| 5 |
A' |
3040 |
2909 |
24.52 |
|
|
|
| 6 |
A' |
3024 |
2893 |
17.01 |
|
|
|
| 7 |
A' |
2979 |
2850 |
58.13 |
|
|
|
| 8 |
A' |
1554 |
1487 |
3.63 |
|
|
|
| 9 |
A' |
1536 |
1469 |
5.85 |
|
|
|
| 10 |
A' |
1526 |
1460 |
0.61 |
|
|
|
| 11 |
A' |
1515 |
1450 |
0.73 |
|
|
|
| 12 |
A' |
1512 |
1447 |
0.14 |
|
|
|
| 13 |
A' |
1479 |
1415 |
8.71 |
|
|
|
| 14 |
A' |
1434 |
1372 |
2.90 |
|
|
|
| 15 |
A' |
1423 |
1361 |
0.38 |
|
|
|
| 16 |
A' |
1371 |
1312 |
12.24 |
|
|
|
| 17 |
A' |
1308 |
1251 |
30.90 |
|
|
|
| 18 |
A' |
1248 |
1194 |
28.09 |
|
|
|
| 19 |
A' |
1140 |
1090 |
6.03 |
|
|
|
| 20 |
A' |
1097 |
1049 |
83.41 |
|
|
|
| 21 |
A' |
1089 |
1042 |
0.39 |
|
|
|
| 22 |
A' |
1056 |
1011 |
0.78 |
|
|
|
| 23 |
A' |
1017 |
973 |
16.32 |
|
|
|
| 24 |
A' |
911 |
872 |
3.88 |
|
|
|
| 25 |
A' |
495 |
474 |
12.28 |
|
|
|
| 26 |
A' |
363 |
348 |
0.06 |
|
|
|
| 27 |
A' |
313 |
300 |
4.59 |
|
|
|
| 28 |
A' |
134 |
128 |
1.53 |
|
|
|
| 29 |
A" |
3124 |
2988 |
66.77 |
|
|
|
| 30 |
A" |
3104 |
2970 |
58.49 |
|
|
|
| 31 |
A" |
3080 |
2947 |
11.65 |
|
|
|
| 32 |
A" |
3055 |
2922 |
3.35 |
|
|
|
| 33 |
A" |
3008 |
2877 |
62.89 |
|
|
|
| 34 |
A" |
1525 |
1459 |
7.36 |
|
|
|
| 35 |
A" |
1347 |
1288 |
0.68 |
|
|
|
| 36 |
A" |
1334 |
1276 |
1.04 |
|
|
|
| 37 |
A" |
1311 |
1254 |
0.06 |
|
|
|
| 38 |
A" |
1248 |
1194 |
0.48 |
|
|
|
| 39 |
A" |
1200 |
1148 |
1.16 |
|
|
|
| 40 |
A" |
998 |
955 |
1.42 |
|
|
|
| 41 |
A" |
868 |
831 |
0.00 |
|
|
|
| 42 |
A" |
779 |
746 |
0.40 |
|
|
|
| 43 |
A" |
741 |
709 |
4.07 |
|
|
|
| 44 |
A" |
289 |
276 |
127.05 |
|
|
|
| 45 |
A" |
249 |
238 |
0.64 |
|
|
|
| 46 |
A" |
152 |
145 |
1.70 |
|
|
|
| 47 |
A" |
100 |
95 |
4.15 |
|
|
|
| 48 |
A" |
76 |
73 |
1.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36588.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 35004.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.630 |
|
|
-0.640 |
| 2 |
H |
0.403 |
|
|
0.407 |
| 3 |
C |
-0.072 |
|
|
0.214 |
| 4 |
H |
0.137 |
|
|
-0.012 |
| 5 |
H |
0.137 |
|
|
-0.012 |
| 6 |
C |
-0.318 |
|
|
0.004 |
| 7 |
H |
0.171 |
|
|
0.039 |
| 8 |
H |
0.171 |
|
|
0.039 |
| 9 |
C |
-0.310 |
|
|
-0.206 |
| 10 |
H |
0.154 |
|
|
0.052 |
| 11 |
H |
0.154 |
|
|
0.052 |
| 12 |
C |
-0.303 |
|
|
0.205 |
| 13 |
H |
0.159 |
|
|
-0.026 |
| 14 |
H |
0.159 |
|
|
-0.026 |
| 15 |
C |
-0.509 |
|
|
-0.307 |
| 16 |
H |
0.164 |
|
|
0.073 |
| 17 |
H |
0.164 |
|
|
0.073 |
| 18 |
H |
0.167 |
|
|
0.072 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.291 |
0.719 |
0.000 |
1.478 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.296 |
0.740 |
0.000 |
1.493 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.101 |
-4.087 |
0.000 |
| y |
-4.087 |
-42.408 |
0.000 |
| z |
0.000 |
0.000 |
-39.154 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.680 |
-4.087 |
0.000 |
| y |
-4.087 |
-5.780 |
0.000 |
| z |
0.000 |
0.000 |
-0.900 |
|
| Polar |
|---|
| 3z2-r2 | -1.799 |
|---|
| x2-y2 | 8.307 |
|---|
| xy | -4.087 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
301.430 |
| (<r2>)1/2 |
17.362 |