return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (1,4-Hexadiene, (Z)-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-234.309125
Energy at 298.15K-234.318779
HF Energy-234.053958
Nuclear repulsion energy217.230608
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3280 3113 17.54      
2 A 3199 3037 7.76      
3 A 3194 3031 44.86      
4 A 3177 3016 39.91      
5 A 3176 3015 7.78      
6 A 3165 3004 1.22      
7 A 3123 2964 22.11      
8 A 3095 2938 21.12      
9 A 3071 2915 22.37      
10 A 3046 2892 28.99      
11 A 1753 1664 1.49      
12 A 1732 1644 11.73      
13 A 1545 1467 7.11      
14 A 1534 1456 4.67      
15 A 1524 1447 4.34      
16 A 1482 1407 2.84      
17 A 1473 1398 4.22      
18 A 1447 1373 3.81      
19 A 1372 1303 6.72      
20 A 1349 1281 1.09      
21 A 1320 1253 0.50      
22 A 1253 1190 1.94      
23 A 1154 1095 1.37      
24 A 1112 1055 9.53      
25 A 1089 1034 0.44      
26 A 1048 995 12.94      
27 A 1023 971 1.48      
28 A 1014 962 0.23      
29 A 982 932 4.82      
30 A 940 893 26.03      
31 A 936 889 8.29      
32 A 894 849 3.31      
33 A 740 702 25.16      
34 A 608 577 5.98      
35 A 570 541 2.10      
36 A 531 504 11.12      
37 A 397 377 0.31      
38 A 297 282 0.01      
39 A 211 201 0.07      
40 A 136 129 0.26      
41 A 128 121 0.11      
42 A 40 38 0.05      

Unscaled Zero Point Vibrational Energy (zpe) 31580.2 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 29975.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.23043 0.06253 0.06078

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.324 0.234 -0.624
H2 -3.264 0.511 -0.145
H3 -1.804 1.143 -0.902
H4 -2.555 -0.321 -1.537
C5 -1.531 -0.597 0.272
H6 -1.959 -1.476 0.577
C7 -0.293 -0.332 0.742
H8 0.117 -1.027 1.367
C9 0.513 0.848 0.452
H10 -0.021 1.522 -0.207
H11 0.724 1.367 1.391
C12 1.787 0.507 -0.186
H13 2.434 1.265 -0.419
C14 2.141 -0.747 -0.478
H15 3.043 -0.932 -0.924
H16 1.522 -1.527 -0.264

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 C7 H8 C9 H10 H11 C12 H13 C14 H15 H16
C11.09101.08341.09281.45762.12092.51293.39413.09652.67193.82624.14384.87314.57455.50064.2455
H21.09101.76181.77052.09972.48423.21374.01143.83943.39784.35885.05165.75445.56026.51705.2035
H31.08341.76181.76372.11653.01082.67583.68072.69971.95063.42003.71624.26684.39505.27224.3120
H41.09281.77051.76372.09722.48113.21104.00933.83873.40384.70914.62205.35274.83285.66414.4377
C51.45762.09972.11652.09721.02381.35012.02492.50972.64573.19303.52664.43453.75074.73923.2357
H62.12092.48423.01082.48111.02382.02772.26703.39543.65473.99324.30705.27324.29655.25093.5820
C72.51293.21372.67583.21101.35012.02771.02091.45842.10062.08432.42743.36702.75433.77662.3943
H83.39414.01143.68074.00932.02492.26701.02092.12372.99932.47042.74863.71672.75303.71712.2098
C93.09653.83942.69973.83872.50973.39541.45842.12371.08361.09391.46442.14982.46143.38502.6773
H102.67193.39781.95063.40382.64573.65472.10062.99931.08361.77002.07332.47733.14603.99023.4173
H113.82624.35883.42004.70913.19303.99322.08432.47041.09391.77002.08732.49273.15834.00303.4282
C124.14385.05163.71624.62203.52664.30702.42742.74861.46442.07332.08731.02341.33602.04782.0528
H134.87315.75444.26685.35274.43455.27323.36703.71672.14982.47732.49271.02342.03452.33512.9412
C144.57455.56024.39504.83283.75074.29652.75432.75302.46143.14603.15831.33602.03451.02271.0183
H155.50066.51705.27225.66414.73925.25093.77663.71713.38503.99024.00302.04782.33511.02271.7614
H164.24555.20354.31204.43773.23573.58202.39432.20982.67733.41733.42822.05282.94121.01831.7614

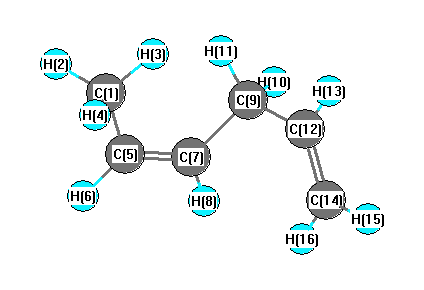
picture of 1,4-Hexadiene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 116.365 C1 C5 C7 126.969
H2 C1 H3 108.241 H2 C1 H4 108.335
H2 C1 C5 110.119 H3 C1 H4 108.279
H3 C1 C5 111.963 H4 C1 C5 109.809
C5 C7 H8 116.620 C5 C7 C9 126.613
H6 C5 C7 116.666 C7 C9 H10 110.593
C7 C9 H11 108.657 C7 C9 C12 112.300
H8 C7 C9 116.767 C9 C12 H13 118.491
C9 C12 C14 122.962 H10 C9 H11 108.752
H10 C9 C12 107.979 H11 C9 C12 108.481
C12 C14 H15 119.911 C12 C14 H16 120.773
H13 C12 C14 118.547 H15 C14 H16 119.316
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.501      
2 H 0.163      
3 H 0.163      
4 H 0.164      
5 C -0.126      
6 H 0.145      
7 C -0.131      
8 H 0.150      
9 C -0.355      
10 H 0.166      
11 H 0.161      
12 C -0.098      
13 H 0.146      
14 C -0.358      
15 H 0.152      
16 H 0.160      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 11.283 0.428 0.176
y 0.428 8.869 -0.460
z 0.176 -0.460 6.956


<r2> (average value of r2) Å2
<r2> 212.360
(<r2>)1/2 14.573