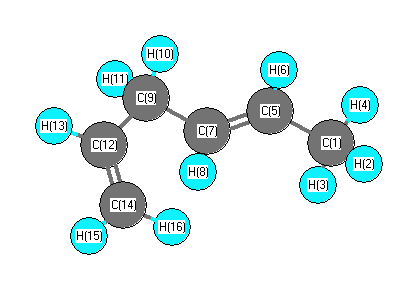
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3270 |
3110 |
17.68 |
|
|
|
| 2 |
A |
3189 |
3032 |
7.80 |
|
|
|
| 3 |
A |
3171 |
3016 |
25.49 |
|
|
|
| 4 |
A |
3168 |
3012 |
56.16 |
|
|
|
| 5 |
A |
3162 |
3007 |
1.04 |
|
|
|
| 6 |
A |
3145 |
2991 |
5.90 |
|
|
|
| 7 |
A |
3113 |
2961 |
20.81 |
|
|
|
| 8 |
A |
3065 |
2915 |
16.95 |
|
|
|
| 9 |
A |
3054 |
2905 |
32.35 |
|
|
|
| 10 |
A |
3027 |
2879 |
30.42 |
|
|
|
| 11 |
A |
1783 |
1696 |
1.17 |
|
|
|
| 12 |
A |
1749 |
1663 |
13.31 |
|
|
|
| 13 |
A |
1520 |
1446 |
9.94 |
|
|
|
| 14 |
A |
1507 |
1433 |
6.70 |
|
|
|
| 15 |
A |
1492 |
1419 |
10.50 |
|
|
|
| 16 |
A |
1462 |
1390 |
2.40 |
|
|
|
| 17 |
A |
1439 |
1369 |
2.71 |
|
|
|
| 18 |
A |
1389 |
1321 |
0.41 |
|
|
|
| 19 |
A |
1347 |
1281 |
1.60 |
|
|
|
| 20 |
A |
1338 |
1273 |
2.24 |
|
|
|
| 21 |
A |
1328 |
1263 |
3.15 |
|
|
|
| 22 |
A |
1237 |
1176 |
1.12 |
|
|
|
| 23 |
A |
1152 |
1095 |
4.39 |
|
|
|
| 24 |
A |
1129 |
1074 |
6.41 |
|
|
|
| 25 |
A |
1078 |
1025 |
0.21 |
|
|
|
| 26 |
A |
1063 |
1010 |
5.27 |
|
|
|
| 27 |
A |
1043 |
992 |
9.44 |
|
|
|
| 28 |
A |
1018 |
968 |
29.38 |
|
|
|
| 29 |
A |
972 |
924 |
13.82 |
|
|
|
| 30 |
A |
945 |
899 |
26.24 |
|
|
|
| 31 |
A |
940 |
894 |
7.19 |
|
|
|
| 32 |
A |
905 |
861 |
3.01 |
|
|
|
| 33 |
A |
786 |
748 |
0.33 |
|
|
|
| 34 |
A |
601 |
572 |
4.79 |
|
|
|
| 35 |
A |
548 |
521 |
6.98 |
|
|
|
| 36 |
A |
470 |
447 |
0.23 |
|
|
|
| 37 |
A |
318 |
302 |
0.35 |
|
|
|
| 38 |
A |
282 |
269 |
1.79 |
|
|
|
| 39 |
A |
216 |
206 |
1.17 |
|
|
|
| 40 |
A |
159 |
151 |
0.62 |
|
|
|
| 41 |
A |
134 |
128 |
0.06 |
|
|
|
| 42 |
A |
77 |
74 |
0.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31395.4 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 29857.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.562 |
|
|
|
| 2 |
H |
0.181 |
|
|
|
| 3 |
H |
0.178 |
|
|
|
| 4 |
H |
0.180 |
|
|
|
| 5 |
C |
-0.124 |
|
|
|
| 6 |
H |
0.151 |
|
|
|
| 7 |
C |
-0.122 |
|
|
|
| 8 |
H |
0.156 |
|
|
|
| 9 |
C |
-0.406 |
|
|
|
| 10 |
H |
0.182 |
|
|
|
| 11 |
H |
0.179 |
|
|
|
| 12 |
C |
-0.097 |
|
|
|
| 13 |
H |
0.155 |
|
|
|
| 14 |
C |
-0.379 |
|
|
|
| 15 |
H |
0.162 |
|
|
|
| 16 |
H |
0.167 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.317 |
-0.278 |
-0.033 |
0.422 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.883 |
1.321 |
-0.423 |
| y |
1.321 |
-37.149 |
0.739 |
| z |
-0.423 |
0.739 |
-38.563 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.972 |
1.321 |
-0.423 |
| y |
1.321 |
0.074 |
0.739 |
| z |
-0.423 |
0.739 |
-2.047 |
|
| Polar |
|---|
| 3z2-r2 | -4.093 |
|---|
| x2-y2 | 1.265 |
|---|
| xy | 1.321 |
|---|
| xz | -0.423 |
|---|
| yz | 0.739 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.983 |
0.327 |
-0.836 |
| y |
0.327 |
9.200 |
0.272 |
| z |
-0.836 |
0.272 |
6.735 |
<r2> (average value of r
2) Å
2
| <r2> |
233.266 |
| (<r2>)1/2 |
15.273 |