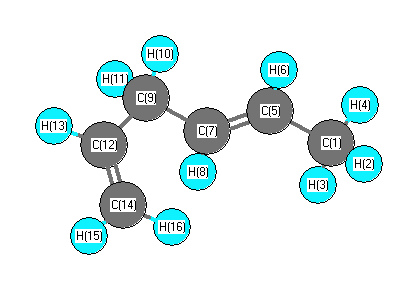
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3176 |
3124 |
18.61 |
|
|
|
| 2 |
A |
3096 |
3045 |
7.03 |
|
|
|
| 3 |
A |
3074 |
3023 |
27.98 |
|
|
|
| 4 |
A |
3071 |
3021 |
64.12 |
|
|
|
| 5 |
A |
3065 |
3014 |
2.26 |
|
|
|
| 6 |
A |
3053 |
3002 |
2.32 |
|
|
|
| 7 |
A |
3018 |
2968 |
20.67 |
|
|
|
| 8 |
A |
2967 |
2918 |
18.68 |
|
|
|
| 9 |
A |
2965 |
2917 |
32.58 |
|
|
|
| 10 |
A |
2930 |
2882 |
31.07 |
|
|
|
| 11 |
A |
1710 |
1682 |
1.24 |
|
|
|
| 12 |
A |
1681 |
1653 |
14.32 |
|
|
|
| 13 |
A |
1474 |
1450 |
9.17 |
|
|
|
| 14 |
A |
1461 |
1437 |
6.24 |
|
|
|
| 15 |
A |
1443 |
1420 |
10.74 |
|
|
|
| 16 |
A |
1414 |
1390 |
1.76 |
|
|
|
| 17 |
A |
1390 |
1367 |
2.22 |
|
|
|
| 18 |
A |
1336 |
1314 |
0.54 |
|
|
|
| 19 |
A |
1304 |
1282 |
1.81 |
|
|
|
| 20 |
A |
1296 |
1274 |
2.94 |
|
|
|
| 21 |
A |
1282 |
1261 |
3.94 |
|
|
|
| 22 |
A |
1195 |
1175 |
1.14 |
|
|
|
| 23 |
A |
1113 |
1095 |
4.77 |
|
|
|
| 24 |
A |
1094 |
1076 |
6.45 |
|
|
|
| 25 |
A |
1037 |
1020 |
0.12 |
|
|
|
| 26 |
A |
1027 |
1010 |
3.85 |
|
|
|
| 27 |
A |
1003 |
986 |
10.31 |
|
|
|
| 28 |
A |
979 |
962 |
26.84 |
|
|
|
| 29 |
A |
939 |
924 |
11.79 |
|
|
|
| 30 |
A |
909 |
894 |
1.82 |
|
|
|
| 31 |
A |
891 |
877 |
28.73 |
|
|
|
| 32 |
A |
872 |
857 |
3.65 |
|
|
|
| 33 |
A |
753 |
741 |
0.26 |
|
|
|
| 34 |
A |
580 |
570 |
4.19 |
|
|
|
| 35 |
A |
529 |
520 |
6.75 |
|
|
|
| 36 |
A |
455 |
448 |
0.20 |
|
|
|
| 37 |
A |
307 |
302 |
0.38 |
|
|
|
| 38 |
A |
274 |
270 |
1.88 |
|
|
|
| 39 |
A |
212 |
208 |
1.25 |
|
|
|
| 40 |
A |
153 |
151 |
0.59 |
|
|
|
| 41 |
A |
133 |
131 |
0.06 |
|
|
|
| 42 |
A |
73 |
71 |
0.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30365.1 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 29864.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.534 |
|
|
|
| 2 |
H |
0.170 |
|
|
|
| 3 |
H |
0.166 |
|
|
|
| 4 |
H |
0.169 |
|
|
|
| 5 |
C |
-0.097 |
|
|
|
| 6 |
H |
0.129 |
|
|
|
| 7 |
C |
-0.091 |
|
|
|
| 8 |
H |
0.132 |
|
|
|
| 9 |
C |
-0.385 |
|
|
|
| 10 |
H |
0.167 |
|
|
|
| 11 |
H |
0.165 |
|
|
|
| 12 |
C |
-0.069 |
|
|
|
| 13 |
H |
0.132 |
|
|
|
| 14 |
C |
-0.347 |
|
|
|
| 15 |
H |
0.144 |
|
|
|
| 16 |
H |
0.149 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.351 |
-0.293 |
-0.027 |
0.458 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.952 |
1.346 |
-0.409 |
| y |
1.346 |
-37.241 |
0.694 |
| z |
-0.409 |
0.694 |
-38.659 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.998 |
1.346 |
-0.409 |
| y |
1.346 |
0.064 |
0.694 |
| z |
-0.409 |
0.694 |
-2.063 |
|
| Polar |
|---|
| 3z2-r2 | -4.126 |
|---|
| x2-y2 | 1.289 |
|---|
| xy | 1.346 |
|---|
| xz | -0.409 |
|---|
| yz | 0.694 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.577 |
0.343 |
-0.798 |
| y |
0.343 |
9.417 |
0.303 |
| z |
-0.798 |
0.303 |
6.873 |
<r2> (average value of r
2) Å
2
| <r2> |
236.697 |
| (<r2>)1/2 |
15.385 |