Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -79.502610 |
| Energy at 298.15K | -79.508572 |
| HF Energy | -79.228573 |
| Nuclear repulsion energy | 42.253865 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3128 |
2890 |
0.00 |
|
|
|
| 2 |
A1g |
1510 |
1395 |
0.00 |
|
|
|
| 3 |
A1g |
1054 |
974 |
0.00 |
|
|
|
| 4 |
A1u |
325 |
301 |
0.00 |
|
|
|
| 5 |
A2u |
3125 |
2886 |
51.25 |
|
|
|
| 6 |
A2u |
1480 |
1367 |
1.61 |
|
|
|
| 7 |
Eg |
3189 |
2946 |
0.00 |
|
|
|
| 7 |
Eg |
3189 |
2946 |
0.00 |
|
|
|
| 8 |
Eg |
1577 |
1456 |
0.00 |
|
|
|
| 8 |
Eg |
1577 |
1456 |
0.00 |
|
|
|
| 9 |
Eg |
1282 |
1184 |
0.00 |
|
|
|
| 9 |
Eg |
1282 |
1184 |
0.00 |
|
|
|
| 10 |
Eu |
3211 |
2966 |
65.72 |
|
|
|
| 10 |
Eu |
3211 |
2966 |
65.73 |
|
|
|
| 11 |
Eu |
1582 |
1461 |
8.37 |
|
|
|
| 11 |
Eu |
1582 |
1461 |
8.37 |
|
|
|
| 12 |
Eu |
856 |
791 |
4.04 |
|
|
|
| 12 |
Eu |
856 |
791 |
4.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17008.2 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 15710.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.763 |
| C2 |
0.000 |
0.000 |
-0.763 |
| H3 |
0.000 |
1.018 |
1.158 |
| H4 |
-0.882 |
-0.509 |
1.158 |
| H5 |
0.882 |
-0.509 |
1.158 |
| H6 |
0.000 |
-1.018 |
-1.158 |
| H7 |
-0.882 |
0.509 |
-1.158 |
| H8 |
0.882 |
0.509 |
-1.158 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5261 | 1.0923 | 1.0923 | 1.0923 | 2.1744 | 2.1744 | 2.1744 |
C2 | 1.5261 | | 2.1744 | 2.1744 | 2.1744 | 1.0923 | 1.0923 | 1.0923 | H3 | 1.0923 | 2.1744 | | 1.7639 | 1.7639 | 3.0844 | 2.5302 | 2.5302 | H4 | 1.0923 | 2.1744 | 1.7639 | | 1.7639 | 2.5302 | 2.5302 | 3.0844 | H5 | 1.0923 | 2.1744 | 1.7639 | 1.7639 | | 2.5302 | 3.0844 | 2.5302 | H6 | 2.1744 | 1.0923 | 3.0844 | 2.5302 | 2.5302 | | 1.7639 | 1.7639 | H7 | 2.1744 | 1.0923 | 2.5302 | 2.5302 | 3.0844 | 1.7639 | | 1.7639 | H8 | 2.1744 | 1.0923 | 2.5302 | 3.0844 | 2.5302 | 1.7639 | 1.7639 | |
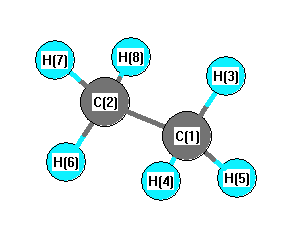 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.201 |
|
C1 |
C2 |
H7 |
111.201 |
| C1 |
C2 |
H8 |
111.201 |
|
C2 |
C1 |
H3 |
111.201 |
| C2 |
C1 |
H4 |
111.201 |
|
C2 |
C1 |
H5 |
111.201 |
| H3 |
C1 |
H4 |
107.687 |
|
H3 |
C1 |
H5 |
107.687 |
| H4 |
C1 |
H5 |
107.687 |
|
H6 |
C2 |
H7 |
107.687 |
| H6 |
C2 |
H8 |
107.687 |
|
H7 |
C2 |
H8 |
107.687 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -79.497793 |
| Energy at 298.15K | |
| HF Energy | -79.223812 |
| Nuclear repulsion energy | 42.089475 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3141 |
2901 |
0.00 |
|
|
|
| 2 |
A1' |
1530 |
1413 |
0.00 |
|
|
|
| 3 |
A1' |
1052 |
972 |
0.00 |
|
|
|
| 4 |
A1" |
316i |
292i |
0.00 |
|
|
|
| 5 |
A2" |
3131 |
2892 |
54.03 |
|
|
|
| 6 |
A2" |
1480 |
1367 |
1.26 |
|
|
|
| 7 |
E' |
3216 |
2971 |
56.73 |
|
|
|
| 7 |
E' |
3216 |
2971 |
56.73 |
|
|
|
| 8 |
E' |
1588 |
1467 |
9.39 |
|
|
|
| 8 |
E' |
1588 |
1467 |
9.39 |
|
|
|
| 9 |
E' |
941 |
869 |
3.69 |
|
|
|
| 9 |
E' |
941 |
869 |
3.69 |
|
|
|
| 10 |
E" |
3196 |
2953 |
0.00 |
|
|
|
| 10 |
E" |
3196 |
2953 |
0.00 |
|
|
|
| 11 |
E" |
1574 |
1454 |
0.00 |
|
|
|
| 11 |
E" |
1574 |
1454 |
0.00 |
|
|
|
| 12 |
E" |
1231 |
1137 |
0.00 |
|
|
|
| 12 |
E" |
1231 |
1137 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16754.7 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 15476.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.770 |
| C2 |
0.000 |
0.000 |
-0.770 |
| H3 |
0.000 |
1.014 |
1.173 |
| H4 |
-0.878 |
-0.507 |
1.173 |
| H5 |
0.878 |
-0.507 |
1.173 |
| H6 |
0.000 |
1.014 |
-1.173 |
| H7 |
0.878 |
-0.507 |
-1.173 |
| H8 |
-0.878 |
-0.507 |
-1.173 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5396 | 1.0914 | 1.0914 | 1.0914 | 2.1915 | 2.1915 | 2.1915 |
C2 | 1.5396 | | 2.1915 | 2.1915 | 2.1915 | 1.0914 | 1.0914 | 1.0914 | H3 | 1.0914 | 2.1915 | | 1.7566 | 1.7566 | 2.3459 | 2.9307 | 2.9307 | H4 | 1.0914 | 2.1915 | 1.7566 | | 1.7566 | 2.9307 | 2.9307 | 2.3459 | H5 | 1.0914 | 2.1915 | 1.7566 | 1.7566 | | 2.9307 | 2.3459 | 2.9307 | H6 | 2.1915 | 1.0914 | 2.3459 | 2.9307 | 2.9307 | | 1.7566 | 1.7566 | H7 | 2.1915 | 1.0914 | 2.9307 | 2.9307 | 2.3459 | 1.7566 | | 1.7566 | H8 | 2.1915 | 1.0914 | 2.9307 | 2.3459 | 2.9307 | 1.7566 | 1.7566 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.679 |
|
C1 |
C2 |
H7 |
111.679 |
| C1 |
C2 |
H8 |
111.679 |
|
C2 |
C1 |
H3 |
111.679 |
| C2 |
C1 |
H4 |
111.679 |
|
C2 |
C1 |
H5 |
111.679 |
| H3 |
C1 |
H4 |
107.176 |
|
H3 |
C1 |
H5 |
107.176 |
| H4 |
C1 |
H5 |
107.176 |
|
H6 |
C2 |
H7 |
107.176 |
| H6 |
C2 |
H8 |
107.176 |
|
H7 |
C2 |
H8 |
107.176 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
