Jump to
S1C2
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -79.528043 |
| Energy at 298.15K | -79.533983 |
| HF Energy | -79.228226 |
| Nuclear repulsion energy | 42.110031 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3067 |
2921 |
0.00 |
|
|
|
| 2 |
A1g |
1483 |
1412 |
0.00 |
|
|
|
| 3 |
A1g |
1039 |
989 |
0.00 |
|
|
|
| 4 |
A1u |
323 |
307 |
0.00 |
|
|
|
| 5 |
A2u |
3065 |
2919 |
52.26 |
|
|
|
| 6 |
A2u |
1456 |
1386 |
1.28 |
|
|
|
| 7 |
Eg |
3127 |
2978 |
0.00 |
|
|
|
| 7 |
Eg |
3127 |
2978 |
0.00 |
|
|
|
| 8 |
Eg |
1551 |
1477 |
0.00 |
|
|
|
| 8 |
Eg |
1551 |
1477 |
0.00 |
|
|
|
| 9 |
Eg |
1261 |
1201 |
0.00 |
|
|
|
| 9 |
Eg |
1261 |
1201 |
0.00 |
|
|
|
| 10 |
Eu |
3148 |
2998 |
68.41 |
|
|
|
| 10 |
Eu |
3148 |
2998 |
68.41 |
|
|
|
| 11 |
Eu |
1556 |
1482 |
7.68 |
|
|
|
| 11 |
Eu |
1556 |
1482 |
7.68 |
|
|
|
| 12 |
Eu |
843 |
803 |
3.83 |
|
|
|
| 12 |
Eu |
843 |
803 |
3.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16702.0 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 15905.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.765 |
| C2 |
0.000 |
0.000 |
-0.765 |
| H3 |
0.000 |
1.023 |
1.162 |
| H4 |
-0.886 |
-0.511 |
1.162 |
| H5 |
0.886 |
-0.511 |
1.162 |
| H6 |
0.000 |
-1.023 |
-1.162 |
| H7 |
-0.886 |
0.511 |
-1.162 |
| H8 |
0.886 |
0.511 |
-1.162 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5296 | 1.0970 | 1.0970 | 1.0970 | 2.1811 | 2.1811 | 2.1811 |
C2 | 1.5296 | | 2.1811 | 2.1811 | 2.1811 | 1.0970 | 1.0970 | 1.0970 | H3 | 1.0970 | 2.1811 | | 1.7714 | 1.7714 | 3.0955 | 2.5385 | 2.5385 | H4 | 1.0970 | 2.1811 | 1.7714 | | 1.7714 | 2.5385 | 2.5385 | 3.0955 | H5 | 1.0970 | 2.1811 | 1.7714 | 1.7714 | | 2.5385 | 3.0955 | 2.5385 | H6 | 2.1811 | 1.0970 | 3.0955 | 2.5385 | 2.5385 | | 1.7714 | 1.7714 | H7 | 2.1811 | 1.0970 | 2.5385 | 2.5385 | 3.0955 | 1.7714 | | 1.7714 | H8 | 2.1811 | 1.0970 | 2.5385 | 3.0955 | 2.5385 | 1.7714 | 1.7714 | |
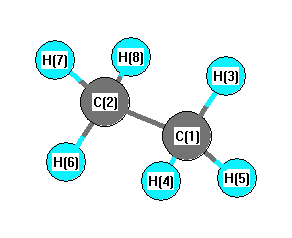 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.209 |
|
C1 |
C2 |
H7 |
111.209 |
| C1 |
C2 |
H8 |
111.209 |
|
C2 |
C1 |
H3 |
111.209 |
| C2 |
C1 |
H4 |
111.209 |
|
C2 |
C1 |
H5 |
111.209 |
| H3 |
C1 |
H4 |
107.679 |
|
H3 |
C1 |
H5 |
107.679 |
| H4 |
C1 |
H5 |
107.679 |
|
H6 |
C2 |
H7 |
107.679 |
| H6 |
C2 |
H8 |
107.679 |
|
H7 |
C2 |
H8 |
107.679 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -79.523264 |
| Energy at 298.15K | |
| HF Energy | -79.223463 |
| Nuclear repulsion energy | 41.946204 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3079 |
2932 |
0.00 |
|
|
|
| 2 |
A1' |
1503 |
1431 |
0.00 |
|
|
|
| 3 |
A1' |
1035 |
986 |
0.00 |
|
|
|
| 4 |
A1" |
314i |
299i |
0.00 |
|
|
|
| 5 |
A2" |
3071 |
2924 |
55.07 |
|
|
|
| 6 |
A2" |
1455 |
1385 |
0.98 |
|
|
|
| 7 |
E' |
3153 |
3003 |
59.59 |
|
|
|
| 7 |
E' |
3153 |
3003 |
59.59 |
|
|
|
| 8 |
E' |
1562 |
1488 |
8.59 |
|
|
|
| 8 |
E' |
1562 |
1488 |
8.59 |
|
|
|
| 9 |
E' |
927 |
883 |
3.49 |
|
|
|
| 9 |
E' |
927 |
883 |
3.49 |
|
|
|
| 10 |
E" |
3134 |
2984 |
0.00 |
|
|
|
| 10 |
E" |
3134 |
2984 |
0.00 |
|
|
|
| 11 |
E" |
1548 |
1474 |
0.00 |
|
|
|
| 11 |
E" |
1548 |
1474 |
0.00 |
|
|
|
| 12 |
E" |
1209 |
1151 |
0.00 |
|
|
|
| 12 |
E" |
1209 |
1151 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16446.8 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 15662.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.772 |
| C2 |
0.000 |
0.000 |
-0.772 |
| H3 |
0.000 |
1.019 |
1.176 |
| H4 |
-0.882 |
-0.509 |
1.176 |
| H5 |
0.882 |
-0.509 |
1.176 |
| H6 |
0.000 |
1.019 |
-1.176 |
| H7 |
0.882 |
-0.509 |
-1.176 |
| H8 |
-0.882 |
-0.509 |
-1.176 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5430 | 1.0961 | 1.0961 | 1.0961 | 2.1982 | 2.1982 | 2.1982 |
C2 | 1.5430 | | 2.1982 | 2.1982 | 2.1982 | 1.0961 | 1.0961 | 1.0961 | H3 | 1.0961 | 2.1982 | | 1.7642 | 1.7642 | 2.3530 | 2.9409 | 2.9409 | H4 | 1.0961 | 2.1982 | 1.7642 | | 1.7642 | 2.9409 | 2.9409 | 2.3530 | H5 | 1.0961 | 2.1982 | 1.7642 | 1.7642 | | 2.9409 | 2.3530 | 2.9409 | H6 | 2.1982 | 1.0961 | 2.3530 | 2.9409 | 2.9409 | | 1.7642 | 1.7642 | H7 | 2.1982 | 1.0961 | 2.9409 | 2.9409 | 2.3530 | 1.7642 | | 1.7642 | H8 | 2.1982 | 1.0961 | 2.9409 | 2.3530 | 2.9409 | 1.7642 | 1.7642 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.682 |
|
C1 |
C2 |
H7 |
111.682 |
| C1 |
C2 |
H8 |
111.682 |
|
C2 |
C1 |
H3 |
111.682 |
| C2 |
C1 |
H4 |
111.682 |
|
C2 |
C1 |
H5 |
111.682 |
| H3 |
C1 |
H4 |
107.173 |
|
H3 |
C1 |
H5 |
107.173 |
| H4 |
C1 |
H5 |
107.173 |
|
H6 |
C2 |
H7 |
107.173 |
| H6 |
C2 |
H8 |
107.173 |
|
H7 |
C2 |
H8 |
107.173 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
