Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -79.726237 |
| Energy at 298.15K | -79.732154 |
| HF Energy | -79.726237 |
| Nuclear repulsion energy | 42.222038 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3070 |
2920 |
0.00 |
213.09 |
0.01 |
0.02 |
| 2 |
A1g |
1455 |
1384 |
0.00 |
7.64 |
0.74 |
0.85 |
| 3 |
A1g |
1036 |
985 |
0.00 |
8.45 |
0.31 |
0.47 |
| 4 |
A1u |
316 |
301 |
0.00 |
0.00 |
0.00 |
0.00 |
| 5 |
A2u |
3071 |
2920 |
55.81 |
0.00 |
0.00 |
0.00 |
| 6 |
A2u |
1427 |
1357 |
1.72 |
0.00 |
0.00 |
0.00 |
| 7 |
Eg |
3133 |
2980 |
0.00 |
135.17 |
0.75 |
0.86 |
| 7 |
Eg |
3133 |
2980 |
0.00 |
135.17 |
0.75 |
0.86 |
| 8 |
Eg |
1531 |
1456 |
0.00 |
39.34 |
0.75 |
0.86 |
| 8 |
Eg |
1531 |
1456 |
0.00 |
39.34 |
0.75 |
0.86 |
| 9 |
Eg |
1234 |
1173 |
0.00 |
2.65 |
0.75 |
0.86 |
| 9 |
Eg |
1234 |
1173 |
0.00 |
2.65 |
0.75 |
0.86 |
| 10 |
Eu |
3157 |
3003 |
62.36 |
0.00 |
0.00 |
0.00 |
| 10 |
Eu |
3157 |
3003 |
62.36 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1535 |
1460 |
9.95 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1535 |
1460 |
9.95 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
832 |
791 |
5.99 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
832 |
791 |
5.99 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 16609.6 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 15795.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.761 |
| C2 |
0.000 |
0.000 |
-0.761 |
| H3 |
0.000 |
1.020 |
1.161 |
| H4 |
-0.883 |
-0.510 |
1.161 |
| H5 |
0.883 |
-0.510 |
1.161 |
| H6 |
0.000 |
-1.020 |
-1.161 |
| H7 |
-0.883 |
0.510 |
-1.161 |
| H8 |
0.883 |
0.510 |
-1.161 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5224 | 1.0954 | 1.0954 | 1.0954 | 2.1758 | 2.1758 | 2.1758 |
C2 | 1.5224 | | 2.1758 | 2.1758 | 2.1758 | 1.0954 | 1.0954 | 1.0954 | H3 | 1.0954 | 2.1758 | | 1.7667 | 1.7667 | 3.0904 | 2.5357 | 2.5357 | H4 | 1.0954 | 2.1758 | 1.7667 | | 1.7667 | 2.5357 | 2.5357 | 3.0904 | H5 | 1.0954 | 2.1758 | 1.7667 | 1.7667 | | 2.5357 | 3.0904 | 2.5357 | H6 | 2.1758 | 1.0954 | 3.0904 | 2.5357 | 2.5357 | | 1.7667 | 1.7667 | H7 | 2.1758 | 1.0954 | 2.5357 | 2.5357 | 3.0904 | 1.7667 | | 1.7667 | H8 | 2.1758 | 1.0954 | 2.5357 | 3.0904 | 2.5357 | 1.7667 | 1.7667 | |
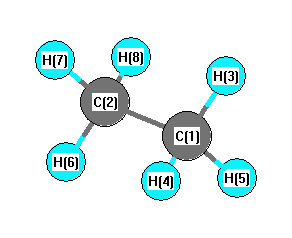 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.389 |
|
C1 |
C2 |
H7 |
111.389 |
| C1 |
C2 |
H8 |
111.389 |
|
C2 |
C1 |
H3 |
111.389 |
| C2 |
C1 |
H4 |
111.389 |
|
C2 |
C1 |
H5 |
111.389 |
| H3 |
C1 |
H4 |
107.487 |
|
H3 |
C1 |
H5 |
107.487 |
| H4 |
C1 |
H5 |
107.487 |
|
H6 |
C2 |
H7 |
107.487 |
| H6 |
C2 |
H8 |
107.487 |
|
H7 |
C2 |
H8 |
107.487 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.509 |
|
|
|
| 2 |
C |
-0.509 |
|
|
|
| 3 |
H |
0.170 |
|
|
|
| 4 |
H |
0.170 |
|
|
|
| 5 |
H |
0.170 |
|
|
|
| 6 |
H |
0.170 |
|
|
|
| 7 |
H |
0.170 |
|
|
|
| 8 |
H |
0.170 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.641 |
0.000 |
0.000 |
| y |
0.000 |
-14.641 |
0.000 |
| z |
0.000 |
0.000 |
-15.158 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.259 |
0.000 |
0.000 |
| y |
0.000 |
0.259 |
0.000 |
| z |
0.000 |
0.000 |
-0.518 |
|
| Polar |
|---|
| 3z2-r2 | -1.035 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.321 |
0.000 |
0.000 |
| y |
0.000 |
3.321 |
0.000 |
| z |
0.000 |
0.000 |
3.549 |
<r2> (average value of r
2) Å
2
| <r2> |
30.532 |
| (<r2>)1/2 |
5.526 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -79.721685 |
| Energy at 298.15K | |
| HF Energy | -79.721685 |
| Nuclear repulsion energy | 42.061298 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3087 |
2935 |
0.00 |
|
|
|
| 2 |
A1' |
1476 |
1404 |
0.00 |
|
|
|
| 3 |
A1' |
1035 |
984 |
0.00 |
|
|
|
| 4 |
A1" |
306i |
291i |
0.00 |
|
|
|
| 5 |
A2" |
3079 |
2928 |
57.93 |
|
|
|
| 6 |
A2" |
1426 |
1356 |
1.62 |
|
|
|
| 7 |
E' |
3165 |
3010 |
51.59 |
|
|
|
| 7 |
E' |
3165 |
3010 |
51.57 |
|
|
|
| 8 |
E' |
1541 |
1466 |
11.34 |
|
|
|
| 8 |
E' |
1541 |
1466 |
11.34 |
|
|
|
| 9 |
E' |
914 |
869 |
5.58 |
|
|
|
| 9 |
E' |
914 |
869 |
5.58 |
|
|
|
| 10 |
E" |
3144 |
2990 |
0.00 |
|
|
|
| 10 |
E" |
3144 |
2990 |
0.00 |
|
|
|
| 11 |
E" |
1531 |
1456 |
0.00 |
|
|
|
| 11 |
E" |
1531 |
1456 |
0.00 |
|
|
|
| 12 |
E" |
1185 |
1127 |
0.00 |
|
|
|
| 12 |
E" |
1185 |
1127 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16378.5 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 15575.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.768 |
| C2 |
0.000 |
0.000 |
-0.768 |
| H3 |
0.000 |
1.016 |
1.176 |
| H4 |
-0.879 |
-0.508 |
1.176 |
| H5 |
0.879 |
-0.508 |
1.176 |
| H6 |
0.000 |
1.016 |
-1.176 |
| H7 |
0.879 |
-0.508 |
-1.176 |
| H8 |
-0.879 |
-0.508 |
-1.176 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5357 | 1.0944 | 1.0944 | 1.0944 | 2.1929 | 2.1929 | 2.1929 |
C2 | 1.5357 | | 2.1929 | 2.1929 | 2.1929 | 1.0944 | 1.0944 | 1.0944 | H3 | 1.0944 | 2.1929 | | 1.7590 | 1.7590 | 2.3514 | 2.9365 | 2.9365 | H4 | 1.0944 | 2.1929 | 1.7590 | | 1.7590 | 2.9365 | 2.9365 | 2.3514 | H5 | 1.0944 | 2.1929 | 1.7590 | 1.7590 | | 2.9365 | 2.3514 | 2.9365 | H6 | 2.1929 | 1.0944 | 2.3514 | 2.9365 | 2.9365 | | 1.7590 | 1.7590 | H7 | 2.1929 | 1.0944 | 2.9365 | 2.9365 | 2.3514 | 1.7590 | | 1.7590 | H8 | 2.1929 | 1.0944 | 2.9365 | 2.3514 | 2.9365 | 1.7590 | 1.7590 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.879 |
|
C1 |
C2 |
H7 |
111.879 |
| C1 |
C2 |
H8 |
111.879 |
|
C2 |
C1 |
H3 |
111.879 |
| C2 |
C1 |
H4 |
111.879 |
|
C2 |
C1 |
H5 |
111.879 |
| H3 |
C1 |
H4 |
106.960 |
|
H3 |
C1 |
H5 |
106.960 |
| H4 |
C1 |
H5 |
106.960 |
|
H6 |
C2 |
H7 |
106.960 |
| H6 |
C2 |
H8 |
106.960 |
|
H7 |
C2 |
H8 |
106.960 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.519 |
|
|
|
| 2 |
C |
-0.519 |
|
|
|
| 3 |
H |
0.173 |
|
|
|
| 4 |
H |
0.173 |
|
|
|
| 5 |
H |
0.173 |
|
|
|
| 6 |
H |
0.173 |
|
|
|
| 7 |
H |
0.173 |
|
|
|
| 8 |
H |
0.173 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.625 |
0.000 |
0.000 |
| y |
0.000 |
-14.625 |
0.000 |
| z |
0.000 |
0.000 |
-15.080 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.228 |
0.000 |
0.000 |
| y |
0.000 |
0.228 |
0.000 |
| z |
0.000 |
0.000 |
-0.455 |
|
| Polar |
|---|
| 3z2-r2 | -0.911 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.255 |
0.000 |
0.000 |
| y |
0.000 |
3.255 |
0.000 |
| z |
0.000 |
0.000 |
3.469 |
<r2> (average value of r
2) Å
2
| <r2> |
30.786 |
| (<r2>)1/2 |
5.549 |
