Jump to
S1C2
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -79.698708 |
| Energy at 298.15K | -79.704579 |
| HF Energy | -79.698708 |
| Nuclear repulsion energy | 41.933794 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
2985 |
2935 |
0.00 |
214.09 |
0.01 |
0.01 |
| 2 |
A1g |
1405 |
1382 |
0.00 |
9.94 |
0.71 |
0.83 |
| 3 |
A1g |
1003 |
987 |
0.00 |
7.11 |
0.31 |
0.47 |
| 4 |
A1u |
310 |
305 |
0.00 |
0.00 |
0.00 |
0.00 |
| 5 |
A2u |
2987 |
2938 |
57.05 |
0.00 |
0.00 |
0.00 |
| 6 |
A2u |
1383 |
1360 |
0.84 |
0.00 |
0.00 |
0.00 |
| 7 |
Eg |
3044 |
2994 |
0.00 |
134.35 |
0.75 |
0.86 |
| 7 |
Eg |
3044 |
2994 |
0.00 |
134.35 |
0.75 |
0.86 |
| 8 |
Eg |
1486 |
1462 |
0.00 |
40.73 |
0.75 |
0.86 |
| 8 |
Eg |
1486 |
1462 |
0.00 |
40.73 |
0.75 |
0.86 |
| 9 |
Eg |
1194 |
1174 |
0.00 |
2.83 |
0.75 |
0.86 |
| 9 |
Eg |
1194 |
1174 |
0.00 |
2.83 |
0.75 |
0.86 |
| 10 |
Eu |
3069 |
3018 |
62.61 |
0.00 |
0.00 |
0.00 |
| 10 |
Eu |
3069 |
3018 |
62.61 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1490 |
1466 |
9.20 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1490 |
1466 |
9.20 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
808 |
795 |
6.42 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
808 |
795 |
6.42 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 16127.7 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 15861.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.766 |
| C2 |
0.000 |
0.000 |
-0.766 |
| H3 |
0.000 |
1.027 |
1.169 |
| H4 |
-0.890 |
-0.514 |
1.169 |
| H5 |
0.890 |
-0.514 |
1.169 |
| H6 |
0.000 |
-1.027 |
-1.169 |
| H7 |
-0.890 |
0.514 |
-1.169 |
| H8 |
0.890 |
0.514 |
-1.169 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5314 | 1.1038 | 1.1038 | 1.1038 | 2.1905 | 2.1905 | 2.1905 |
C2 | 1.5314 | | 2.1905 | 2.1905 | 2.1905 | 1.1038 | 1.1038 | 1.1038 | H3 | 1.1038 | 2.1905 | | 1.7797 | 1.7797 | 3.1126 | 2.5536 | 2.5536 | H4 | 1.1038 | 2.1905 | 1.7797 | | 1.7797 | 2.5536 | 2.5536 | 3.1126 | H5 | 1.1038 | 2.1905 | 1.7797 | 1.7797 | | 2.5536 | 3.1126 | 2.5536 | H6 | 2.1905 | 1.1038 | 3.1126 | 2.5536 | 2.5536 | | 1.7797 | 1.7797 | H7 | 2.1905 | 1.1038 | 2.5536 | 2.5536 | 3.1126 | 1.7797 | | 1.7797 | H8 | 2.1905 | 1.1038 | 2.5536 | 3.1126 | 2.5536 | 1.7797 | 1.7797 | |
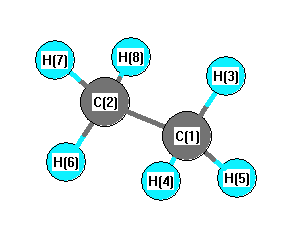 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.425 |
|
C1 |
C2 |
H7 |
111.425 |
| C1 |
C2 |
H8 |
111.425 |
|
C2 |
C1 |
H3 |
111.425 |
| C2 |
C1 |
H4 |
111.425 |
|
C2 |
C1 |
H5 |
111.425 |
| H3 |
C1 |
H4 |
107.449 |
|
H3 |
C1 |
H5 |
107.449 |
| H4 |
C1 |
H5 |
107.449 |
|
H6 |
C2 |
H7 |
107.449 |
| H6 |
C2 |
H8 |
107.449 |
|
H7 |
C2 |
H8 |
107.449 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.476 |
|
|
|
| 2 |
C |
-0.476 |
|
|
|
| 3 |
H |
0.159 |
|
|
|
| 4 |
H |
0.159 |
|
|
|
| 5 |
H |
0.159 |
|
|
|
| 6 |
H |
0.159 |
|
|
|
| 7 |
H |
0.159 |
|
|
|
| 8 |
H |
0.159 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.684 |
0.000 |
0.000 |
| y |
0.000 |
-14.684 |
0.000 |
| z |
0.000 |
0.000 |
-15.166 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.241 |
0.000 |
0.000 |
| y |
0.000 |
0.241 |
0.000 |
| z |
0.000 |
0.000 |
-0.482 |
|
| Polar |
|---|
| 3z2-r2 | -0.963 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.385 |
0.000 |
0.000 |
| y |
0.000 |
3.385 |
0.000 |
| z |
0.000 |
0.000 |
3.653 |
<r2> (average value of r
2) Å
2
| <r2> |
30.839 |
| (<r2>)1/2 |
5.553 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -79.694263 |
| Energy at 298.15K | |
| HF Energy | -79.694263 |
| Nuclear repulsion energy | 41.766556 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3000 |
2950 |
0.00 |
|
|
|
| 2 |
A1' |
1425 |
1402 |
0.00 |
|
|
|
| 3 |
A1' |
1000 |
984 |
0.00 |
|
|
|
| 4 |
A1" |
301i |
296i |
0.00 |
|
|
|
| 5 |
A2" |
2994 |
2945 |
58.59 |
|
|
|
| 6 |
A2" |
1382 |
1359 |
0.83 |
|
|
|
| 7 |
E' |
3075 |
3024 |
51.29 |
|
|
|
| 7 |
E' |
3075 |
3024 |
51.29 |
|
|
|
| 8 |
E' |
1495 |
1471 |
10.58 |
|
|
|
| 8 |
E' |
1495 |
1471 |
10.58 |
|
|
|
| 9 |
E' |
887 |
872 |
6.04 |
|
|
|
| 9 |
E' |
887 |
872 |
6.04 |
|
|
|
| 10 |
E" |
3054 |
3004 |
0.00 |
|
|
|
| 10 |
E" |
3054 |
3004 |
0.00 |
|
|
|
| 11 |
E" |
1488 |
1463 |
0.00 |
|
|
|
| 11 |
E" |
1488 |
1463 |
0.00 |
|
|
|
| 12 |
E" |
1144 |
1125 |
0.00 |
|
|
|
| 12 |
E" |
1144 |
1125 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15893.0 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 15630.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.773 |
| C2 |
0.000 |
0.000 |
-0.773 |
| H3 |
0.000 |
1.023 |
1.184 |
| H4 |
-0.886 |
-0.511 |
1.184 |
| H5 |
0.886 |
-0.511 |
1.184 |
| H6 |
0.000 |
1.023 |
-1.184 |
| H7 |
0.886 |
-0.511 |
-1.184 |
| H8 |
-0.886 |
-0.511 |
-1.184 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5455 | 1.1026 | 1.1026 | 1.1026 | 2.2082 | 2.2082 | 2.2082 |
C2 | 1.5455 | | 2.2082 | 2.2082 | 2.2082 | 1.1026 | 1.1026 | 1.1026 | H3 | 1.1026 | 2.2082 | | 1.7718 | 1.7718 | 2.3685 | 2.9579 | 2.9579 | H4 | 1.1026 | 2.2082 | 1.7718 | | 1.7718 | 2.9579 | 2.9579 | 2.3685 | H5 | 1.1026 | 2.2082 | 1.7718 | 1.7718 | | 2.9579 | 2.3685 | 2.9579 | H6 | 2.2082 | 1.1026 | 2.3685 | 2.9579 | 2.9579 | | 1.7718 | 1.7718 | H7 | 2.2082 | 1.1026 | 2.9579 | 2.9579 | 2.3685 | 1.7718 | | 1.7718 | H8 | 2.2082 | 1.1026 | 2.9579 | 2.3685 | 2.9579 | 1.7718 | 1.7718 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.914 |
|
C1 |
C2 |
H7 |
111.914 |
| C1 |
C2 |
H8 |
111.914 |
|
C2 |
C1 |
H3 |
111.914 |
| C2 |
C1 |
H4 |
111.914 |
|
C2 |
C1 |
H5 |
111.914 |
| H3 |
C1 |
H4 |
106.922 |
|
H3 |
C1 |
H5 |
106.922 |
| H4 |
C1 |
H5 |
106.922 |
|
H6 |
C2 |
H7 |
106.922 |
| H6 |
C2 |
H8 |
106.922 |
|
H7 |
C2 |
H8 |
106.922 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.485 |
|
|
|
| 2 |
C |
-0.485 |
|
|
|
| 3 |
H |
0.162 |
|
|
|
| 4 |
H |
0.162 |
|
|
|
| 5 |
H |
0.162 |
|
|
|
| 6 |
H |
0.162 |
|
|
|
| 7 |
H |
0.162 |
|
|
|
| 8 |
H |
0.162 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.669 |
0.000 |
0.000 |
| y |
0.000 |
-14.669 |
0.000 |
| z |
0.000 |
0.000 |
-15.093 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.212 |
0.000 |
0.000 |
| y |
0.000 |
0.212 |
0.000 |
| z |
0.000 |
0.000 |
-0.424 |
|
| Polar |
|---|
| 3z2-r2 | -0.848 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.314 |
0.000 |
0.000 |
| y |
0.000 |
3.314 |
0.000 |
| z |
0.000 |
0.000 |
3.568 |
<r2> (average value of r
2) Å
2
| <r2> |
31.109 |
| (<r2>)1/2 |
5.578 |
