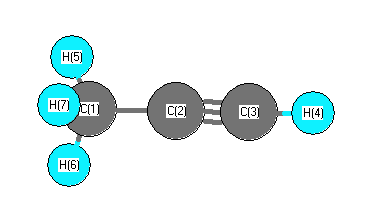
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3515 |
3342 |
53.86 |
31.35 |
0.21 |
0.34 |
| 2 |
A1 |
3067 |
2916 |
16.77 |
177.54 |
0.01 |
0.02 |
| 3 |
A1 |
2267 |
2156 |
3.90 |
93.85 |
0.30 |
0.46 |
| 4 |
A1 |
1439 |
1369 |
0.02 |
23.65 |
0.53 |
0.70 |
| 5 |
A1 |
970 |
922 |
0.62 |
3.31 |
0.21 |
0.35 |
| 6 |
E |
3141 |
2987 |
8.27 |
88.14 |
0.75 |
0.86 |
| 6 |
E |
3141 |
2987 |
8.27 |
88.14 |
0.75 |
0.86 |
| 7 |
E |
1505 |
1431 |
7.76 |
19.91 |
0.75 |
0.86 |
| 7 |
E |
1505 |
1431 |
7.76 |
19.91 |
0.75 |
0.86 |
| 8 |
E |
1069 |
1017 |
2.30 |
0.00 |
0.75 |
0.86 |
| 8 |
E |
1069 |
1017 |
2.30 |
0.00 |
0.75 |
0.86 |
| 9 |
E |
619 |
589 |
52.76 |
0.52 |
0.75 |
0.86 |
| 9 |
E |
619 |
589 |
52.76 |
0.52 |
0.75 |
0.86 |
| 10 |
E |
349 |
332 |
4.10 |
9.77 |
0.75 |
0.86 |
| 10 |
E |
349 |
332 |
4.10 |
9.77 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12311.9 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 11708.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.638 |
|
|
|
| 2 |
C |
0.271 |
|
|
|
| 3 |
C |
-0.481 |
|
|
|
| 4 |
H |
0.222 |
|
|
|
| 5 |
H |
0.209 |
|
|
|
| 6 |
H |
0.209 |
|
|
|
| 7 |
H |
0.209 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.708 |
0.708 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.058 |
0.000 |
0.000 |
| y |
0.000 |
-19.058 |
0.000 |
| z |
0.000 |
0.000 |
-13.424 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.817 |
0.000 |
0.000 |
| y |
0.000 |
-2.817 |
0.000 |
| z |
0.000 |
0.000 |
5.634 |
|
| Polar |
|---|
| 3z2-r2 | 11.269 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.645 |
0.000 |
0.000 |
| y |
0.000 |
2.646 |
0.000 |
| z |
0.000 |
0.000 |
6.624 |
<r2> (average value of r
2) Å
2
| <r2> |
49.686 |
| (<r2>)1/2 |
7.049 |