Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -134.660337 |
| Energy at 298.15K | -134.668623 |
| HF Energy | -134.247225 |
| Nuclear repulsion energy | 82.955122 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3583 |
3309 |
0.44 |
|
|
|
| 2 |
A' |
3211 |
2966 |
41.36 |
|
|
|
| 3 |
A' |
3152 |
2911 |
32.74 |
|
|
|
| 4 |
A' |
3130 |
2891 |
22.12 |
|
|
|
| 5 |
A' |
1765 |
1630 |
27.52 |
|
|
|
| 6 |
A' |
1593 |
1471 |
2.20 |
|
|
|
| 7 |
A' |
1575 |
1455 |
0.12 |
|
|
|
| 8 |
A' |
1500 |
1385 |
9.89 |
|
|
|
| 9 |
A' |
1456 |
1345 |
13.41 |
|
|
|
| 10 |
A' |
1219 |
1126 |
12.84 |
|
|
|
| 11 |
A' |
1136 |
1050 |
33.15 |
|
|
|
| 12 |
A' |
960 |
887 |
130.81 |
|
|
|
| 13 |
A' |
927 |
856 |
55.50 |
|
|
|
| 14 |
A' |
420 |
388 |
7.04 |
|
|
|
| 15 |
A" |
3670 |
3390 |
0.01 |
|
|
|
| 16 |
A" |
3213 |
2968 |
65.68 |
|
|
|
| 17 |
A" |
3187 |
2944 |
2.63 |
|
|
|
| 18 |
A" |
1580 |
1460 |
6.95 |
|
|
|
| 19 |
A" |
1457 |
1346 |
0.00 |
|
|
|
| 20 |
A" |
1340 |
1238 |
0.02 |
|
|
|
| 21 |
A" |
1053 |
973 |
1.44 |
|
|
|
| 22 |
A" |
811 |
750 |
0.68 |
|
|
|
| 23 |
A" |
319 |
294 |
47.93 |
|
|
|
| 24 |
A" |
270 |
249 |
13.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21262.8 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 19640.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.303 |
-0.085 |
0.000 |
| C2 |
0.000 |
0.575 |
0.000 |
| C3 |
1.209 |
-0.356 |
0.000 |
| H4 |
2.145 |
0.206 |
0.000 |
| H5 |
1.203 |
-1.000 |
0.883 |
| H6 |
1.203 |
-1.000 |
-0.883 |
| H7 |
0.037 |
1.229 |
-0.874 |
| H8 |
0.037 |
1.229 |
0.874 |
| H9 |
-1.379 |
-0.690 |
0.810 |
| H10 |
-1.379 |
-0.690 |
-0.810 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| N1 | | 1.4607 | 2.5260 | 3.4599 | 2.8096 | 2.8096 | 2.0703 | 2.0703 | 1.0137 | 1.0137 |
C2 | 1.4607 | | 1.5261 | 2.1764 | 2.1694 | 2.1694 | 1.0918 | 1.0918 | 2.0392 | 2.0392 | C3 | 2.5260 | 1.5261 | | 1.0919 | 1.0922 | 1.0922 | 2.1562 | 2.1562 | 2.7318 | 2.7318 | H4 | 3.4599 | 2.1764 | 1.0919 | | 1.7660 | 1.7660 | 2.5006 | 2.5006 | 3.7248 | 3.7248 | H5 | 2.8096 | 2.1694 | 1.0922 | 1.7660 | | 1.7656 | 3.0677 | 2.5152 | 2.6013 | 3.1027 | H6 | 2.8096 | 2.1694 | 1.0922 | 1.7660 | 1.7656 | | 2.5152 | 3.0677 | 3.1027 | 2.6013 | H7 | 2.0703 | 1.0918 | 2.1562 | 2.5006 | 3.0677 | 2.5152 | | 1.7471 | 2.9193 | 2.3858 | H8 | 2.0703 | 1.0918 | 2.1562 | 2.5006 | 2.5152 | 3.0677 | 1.7471 | | 2.3858 | 2.9193 | H9 | 1.0137 | 2.0392 | 2.7318 | 3.7248 | 2.6013 | 3.1027 | 2.9193 | 2.3858 | | 1.6200 | H10 | 1.0137 | 2.0392 | 2.7318 | 3.7248 | 3.1027 | 2.6013 | 2.3858 | 2.9193 | 1.6200 | |
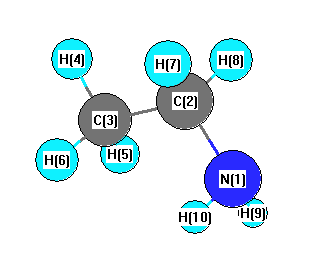 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
115.484 |
|
N1 |
C2 |
H7 |
107.528 |
| N1 |
C2 |
H8 |
107.528 |
|
C2 |
N1 |
H9 |
109.675 |
| C2 |
N1 |
H10 |
109.675 |
|
C2 |
C3 |
H4 |
111.394 |
| C2 |
C3 |
H5 |
110.813 |
|
C2 |
C3 |
H6 |
110.813 |
| C3 |
C2 |
H7 |
109.796 |
|
C3 |
C2 |
H8 |
109.796 |
| H4 |
C3 |
H5 |
107.915 |
|
H4 |
C3 |
H6 |
107.915 |
| H5 |
C3 |
H6 |
107.848 |
|
H7 |
C2 |
H8 |
106.287 |
| H9 |
N1 |
H10 |
106.090 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -134.660167 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
