Jump to
S1C2
Energy calculated at B3LYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -135.170732 |
| Energy at 298.15K | -135.178906 |
| Nuclear repulsion energy | 82.494651 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3449 |
3312 |
2.97 |
|
|
|
| 2 |
A' |
3102 |
2978 |
50.53 |
|
|
|
| 3 |
A' |
3046 |
2925 |
37.47 |
|
|
|
| 4 |
A' |
3032 |
2912 |
24.26 |
|
|
|
| 5 |
A' |
1703 |
1635 |
19.89 |
|
|
|
| 6 |
A' |
1535 |
1474 |
1.49 |
|
|
|
| 7 |
A' |
1515 |
1455 |
0.00 |
|
|
|
| 8 |
A' |
1437 |
1380 |
5.14 |
|
|
|
| 9 |
A' |
1401 |
1345 |
18.51 |
|
|
|
| 10 |
A' |
1168 |
1121 |
11.03 |
|
|
|
| 11 |
A' |
1068 |
1026 |
27.45 |
|
|
|
| 12 |
A' |
918 |
881 |
115.90 |
|
|
|
| 13 |
A' |
875 |
840 |
68.05 |
|
|
|
| 14 |
A' |
402 |
386 |
6.29 |
|
|
|
| 15 |
A" |
3532 |
3392 |
1.54 |
|
|
|
| 16 |
A" |
3106 |
2982 |
70.66 |
|
|
|
| 17 |
A" |
3075 |
2953 |
4.56 |
|
|
|
| 18 |
A" |
1526 |
1465 |
6.27 |
|
|
|
| 19 |
A" |
1409 |
1353 |
0.00 |
|
|
|
| 20 |
A" |
1288 |
1236 |
0.04 |
|
|
|
| 21 |
A" |
1019 |
979 |
1.42 |
|
|
|
| 22 |
A" |
785 |
754 |
1.08 |
|
|
|
| 23 |
A" |
306 |
294 |
45.07 |
|
|
|
| 24 |
A" |
259 |
249 |
10.55 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20476.2 cm
-1
Scaled (by 0.9603) Zero Point Vibrational Energy (zpe) 19663.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.314 |
-0.079 |
0.000 |
| C2 |
0.000 |
0.574 |
0.000 |
| C3 |
1.217 |
-0.361 |
0.000 |
| H4 |
2.158 |
0.204 |
0.000 |
| H5 |
1.215 |
-1.008 |
0.886 |
| H6 |
1.215 |
-1.008 |
-0.886 |
| H7 |
0.041 |
1.233 |
-0.877 |
| H8 |
0.041 |
1.233 |
0.877 |
| H9 |
-1.388 |
-0.691 |
0.812 |
| H10 |
-1.388 |
-0.691 |
-0.812 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| N1 | | 1.4671 | 2.5462 | 3.4832 | 2.8364 | 2.8364 | 2.0795 | 2.0795 | 1.0200 | 1.0200 |
C2 | 1.4671 | | 1.5345 | 2.1895 | 2.1833 | 2.1833 | 1.0975 | 1.0975 | 2.0463 | 2.0463 | C3 | 2.5462 | 1.5345 | | 1.0975 | 1.0977 | 1.0977 | 2.1659 | 2.1659 | 2.7481 | 2.7481 | H4 | 3.4832 | 2.1895 | 1.0975 | | 1.7731 | 1.7731 | 2.5119 | 2.5119 | 3.7460 | 3.7460 | H5 | 2.8364 | 2.1833 | 1.0977 | 1.7731 | | 1.7727 | 3.0840 | 2.5301 | 2.6231 | 3.1242 | H6 | 2.8364 | 2.1833 | 1.0977 | 1.7731 | 1.7727 | | 2.5301 | 3.0840 | 3.1242 | 2.6231 | H7 | 2.0795 | 1.0975 | 2.1659 | 2.5119 | 3.0840 | 2.5301 | | 1.7542 | 2.9320 | 2.3972 | H8 | 2.0795 | 1.0975 | 2.1659 | 2.5119 | 2.5301 | 3.0840 | 1.7542 | | 2.3972 | 2.9320 | H9 | 1.0200 | 2.0463 | 2.7481 | 3.7460 | 2.6231 | 3.1242 | 2.9320 | 2.3972 | | 1.6246 | H10 | 1.0200 | 2.0463 | 2.7481 | 3.7460 | 3.1242 | 2.6231 | 2.3972 | 2.9320 | 1.6246 | |
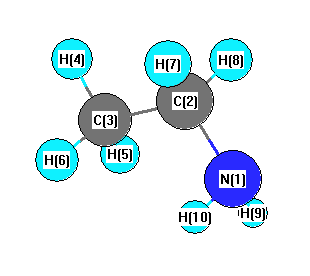 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
116.025 |
|
N1 |
C2 |
H7 |
107.482 |
| N1 |
C2 |
H8 |
107.482 |
|
C2 |
N1 |
H9 |
109.402 |
| C2 |
N1 |
H10 |
109.402 |
|
C2 |
C3 |
H4 |
111.499 |
| C2 |
C3 |
H5 |
110.998 |
|
C2 |
C3 |
H6 |
110.998 |
| C3 |
C2 |
H7 |
109.635 |
|
C3 |
C2 |
H8 |
109.635 |
| H4 |
C3 |
H5 |
107.741 |
|
H4 |
C3 |
H6 |
107.741 |
| H5 |
C3 |
H6 |
107.695 |
|
H7 |
C2 |
H8 |
106.106 |
| H9 |
N1 |
H10 |
105.572 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.699 |
-0.237 |
-0.019 |
-0.973 |
| 2 |
C |
-0.137 |
0.027 |
0.007 |
0.448 |
| 3 |
C |
-0.445 |
0.068 |
0.022 |
-0.373 |
| 4 |
H |
0.141 |
0.066 |
-0.002 |
0.064 |
| 5 |
H |
0.137 |
-0.956 |
-1.018 |
0.085 |
| 6 |
H |
0.137 |
-0.027 |
0.006 |
0.085 |
| 7 |
H |
0.143 |
-0.101 |
-0.022 |
-0.020 |
| 8 |
H |
0.143 |
0.478 |
0.353 |
-0.020 |
| 9 |
H |
0.290 |
0.339 |
0.334 |
0.351 |
| 10 |
H |
0.290 |
0.343 |
0.339 |
0.351 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.975 |
-1.060 |
0.000 |
1.440 |
| CHELPG |
0.280 |
0.596 |
1.205 |
1.373 |
| AIM |
0.321 |
-0.229 |
-0.053 |
0.398 |
| ESP |
0.924 |
-1.076 |
0.000 |
1.419 |
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.914 |
-0.056 |
-0.023 |
| y |
-0.056 |
4.358 |
-0.174 |
| z |
-0.023 |
-0.174 |
3.975 |
<r2> (average value of r
2) Å
2
| <r2> |
58.842 |
| (<r2>)1/2 |
7.671 |
Jump to
S1C1
Energy calculated at B3LYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -135.170043 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/6-31G*
Geometric Data calculated at B3LYP/6-31G*
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
