Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -477.112004 |
| Energy at 298.15K | |
| HF Energy | -476.735460 |
| Nuclear repulsion energy | 107.593445 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3226 |
2980 |
23.89 |
|
|
|
| 2 |
A' |
3175 |
2932 |
22.30 |
|
|
|
| 3 |
A' |
3147 |
2907 |
18.69 |
|
|
|
| 4 |
A' |
2813 |
2599 |
23.02 |
|
|
|
| 5 |
A' |
1588 |
1467 |
1.77 |
|
|
|
| 6 |
A' |
1580 |
1459 |
4.62 |
|
|
|
| 7 |
A' |
1504 |
1389 |
4.76 |
|
|
|
| 8 |
A' |
1399 |
1293 |
38.66 |
|
|
|
| 9 |
A' |
1177 |
1087 |
2.94 |
|
|
|
| 10 |
A' |
1053 |
972 |
2.10 |
|
|
|
| 11 |
A' |
920 |
849 |
3.58 |
|
|
|
| 12 |
A' |
726 |
671 |
0.85 |
|
|
|
| 13 |
A' |
320 |
296 |
2.47 |
|
|
|
| 14 |
A" |
3239 |
2992 |
31.19 |
|
|
|
| 15 |
A" |
3218 |
2973 |
0.03 |
|
|
|
| 16 |
A" |
1577 |
1457 |
7.97 |
|
|
|
| 17 |
A" |
1341 |
1238 |
0.49 |
|
|
|
| 18 |
A" |
1118 |
1033 |
0.45 |
|
|
|
| 19 |
A" |
831 |
768 |
3.51 |
|
|
|
| 20 |
A" |
269 |
249 |
1.52 |
|
|
|
| 21 |
A" |
188 |
174 |
21.80 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17204.2 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 15891.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.516 |
0.688 |
0.000 |
| C2 |
0.000 |
0.824 |
0.000 |
| S3 |
-0.757 |
-0.835 |
0.000 |
| H4 |
1.980 |
1.675 |
0.000 |
| H5 |
1.862 |
0.150 |
0.883 |
| H6 |
1.862 |
0.150 |
-0.883 |
| H7 |
-0.328 |
1.371 |
0.883 |
| H8 |
-0.328 |
1.371 |
-0.883 |
| H9 |
-2.034 |
-0.435 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5225 | 2.7359 | 1.0906 | 1.0903 | 1.0903 | 2.1561 | 2.1561 | 3.7240 |
C2 | 1.5225 | | 1.8237 | 2.1550 | 2.1681 | 2.1681 | 1.0890 | 1.0890 | 2.3928 | S3 | 2.7359 | 1.8237 | | 3.7134 | 2.9336 | 2.9336 | 2.4143 | 2.4143 | 1.3384 | H4 | 1.0906 | 2.1550 | 3.7134 | | 1.7663 | 1.7663 | 2.4900 | 2.4900 | 4.5352 | H5 | 1.0903 | 2.1681 | 2.9336 | 1.7663 | | 1.7670 | 2.5072 | 3.0669 | 4.0374 | H6 | 1.0903 | 2.1681 | 2.9336 | 1.7663 | 1.7670 | | 3.0669 | 2.5072 | 4.0374 | H7 | 2.1561 | 1.0890 | 2.4143 | 2.4900 | 2.5072 | 3.0669 | | 1.7658 | 2.6369 | H8 | 2.1561 | 1.0890 | 2.4143 | 2.4900 | 3.0669 | 2.5072 | 1.7658 | | 2.6369 | H9 | 3.7240 | 2.3928 | 1.3384 | 4.5352 | 4.0374 | 4.0374 | 2.6369 | 2.6369 | |
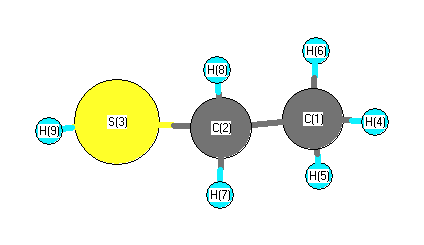 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
109.367 |
|
C1 |
C2 |
H7 |
110.196 |
| C1 |
C2 |
H8 |
110.196 |
|
C2 |
C1 |
H4 |
110.012 |
| C2 |
C1 |
H5 |
111.075 |
|
C2 |
C1 |
H6 |
111.075 |
| C2 |
S3 |
H9 |
97.156 |
|
S3 |
C2 |
H7 |
109.364 |
| S3 |
C2 |
H8 |
109.364 |
|
H4 |
C1 |
H5 |
108.164 |
| H4 |
C1 |
H6 |
108.164 |
|
H5 |
C1 |
H6 |
108.250 |
| H7 |
C2 |
H8 |
108.334 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -477.112724 |
| Energy at 298.15K | -477.119161 |
| HF Energy | -476.735958 |
| Nuclear repulsion energy | 107.357732 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3240 |
2992 |
28.87 |
|
|
|
| 2 |
A |
3221 |
2975 |
9.91 |
|
|
|
| 3 |
A |
3211 |
2966 |
20.12 |
|
|
|
| 4 |
A |
3174 |
2932 |
15.31 |
|
|
|
| 5 |
A |
3141 |
2901 |
22.27 |
|
|
|
| 6 |
A |
2807 |
2593 |
20.58 |
|
|
|
| 7 |
A |
1582 |
1462 |
3.10 |
|
|
|
| 8 |
A |
1576 |
1455 |
8.99 |
|
|
|
| 9 |
A |
1566 |
1447 |
1.74 |
|
|
|
| 10 |
A |
1502 |
1387 |
3.86 |
|
|
|
| 11 |
A |
1404 |
1297 |
19.52 |
|
|
|
| 12 |
A |
1352 |
1249 |
2.91 |
|
|
|
| 13 |
A |
1194 |
1103 |
13.26 |
|
|
|
| 14 |
A |
1130 |
1044 |
0.24 |
|
|
|
| 15 |
A |
1046 |
966 |
4.96 |
|
|
|
| 16 |
A |
925 |
854 |
9.57 |
|
|
|
| 17 |
A |
778 |
719 |
1.70 |
|
|
|
| 18 |
A |
714 |
660 |
3.12 |
|
|
|
| 19 |
A |
346 |
319 |
1.99 |
|
|
|
| 20 |
A |
276 |
255 |
2.53 |
|
|
|
| 21 |
A |
226 |
209 |
19.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17205.1 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 15892.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.635 |
-0.350 |
-0.053 |
| C2 |
0.492 |
0.644 |
0.090 |
| S3 |
-1.161 |
-0.099 |
-0.079 |
| H4 |
2.595 |
0.165 |
0.018 |
| H5 |
1.601 |
-1.105 |
0.734 |
| H6 |
1.586 |
-0.863 |
-1.014 |
| H7 |
0.553 |
1.173 |
1.040 |
| H8 |
0.541 |
1.396 |
-0.697 |
| H9 |
-1.065 |
-0.935 |
0.963 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5216 | 2.8075 | 1.0920 | 1.0911 | 1.0895 | 2.1656 | 2.1589 | 2.9434 |
C2 | 1.5216 | | 1.8206 | 2.1583 | 2.1683 | 2.1645 | 1.0893 | 1.0900 | 2.3830 | S3 | 2.8075 | 1.8206 | | 3.7669 | 3.0494 | 3.0005 | 2.4103 | 2.3481 | 1.3390 | H4 | 1.0920 | 2.1583 | 3.7669 | | 1.7649 | 1.7711 | 2.4974 | 2.4996 | 3.9370 | H5 | 1.0911 | 2.1683 | 3.0494 | 1.7649 | | 1.7644 | 2.5266 | 3.0701 | 2.6806 | H6 | 1.0895 | 2.1645 | 3.0005 | 1.7711 | 1.7644 | | 3.0712 | 2.5084 | 3.3073 | H7 | 2.1656 | 1.0893 | 2.4103 | 2.4974 | 2.5266 | 3.0712 | | 1.7521 | 2.6586 | H8 | 2.1589 | 1.0900 | 2.3481 | 2.4996 | 3.0701 | 2.5084 | 1.7521 | | 3.2812 | H9 | 2.9434 | 2.3830 | 1.3390 | 3.9370 | 2.6806 | 3.3073 | 2.6586 | 3.2812 | |
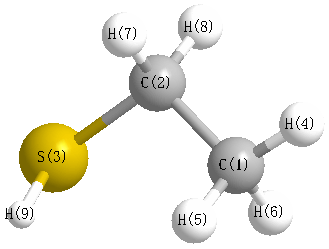 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
113.986 |
|
C1 |
C2 |
H7 |
110.996 |
| C1 |
C2 |
H8 |
110.424 |
|
C2 |
C1 |
H4 |
110.258 |
| C2 |
C1 |
H5 |
111.109 |
|
C2 |
C1 |
H6 |
110.892 |
| C2 |
S3 |
H9 |
96.731 |
|
S3 |
C2 |
H7 |
109.260 |
| S3 |
C2 |
H8 |
104.760 |
|
H4 |
C1 |
H5 |
107.895 |
| H4 |
C1 |
H6 |
108.561 |
|
H5 |
C1 |
H6 |
108.023 |
| H7 |
C2 |
H8 |
107.022 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
