Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -477.934755 |
| Energy at 298.15K | -477.940971 |
| HF Energy | -477.934755 |
| Nuclear repulsion energy | 107.132400 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3142 |
3006 |
22.15 |
|
|
|
| 2 |
A' |
3084 |
2950 |
24.05 |
|
|
|
| 3 |
A' |
3065 |
2932 |
19.77 |
|
|
|
| 4 |
A' |
2704 |
2587 |
24.36 |
|
|
|
| 5 |
A' |
1530 |
1463 |
2.90 |
|
|
|
| 6 |
A' |
1514 |
1449 |
3.45 |
|
|
|
| 7 |
A' |
1438 |
1376 |
5.24 |
|
|
|
| 8 |
A' |
1324 |
1267 |
42.96 |
|
|
|
| 9 |
A' |
1129 |
1080 |
2.33 |
|
|
|
| 10 |
A' |
1012 |
969 |
3.98 |
|
|
|
| 11 |
A' |
871 |
834 |
2.69 |
|
|
|
| 12 |
A' |
684 |
655 |
1.36 |
|
|
|
| 13 |
A' |
302 |
289 |
2.83 |
|
|
|
| 14 |
A" |
3155 |
3018 |
29.99 |
|
|
|
| 15 |
A" |
3130 |
2995 |
0.47 |
|
|
|
| 16 |
A" |
1520 |
1454 |
9.00 |
|
|
|
| 17 |
A" |
1285 |
1229 |
0.74 |
|
|
|
| 18 |
A" |
1063 |
1017 |
0.34 |
|
|
|
| 19 |
A" |
800 |
765 |
5.18 |
|
|
|
| 20 |
A" |
260 |
249 |
0.83 |
|
|
|
| 21 |
A" |
163 |
156 |
22.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16587.6 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 15869.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.517 |
0.696 |
0.000 |
| C2 |
0.000 |
0.831 |
0.000 |
| S3 |
-0.756 |
-0.842 |
0.000 |
| H4 |
1.981 |
1.688 |
0.000 |
| H5 |
1.868 |
0.158 |
0.887 |
| H6 |
1.868 |
0.158 |
-0.887 |
| H7 |
-0.335 |
1.376 |
0.888 |
| H8 |
-0.335 |
1.376 |
-0.888 |
| H9 |
-2.043 |
-0.440 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5226 | 2.7443 | 1.0957 | 1.0951 | 1.0951 | 2.1632 | 2.1632 | 3.7368 |
C2 | 1.5226 | | 1.8357 | 2.1587 | 2.1743 | 2.1743 | 1.0944 | 1.0944 | 2.4064 | S3 | 2.7443 | 1.8357 | | 3.7276 | 2.9446 | 2.9446 | 2.4259 | 2.4259 | 1.3481 | H4 | 1.0957 | 2.1587 | 3.7276 | | 1.7725 | 1.7725 | 2.5000 | 2.5000 | 4.5526 | H5 | 1.0951 | 2.1743 | 2.9446 | 1.7725 | | 1.7738 | 2.5170 | 3.0798 | 4.0545 | H6 | 1.0951 | 2.1743 | 2.9446 | 1.7725 | 1.7738 | | 3.0798 | 2.5170 | 4.0545 | H7 | 2.1632 | 1.0944 | 2.4259 | 2.5000 | 2.5170 | 3.0798 | | 1.7756 | 2.6468 | H8 | 2.1632 | 1.0944 | 2.4259 | 2.5000 | 3.0798 | 2.5170 | 1.7756 | | 2.6468 | H9 | 3.7368 | 2.4064 | 1.3481 | 4.5526 | 4.0545 | 4.0545 | 2.6468 | 2.6468 | |
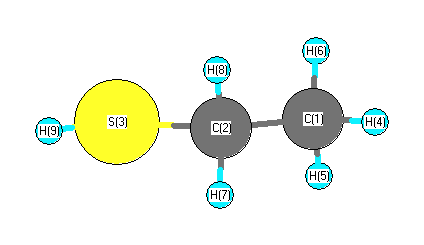 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
109.245 |
|
C1 |
C2 |
H7 |
110.430 |
| C1 |
C2 |
H8 |
110.430 |
|
C2 |
C1 |
H4 |
109.997 |
| C2 |
C1 |
H5 |
111.269 |
|
C2 |
C1 |
H6 |
111.269 |
| C2 |
S3 |
H9 |
96.999 |
|
S3 |
C2 |
H7 |
109.137 |
| S3 |
C2 |
H8 |
109.137 |
|
H4 |
C1 |
H5 |
108.008 |
| H4 |
C1 |
H6 |
108.008 |
|
H5 |
C1 |
H6 |
108.168 |
| H7 |
C2 |
H8 |
108.434 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.521 |
|
|
|
| 2 |
C |
-0.456 |
|
|
|
| 3 |
S |
-0.089 |
|
|
|
| 4 |
H |
0.181 |
|
|
|
| 5 |
H |
0.190 |
|
|
|
| 6 |
H |
0.190 |
|
|
|
| 7 |
H |
0.201 |
|
|
|
| 8 |
H |
0.201 |
|
|
|
| 9 |
H |
0.103 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.052 |
1.830 |
0.000 |
1.831 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.020 |
1.846 |
0.000 |
1.846 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.283 |
-0.208 |
0.000 |
| y |
-0.208 |
-28.080 |
0.000 |
| z |
0.000 |
0.000 |
-28.705 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.110 |
-0.208 |
0.000 |
| y |
-0.208 |
-1.587 |
0.000 |
| z |
0.000 |
0.000 |
-2.523 |
|
| Polar |
|---|
| 3z2-r2 | -5.046 |
|---|
| x2-y2 | 3.798 |
|---|
| xy | -0.208 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.246 |
0.803 |
0.000 |
| y |
0.803 |
5.881 |
0.000 |
| z |
0.000 |
0.000 |
4.482 |
<r2> (average value of r
2) Å
2
| <r2> |
83.548 |
| (<r2>)1/2 |
9.140 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -477.935663 |
| Energy at 298.15K | -477.942004 |
| HF Energy | -477.935663 |
| Nuclear repulsion energy | 106.856737 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3157 |
3020 |
26.93 |
|
|
|
| 2 |
A |
3135 |
2999 |
12.82 |
|
|
|
| 3 |
A |
3125 |
2990 |
17.15 |
|
|
|
| 4 |
A |
3086 |
2952 |
15.83 |
|
|
|
| 5 |
A |
3056 |
2924 |
25.43 |
|
|
|
| 6 |
A |
2704 |
2587 |
21.76 |
|
|
|
| 7 |
A |
1524 |
1458 |
3.27 |
|
|
|
| 8 |
A |
1517 |
1451 |
10.16 |
|
|
|
| 9 |
A |
1503 |
1438 |
1.98 |
|
|
|
| 10 |
A |
1435 |
1373 |
3.99 |
|
|
|
| 11 |
A |
1329 |
1271 |
21.04 |
|
|
|
| 12 |
A |
1295 |
1239 |
4.57 |
|
|
|
| 13 |
A |
1139 |
1090 |
12.21 |
|
|
|
| 14 |
A |
1082 |
1035 |
0.26 |
|
|
|
| 15 |
A |
1002 |
959 |
7.70 |
|
|
|
| 16 |
A |
884 |
846 |
9.73 |
|
|
|
| 17 |
A |
747 |
714 |
2.41 |
|
|
|
| 18 |
A |
669 |
640 |
4.10 |
|
|
|
| 19 |
A |
329 |
315 |
2.68 |
|
|
|
| 20 |
A |
269 |
257 |
3.82 |
|
|
|
| 21 |
A |
232 |
222 |
18.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16609.7 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 15890.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.642 |
-0.350 |
-0.054 |
| C2 |
0.499 |
0.643 |
0.092 |
| S3 |
-1.168 |
-0.099 |
-0.079 |
| H4 |
2.607 |
0.168 |
0.014 |
| H5 |
1.614 |
-1.107 |
0.738 |
| H6 |
1.594 |
-0.868 |
-1.017 |
| H7 |
0.555 |
1.179 |
1.045 |
| H8 |
0.538 |
1.397 |
-0.702 |
| H9 |
-1.070 |
-0.944 |
0.967 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5217 | 2.8216 | 1.0973 | 1.0959 | 1.0943 | 2.1744 | 2.1661 | 2.9582 |
C2 | 1.5217 | | 1.8323 | 2.1629 | 2.1737 | 2.1705 | 1.0947 | 1.0951 | 2.3967 | S3 | 2.8216 | 1.8323 | | 3.7858 | 3.0698 | 3.0164 | 2.4219 | 2.3523 | 1.3488 | H4 | 1.0973 | 2.1629 | 3.7858 | | 1.7708 | 1.7782 | 2.5092 | 2.5113 | 3.9578 | H5 | 1.0959 | 2.1737 | 3.0698 | 1.7708 | | 1.7712 | 2.5384 | 3.0824 | 2.6985 | H6 | 1.0943 | 2.1705 | 3.0164 | 1.7782 | 1.7712 | | 3.0853 | 2.5186 | 3.3227 | H7 | 2.1744 | 1.0947 | 2.4219 | 2.5092 | 2.5384 | 3.0853 | | 1.7598 | 2.6744 | H8 | 2.1661 | 1.0951 | 2.3523 | 2.5113 | 3.0824 | 2.5186 | 1.7598 | | 3.2936 | H9 | 2.9582 | 2.3967 | 1.3488 | 3.9578 | 2.6985 | 3.3227 | 2.6744 | 3.2936 | |
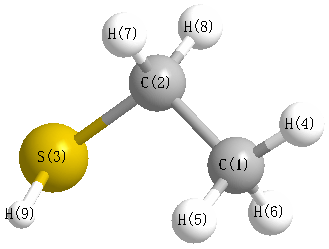 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
114.228 |
|
C1 |
C2 |
H7 |
111.363 |
| C1 |
C2 |
H8 |
110.682 |
|
C2 |
C1 |
H4 |
110.296 |
| C2 |
C1 |
H5 |
111.241 |
|
C2 |
C1 |
H6 |
111.082 |
| C2 |
S3 |
H9 |
96.596 |
|
S3 |
C2 |
H7 |
109.055 |
| S3 |
C2 |
H8 |
104.087 |
|
H4 |
C1 |
H5 |
107.697 |
| H4 |
C1 |
H6 |
108.463 |
|
H5 |
C1 |
H6 |
107.941 |
| H7 |
C2 |
H8 |
106.953 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.499 |
|
|
|
| 2 |
C |
-0.470 |
|
|
|
| 3 |
S |
-0.086 |
|
|
|
| 4 |
H |
0.175 |
|
|
|
| 5 |
H |
0.173 |
|
|
|
| 6 |
H |
0.191 |
|
|
|
| 7 |
H |
0.204 |
|
|
|
| 8 |
H |
0.207 |
|
|
|
| 9 |
H |
0.104 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.671 |
0.058 |
0.826 |
1.865 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.946 |
1.398 |
-0.870 |
| y |
1.398 |
-26.704 |
-1.722 |
| z |
-0.870 |
-1.722 |
-26.730 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.229 |
1.398 |
-0.870 |
| y |
1.398 |
1.135 |
-1.722 |
| z |
-0.870 |
-1.722 |
1.095 |
|
| Polar |
|---|
| 3z2-r2 | 2.189 |
|---|
| x2-y2 | -2.243 |
|---|
| xy | 1.398 |
|---|
| xz | -0.870 |
|---|
| yz | -1.722 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.715 |
0.444 |
0.029 |
| y |
0.444 |
5.105 |
-0.331 |
| z |
0.029 |
-0.331 |
4.904 |
<r2> (average value of r
2) Å
2
| <r2> |
84.217 |
| (<r2>)1/2 |
9.177 |
