Jump to
S1C2
Energy calculated at MP4=FULL/6-31G*
| | hartrees |
|---|
| Energy at 0K | -477.193416 |
| Energy at 298.15K | |
| HF Energy | -476.734809 |
| Nuclear repulsion energy | 107.168452 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3148 |
3027 |
|
|
|
|
| 2 |
A' |
3085 |
2967 |
|
|
|
|
| 3 |
A' |
3066 |
2948 |
|
|
|
|
| 4 |
A' |
2717 |
2613 |
|
|
|
|
| 5 |
A' |
1550 |
1491 |
|
|
|
|
| 6 |
A' |
1539 |
1480 |
|
|
|
|
| 7 |
A' |
1462 |
1406 |
|
|
|
|
| 8 |
A' |
1357 |
1305 |
|
|
|
|
| 9 |
A' |
1144 |
1100 |
|
|
|
|
| 10 |
A' |
1026 |
987 |
|
|
|
|
| 11 |
A' |
888 |
854 |
|
|
|
|
| 12 |
A' |
701 |
674 |
|
|
|
|
| 13 |
A' |
312 |
300 |
|
|
|
|
| 14 |
A" |
3157 |
3036 |
|
|
|
|
| 15 |
A" |
3134 |
3013 |
|
|
|
|
| 16 |
A" |
1540 |
1481 |
|
|
|
|
| 17 |
A" |
1306 |
1256 |
|
|
|
|
| 18 |
A" |
1088 |
1046 |
|
|
|
|
| 19 |
A" |
811 |
780 |
|
|
|
|
| 20 |
A" |
267 |
257 |
|
|
|
|
| 21 |
A" |
177 |
170 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16737.4 cm
-1
Scaled (by 0.9616) Zero Point Vibrational Energy (zpe) 16094.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.520 |
0.687 |
0.000 |
| C2 |
0.000 |
0.830 |
0.000 |
| S3 |
-0.758 |
-0.838 |
0.000 |
| H4 |
1.991 |
1.678 |
0.000 |
| H5 |
1.863 |
0.144 |
0.889 |
| H6 |
1.863 |
0.144 |
-0.889 |
| H7 |
-0.330 |
1.380 |
0.889 |
| H8 |
-0.330 |
1.380 |
-0.889 |
| H9 |
-2.042 |
-0.425 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5263 | 2.7413 | 1.0970 | 1.0967 | 1.0967 | 2.1659 | 2.1659 | 3.7310 |
C2 | 1.5263 | | 1.8318 | 2.1644 | 2.1753 | 2.1753 | 1.0962 | 1.0962 | 2.3966 | S3 | 2.7413 | 1.8318 | | 3.7267 | 2.9373 | 2.9373 | 2.4270 | 2.4270 | 1.3478 | H4 | 1.0970 | 2.1644 | 3.7267 | | 1.7770 | 1.7770 | 2.5036 | 2.5036 | 4.5484 | H5 | 1.0967 | 2.1753 | 2.9373 | 1.7770 | | 1.7774 | 2.5175 | 3.0818 | 4.0451 | H6 | 1.0967 | 2.1753 | 2.9373 | 1.7770 | 1.7774 | | 3.0818 | 2.5175 | 4.0451 | H7 | 2.1659 | 1.0962 | 2.4270 | 2.5036 | 2.5175 | 3.0818 | | 1.7779 | 2.6416 | H8 | 2.1659 | 1.0962 | 2.4270 | 2.5036 | 3.0818 | 2.5175 | 1.7779 | | 2.6416 | H9 | 3.7310 | 2.3966 | 1.3478 | 4.5484 | 4.0451 | 4.0451 | 2.6416 | 2.6416 | |
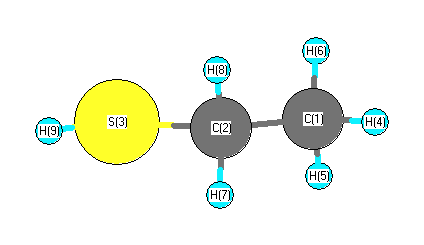 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
109.097 |
|
C1 |
C2 |
H7 |
110.284 |
| C1 |
C2 |
H8 |
110.284 |
|
C2 |
C1 |
H4 |
110.108 |
| C2 |
C1 |
H5 |
110.995 |
|
C2 |
C1 |
H6 |
110.995 |
| C2 |
S3 |
H9 |
96.650 |
|
S3 |
C2 |
H7 |
109.390 |
| S3 |
C2 |
H8 |
109.390 |
|
H4 |
C1 |
H5 |
108.196 |
| H4 |
C1 |
H6 |
108.196 |
|
H5 |
C1 |
H6 |
108.252 |
| H7 |
C2 |
H8 |
108.378 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4=FULL/6-31G*
| | hartrees |
|---|
| Energy at 0K | -477.194305 |
| Energy at 298.15K | -477.200686 |
| HF Energy | -476.735293 |
| Nuclear repulsion energy | 106.928815 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3159 |
3038 |
|
|
|
|
| 2 |
A |
3139 |
3019 |
|
|
|
|
| 3 |
A |
3130 |
3010 |
|
|
|
|
| 4 |
A |
3085 |
2967 |
|
|
|
|
| 5 |
A |
3059 |
2942 |
|
|
|
|
| 6 |
A |
2708 |
2604 |
|
|
|
|
| 7 |
A |
1545 |
1486 |
|
|
|
|
| 8 |
A |
1538 |
1479 |
|
|
|
|
| 9 |
A |
1526 |
1467 |
|
|
|
|
| 10 |
A |
1459 |
1403 |
|
|
|
|
| 11 |
A |
1362 |
1310 |
|
|
|
|
| 12 |
A |
1317 |
1267 |
|
|
|
|
| 13 |
A |
1161 |
1116 |
|
|
|
|
| 14 |
A |
1100 |
1058 |
|
|
|
|
| 15 |
A |
1019 |
980 |
|
|
|
|
| 16 |
A |
898 |
864 |
|
|
|
|
| 17 |
A |
759 |
730 |
|
|
|
|
| 18 |
A |
688 |
662 |
|
|
|
|
| 19 |
A |
338 |
325 |
|
|
|
|
| 20 |
A |
273 |
263 |
|
|
|
|
| 21 |
A |
227 |
218 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16744.9 cm
-1
Scaled (by 0.9616) Zero Point Vibrational Energy (zpe) 16101.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.638 |
-0.352 |
-0.054 |
| C2 |
0.495 |
0.647 |
0.091 |
| S3 |
-1.165 |
-0.100 |
-0.079 |
| H4 |
2.606 |
0.163 |
0.008 |
| H5 |
1.605 |
-1.107 |
0.741 |
| H6 |
1.581 |
-0.872 |
-1.018 |
| H7 |
0.558 |
1.180 |
1.048 |
| H8 |
0.543 |
1.403 |
-0.703 |
| H9 |
-1.050 |
-0.945 |
0.966 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5254 | 2.8148 | 1.0984 | 1.0976 | 1.0959 | 2.1747 | 2.1683 | 2.9360 |
C2 | 1.5254 | | 1.8285 | 2.1673 | 2.1756 | 2.1716 | 1.0965 | 1.0971 | 2.3849 | S3 | 2.8148 | 1.8285 | | 3.7814 | 3.0595 | 3.0030 | 2.4236 | 2.3589 | 1.3487 | H4 | 1.0984 | 2.1673 | 3.7814 | | 1.7761 | 1.7818 | 2.5120 | 2.5100 | 3.9387 | H5 | 1.0976 | 2.1756 | 3.0595 | 1.7761 | | 1.7747 | 2.5342 | 3.0848 | 2.6695 | H6 | 1.0959 | 2.1716 | 3.0030 | 1.7818 | 1.7747 | | 3.0857 | 2.5205 | 3.2959 | H7 | 2.1747 | 1.0965 | 2.4236 | 2.5120 | 2.5342 | 3.0857 | | 1.7644 | 2.6655 | H8 | 2.1683 | 1.0971 | 2.3589 | 2.5100 | 3.0848 | 2.5205 | 1.7644 | | 3.2915 | H9 | 2.9360 | 2.3849 | 1.3487 | 3.9387 | 2.6695 | 3.2959 | 2.6655 | 3.2915 | |
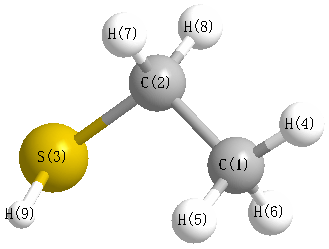 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
113.823 |
|
C1 |
C2 |
H7 |
111.028 |
| C1 |
C2 |
H8 |
110.482 |
|
C2 |
C1 |
H4 |
110.332 |
| C2 |
C1 |
H5 |
111.032 |
|
C2 |
C1 |
H6 |
110.815 |
| C2 |
S3 |
H9 |
96.115 |
|
S3 |
C2 |
H7 |
109.341 |
| S3 |
C2 |
H8 |
104.692 |
|
H4 |
C1 |
H5 |
107.960 |
| H4 |
C1 |
H6 |
108.590 |
|
H5 |
C1 |
H6 |
108.013 |
| H7 |
C2 |
H8 |
107.091 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
