Jump to
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -477.733828 |
| Energy at 298.15K | -477.740092 |
| HF Energy | -477.606764 |
| Nuclear repulsion energy | 107.126461 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3165 |
3004 |
23.44 |
|
|
|
| 2 |
A' |
3110 |
2952 |
23.70 |
|
|
|
| 3 |
A' |
3090 |
2933 |
18.83 |
|
|
|
| 4 |
A' |
2729 |
2591 |
23.71 |
|
|
|
| 5 |
A' |
1555 |
1476 |
1.84 |
|
|
|
| 6 |
A' |
1542 |
1463 |
3.98 |
|
|
|
| 7 |
A' |
1463 |
1389 |
3.47 |
|
|
|
| 8 |
A' |
1350 |
1282 |
44.91 |
|
|
|
| 9 |
A' |
1142 |
1084 |
2.41 |
|
|
|
| 10 |
A' |
1019 |
967 |
2.92 |
|
|
|
| 11 |
A' |
882 |
838 |
2.77 |
|
|
|
| 12 |
A' |
689 |
654 |
1.43 |
|
|
|
| 13 |
A' |
308 |
293 |
2.65 |
|
|
|
| 14 |
A" |
3177 |
3016 |
32.06 |
|
|
|
| 15 |
A" |
3155 |
2995 |
0.27 |
|
|
|
| 16 |
A" |
1545 |
1467 |
7.49 |
|
|
|
| 17 |
A" |
1305 |
1239 |
0.54 |
|
|
|
| 18 |
A" |
1082 |
1027 |
0.45 |
|
|
|
| 19 |
A" |
812 |
770 |
3.90 |
|
|
|
| 20 |
A" |
264 |
251 |
1.14 |
|
|
|
| 21 |
A" |
172 |
163 |
21.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16777.7 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 15925.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.519 |
0.691 |
0.000 |
| C2 |
0.000 |
0.831 |
0.000 |
| S3 |
-0.758 |
-0.840 |
0.000 |
| H4 |
1.983 |
1.682 |
0.000 |
| H5 |
1.865 |
0.152 |
0.886 |
| H6 |
1.865 |
0.152 |
-0.886 |
| H7 |
-0.329 |
1.378 |
0.886 |
| H8 |
-0.329 |
1.378 |
-0.886 |
| H9 |
-2.041 |
-0.434 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5258 | 2.7444 | 1.0935 | 1.0930 | 1.0930 | 2.1616 | 2.1616 | 3.7338 |
C2 | 1.5258 | | 1.8351 | 2.1575 | 2.1735 | 2.1735 | 1.0922 | 1.0922 | 2.4011 | S3 | 2.7444 | 1.8351 | | 3.7244 | 2.9413 | 2.9413 | 2.4270 | 2.4270 | 1.3457 | H4 | 1.0935 | 2.1575 | 3.7244 | | 1.7712 | 1.7712 | 2.4940 | 2.4940 | 4.5459 | H5 | 1.0930 | 2.1735 | 2.9413 | 1.7712 | | 1.7720 | 2.5129 | 3.0750 | 4.0479 | H6 | 1.0930 | 2.1735 | 2.9413 | 1.7712 | 1.7720 | | 3.0750 | 2.5129 | 4.0479 | H7 | 2.1616 | 1.0922 | 2.4270 | 2.4940 | 2.5129 | 3.0750 | | 1.7727 | 2.6458 | H8 | 2.1616 | 1.0922 | 2.4270 | 2.4940 | 3.0750 | 2.5129 | 1.7727 | | 2.6458 | H9 | 3.7338 | 2.4011 | 1.3457 | 4.5459 | 4.0479 | 4.0479 | 2.6458 | 2.6458 | |
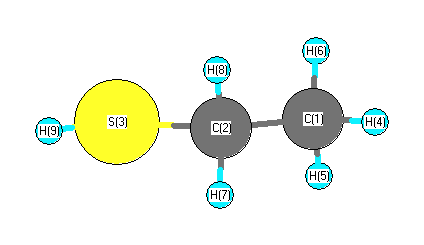 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
109.141 |
|
C1 |
C2 |
H7 |
110.216 |
| C1 |
C2 |
H8 |
110.216 |
|
C2 |
C1 |
H4 |
109.813 |
| C2 |
C1 |
H5 |
111.109 |
|
C2 |
C1 |
H6 |
111.109 |
| C2 |
S3 |
H9 |
96.825 |
|
S3 |
C2 |
H7 |
109.378 |
| S3 |
C2 |
H8 |
109.378 |
|
H4 |
C1 |
H5 |
108.205 |
| H4 |
C1 |
H6 |
108.205 |
|
H5 |
C1 |
H6 |
108.303 |
| H7 |
C2 |
H8 |
108.494 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.470 |
|
|
|
| 2 |
C |
-0.405 |
|
|
|
| 3 |
S |
-0.083 |
|
|
|
| 4 |
H |
0.163 |
|
|
|
| 5 |
H |
0.172 |
|
|
|
| 6 |
H |
0.172 |
|
|
|
| 7 |
H |
0.180 |
|
|
|
| 8 |
H |
0.180 |
|
|
|
| 9 |
H |
0.092 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.139 |
0.793 |
0.000 |
| y |
0.793 |
5.814 |
0.000 |
| z |
0.000 |
0.000 |
4.430 |
<r2> (average value of r
2) Å
2
| <r2> |
83.795 |
| (<r2>)1/2 |
9.154 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -477.734675 |
| Energy at 298.15K | -477.741047 |
| HF Energy | -477.607455 |
| Nuclear repulsion energy | 106.863185 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3179 |
3017 |
28.88 |
|
|
|
| 2 |
A |
3158 |
2998 |
11.68 |
|
|
|
| 3 |
A |
3148 |
2988 |
19.51 |
|
|
|
| 4 |
A |
3111 |
2953 |
15.86 |
|
|
|
| 5 |
A |
3081 |
2925 |
23.67 |
|
|
|
| 6 |
A |
2727 |
2589 |
21.40 |
|
|
|
| 7 |
A |
1550 |
1471 |
2.53 |
|
|
|
| 8 |
A |
1542 |
1464 |
8.45 |
|
|
|
| 9 |
A |
1529 |
1452 |
1.44 |
|
|
|
| 10 |
A |
1460 |
1386 |
2.62 |
|
|
|
| 11 |
A |
1355 |
1286 |
23.62 |
|
|
|
| 12 |
A |
1316 |
1249 |
3.51 |
|
|
|
| 13 |
A |
1157 |
1098 |
12.58 |
|
|
|
| 14 |
A |
1096 |
1040 |
0.20 |
|
|
|
| 15 |
A |
1011 |
960 |
6.05 |
|
|
|
| 16 |
A |
896 |
850 |
8.97 |
|
|
|
| 17 |
A |
757 |
719 |
1.81 |
|
|
|
| 18 |
A |
674 |
640 |
4.04 |
|
|
|
| 19 |
A |
334 |
317 |
2.44 |
|
|
|
| 20 |
A |
272 |
258 |
3.02 |
|
|
|
| 21 |
A |
232 |
220 |
18.86 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16793.2 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 15940.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.643 |
-0.352 |
-0.054 |
| C2 |
0.499 |
0.646 |
0.091 |
| S3 |
-1.168 |
-0.099 |
-0.079 |
| H4 |
2.606 |
0.163 |
0.015 |
| H5 |
1.609 |
-1.107 |
0.736 |
| H6 |
1.592 |
-0.867 |
-1.016 |
| H7 |
0.558 |
1.178 |
1.043 |
| H8 |
0.542 |
1.398 |
-0.701 |
| H9 |
-1.067 |
-0.942 |
0.964 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5242 | 2.8222 | 1.0949 | 1.0937 | 1.0921 | 2.1725 | 2.1657 | 2.9541 |
C2 | 1.5242 | | 1.8341 | 2.1626 | 2.1726 | 2.1695 | 1.0922 | 1.0928 | 2.3952 | S3 | 2.8222 | 1.8341 | | 3.7845 | 3.0649 | 3.0143 | 2.4225 | 2.3559 | 1.3456 | H4 | 1.0949 | 2.1626 | 3.7845 | | 1.7688 | 1.7756 | 2.5060 | 2.5093 | 3.9514 | H5 | 1.0937 | 2.1726 | 3.0649 | 1.7688 | | 1.7684 | 2.5338 | 3.0786 | 2.6905 | H6 | 1.0921 | 2.1695 | 3.0143 | 1.7756 | 1.7684 | | 3.0803 | 2.5159 | 3.3162 | H7 | 2.1725 | 1.0922 | 2.4225 | 2.5060 | 2.5338 | 3.0803 | | 1.7578 | 2.6721 | H8 | 2.1657 | 1.0928 | 2.3559 | 2.5093 | 3.0786 | 2.5159 | 1.7578 | | 3.2918 | H9 | 2.9541 | 2.3952 | 1.3456 | 3.9514 | 2.6905 | 3.3162 | 2.6721 | 3.2918 | |
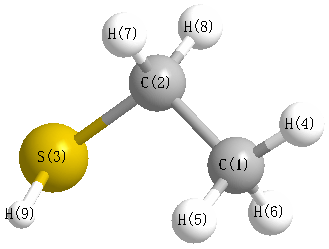 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
114.041 |
|
C1 |
C2 |
H7 |
111.194 |
| C1 |
C2 |
H8 |
110.617 |
|
C2 |
C1 |
H4 |
110.243 |
| C2 |
C1 |
H5 |
111.114 |
|
C2 |
C1 |
H6 |
110.965 |
| C2 |
S3 |
H9 |
96.547 |
|
S3 |
C2 |
H7 |
109.110 |
| S3 |
C2 |
H8 |
104.334 |
|
H4 |
C1 |
H5 |
107.842 |
| H4 |
C1 |
H6 |
108.561 |
|
H5 |
C1 |
H6 |
108.009 |
| H7 |
C2 |
H8 |
107.121 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.448 |
|
|
|
| 2 |
C |
-0.418 |
|
|
|
| 3 |
S |
-0.080 |
|
|
|
| 4 |
H |
0.157 |
|
|
|
| 5 |
H |
0.155 |
|
|
|
| 6 |
H |
0.173 |
|
|
|
| 7 |
H |
0.183 |
|
|
|
| 8 |
H |
0.185 |
|
|
|
| 9 |
H |
0.094 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.627 |
0.055 |
0.810 |
1.818 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.283 |
1.404 |
-0.848 |
| y |
1.404 |
-27.039 |
-1.709 |
| z |
-0.848 |
-1.709 |
-27.042 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.243 |
1.404 |
-0.848 |
| y |
1.404 |
1.123 |
-1.709 |
| z |
-0.848 |
-1.709 |
1.119 |
|
| Polar |
|---|
| 3z2-r2 | 2.239 |
|---|
| x2-y2 | -2.244 |
|---|
| xy | 1.404 |
|---|
| xz | -0.848 |
|---|
| yz | -1.709 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.593 |
0.459 |
0.031 |
| y |
0.459 |
5.039 |
-0.334 |
| z |
0.031 |
-0.334 |
4.850 |
<r2> (average value of r
2) Å
2
| <r2> |
84.415 |
| (<r2>)1/2 |
9.188 |
