Jump to
S1C2
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -169.401688 |
| Energy at 298.15K | |
| HF Energy | -168.929246 |
| Nuclear repulsion energy | 71.169108 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3799 |
3565 |
52.48 |
|
|
|
| 2 |
A' |
3667 |
3442 |
55.59 |
|
|
|
| 3 |
A' |
3062 |
2874 |
87.91 |
|
|
|
| 4 |
A' |
1898 |
1782 |
397.15 |
|
|
|
| 5 |
A' |
1671 |
1568 |
84.66 |
|
|
|
| 6 |
A' |
1476 |
1385 |
5.23 |
|
|
|
| 7 |
A' |
1320 |
1239 |
118.12 |
|
|
|
| 8 |
A' |
1093 |
1026 |
4.35 |
|
|
|
| 9 |
A' |
578 |
542 |
13.34 |
|
|
|
| 10 |
A" |
1073 |
1007 |
0.20 |
|
|
|
| 11 |
A" |
644 |
605 |
22.15 |
|
|
|
| 12 |
A" |
177i |
166i |
310.54 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10051.0 cm
-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 9433.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.421 |
0.000 |
| O2 |
1.199 |
0.230 |
0.000 |
| N3 |
-0.940 |
-0.560 |
0.000 |
| H4 |
-0.441 |
1.433 |
0.000 |
| H5 |
-0.642 |
-1.524 |
0.000 |
| H6 |
-1.925 |
-0.352 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2140 | 1.3594 | 1.1039 | 2.0483 | 2.0745 |
O2 | 1.2140 | | 2.2804 | 2.0340 | 2.5421 | 3.1774 | N3 | 1.3594 | 2.2804 | | 2.0554 | 1.0087 | 1.0064 | H4 | 1.1039 | 2.0340 | 2.0554 | | 2.9641 | 2.3217 | H5 | 2.0483 | 2.5421 | 1.0087 | 2.9641 | | 1.7379 | H6 | 2.0745 | 3.1774 | 1.0064 | 2.3217 | 1.7379 | |
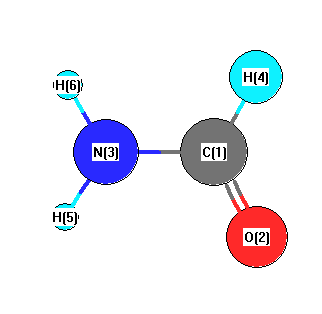 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.011 |
|
C1 |
N3 |
H6 |
121.815 |
| O2 |
C1 |
N3 |
124.683 |
|
O2 |
C1 |
H4 |
122.620 |
| N3 |
C1 |
H4 |
112.697 |
|
H5 |
N3 |
H6 |
119.173 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -169.401734 |
| Energy at 298.15K | -169.405470 |
| HF Energy | -168.929144 |
| Nuclear repulsion energy | 71.140577 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3783 |
3551 |
46.85 |
|
|
|
| 2 |
A |
3655 |
3431 |
48.94 |
|
|
|
| 3 |
A |
3063 |
2875 |
86.52 |
|
|
|
| 4 |
A |
1899 |
1783 |
379.75 |
|
|
|
| 5 |
A |
1675 |
1572 |
79.35 |
|
|
|
| 6 |
A |
1476 |
1386 |
4.76 |
|
|
|
| 7 |
A |
1319 |
1238 |
114.43 |
|
|
|
| 8 |
A |
1104 |
1036 |
4.84 |
|
|
|
| 9 |
A |
1069 |
1003 |
0.53 |
|
|
|
| 10 |
A |
638 |
598 |
37.90 |
|
|
|
| 11 |
A |
577 |
542 |
13.28 |
|
|
|
| 12 |
A |
246 |
231 |
317.44 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10252.5 cm
-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 9623.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.161 |
0.390 |
0.001 |
| O2 |
1.194 |
-0.248 |
0.006 |
| N3 |
-1.086 |
-0.158 |
-0.039 |
| H4 |
0.144 |
1.494 |
0.000 |
| H5 |
-1.166 |
-1.157 |
0.079 |
| H6 |
-1.900 |
0.408 |
0.139 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2136 | 1.3628 | 1.1038 | 2.0399 | 2.0664 |
O2 | 1.2136 | | 2.2824 | 2.0333 | 2.5304 | 3.1660 | N3 | 1.3628 | 2.2824 | | 2.0592 | 1.0097 | 1.0075 | H4 | 1.1038 | 2.0333 | 2.0592 | | 2.9578 | 2.3188 | H5 | 2.0399 | 2.5304 | 1.0097 | 2.9578 | | 1.7300 | H6 | 2.0664 | 3.1660 | 1.0075 | 2.3188 | 1.7300 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
117.822 |
|
C1 |
N3 |
H6 |
120.600 |
| O2 |
C1 |
N3 |
124.615 |
|
O2 |
C1 |
H4 |
122.584 |
| N3 |
C1 |
H4 |
112.779 |
|
H5 |
N3 |
H6 |
118.106 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
