Jump to
S1C2
Energy calculated at B3LYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -169.888843 |
| Energy at 298.15K | -169.892311 |
| HF Energy | -169.888843 |
| Nuclear repulsion energy | 71.001858 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3717 |
3570 |
33.54 |
|
|
|
| 2 |
A' |
3586 |
3444 |
29.58 |
|
|
|
| 3 |
A' |
2963 |
2845 |
114.14 |
|
|
|
| 4 |
A' |
1840 |
1767 |
356.29 |
|
|
|
| 5 |
A' |
1639 |
1574 |
65.58 |
|
|
|
| 6 |
A' |
1438 |
1381 |
4.98 |
|
|
|
| 7 |
A' |
1281 |
1230 |
96.73 |
|
|
|
| 8 |
A' |
1061 |
1019 |
2.97 |
|
|
|
| 9 |
A' |
564 |
542 |
11.42 |
|
|
|
| 10 |
A" |
1045 |
1004 |
0.06 |
|
|
|
| 11 |
A" |
652 |
626 |
19.43 |
|
|
|
| 12 |
A" |
86 |
82 |
265.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9935.9 cm
-1
Scaled (by 0.9603) Zero Point Vibrational Energy (zpe) 9541.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.420 |
0.000 |
| O2 |
1.202 |
0.237 |
0.000 |
| N3 |
-0.941 |
-0.565 |
0.000 |
| H4 |
-0.459 |
1.429 |
0.000 |
| H5 |
-0.643 |
-1.532 |
0.000 |
| H6 |
-1.928 |
-0.358 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2157 | 1.3624 | 1.1088 | 2.0550 | 2.0790 |
O2 | 1.2157 | | 2.2880 | 2.0445 | 2.5561 | 3.1859 | N3 | 1.3624 | 2.2880 | | 2.0522 | 1.0110 | 1.0089 | H4 | 1.1088 | 2.0445 | 2.0522 | | 2.9669 | 2.3134 | H5 | 2.0550 | 2.5561 | 1.0110 | 2.9669 | | 1.7404 | H6 | 2.0790 | 3.1859 | 1.0089 | 2.3134 | 1.7404 | |
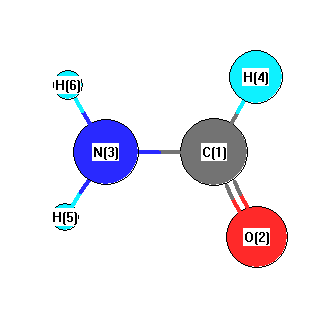 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.221 |
|
C1 |
N3 |
H6 |
121.784 |
| O2 |
C1 |
N3 |
125.018 |
|
O2 |
C1 |
H4 |
123.103 |
| N3 |
C1 |
H4 |
111.879 |
|
H5 |
N3 |
H6 |
118.995 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.363 |
|
|
|
| 2 |
O |
-0.447 |
|
|
|
| 3 |
N |
-0.698 |
|
|
|
| 4 |
H |
0.103 |
|
|
|
| 5 |
H |
0.344 |
|
|
|
| 6 |
H |
0.336 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.752 |
-0.773 |
0.000 |
3.831 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-3.752 |
-0.756 |
0.000 |
3.828 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.687 |
-0.025 |
0.000 |
| y |
-0.025 |
-14.607 |
0.000 |
| z |
0.000 |
0.000 |
-18.404 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.181 |
-0.025 |
0.000 |
| y |
-0.025 |
3.438 |
0.000 |
| z |
0.000 |
0.000 |
-2.257 |
|
| Polar |
|---|
| 3z2-r2 | -4.514 |
|---|
| x2-y2 | -3.080 |
|---|
| xy | -0.025 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.144 |
0.281 |
0.000 |
| y |
0.281 |
3.269 |
0.000 |
| z |
0.000 |
0.000 |
1.432 |
<r2> (average value of r
2) Å
2
| <r2> |
40.911 |
| (<r2>)1/2 |
6.396 |
Jump to
S1C1
Energy calculated at B3LYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -169.888843 |
| Energy at 298.15K | -169.892295 |
| HF Energy | -169.888843 |
| Nuclear repulsion energy | 71.002425 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3718 |
3570 |
33.56 |
|
|
|
| 2 |
A |
3586 |
3444 |
29.61 |
|
|
|
| 3 |
A |
2964 |
2846 |
113.93 |
|
|
|
| 4 |
A |
1839 |
1766 |
356.36 |
|
|
|
| 5 |
A |
1639 |
1574 |
65.37 |
|
|
|
| 6 |
A |
1437 |
1380 |
4.98 |
|
|
|
| 7 |
A |
1281 |
1231 |
96.58 |
|
|
|
| 8 |
A |
1061 |
1019 |
2.92 |
|
|
|
| 9 |
A |
1046 |
1004 |
0.06 |
|
|
|
| 10 |
A |
652 |
626 |
19.84 |
|
|
|
| 11 |
A |
565 |
542 |
11.45 |
|
|
|
| 12 |
A |
77 |
74 |
265.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9932.1 cm
-1
Scaled (by 0.9603) Zero Point Vibrational Energy (zpe) 9537.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.162 |
0.388 |
0.000 |
| O2 |
1.200 |
-0.246 |
-0.000 |
| N3 |
-1.086 |
-0.158 |
0.000 |
| H4 |
0.129 |
1.496 |
0.000 |
| H5 |
-1.184 |
-1.164 |
-0.000 |
| H6 |
-1.917 |
0.414 |
-0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2159 | 1.3622 | 1.1087 | 2.0545 | 2.0790 |
O2 | 1.2159 | | 2.2878 | 2.0443 | 2.5552 | 3.1860 | N3 | 1.3622 | 2.2878 | | 2.0525 | 1.0110 | 1.0089 | H4 | 1.1087 | 2.0443 | 2.0525 | | 2.9668 | 2.3144 | H5 | 2.0545 | 2.5552 | 1.0110 | 2.9668 | | 1.7405 | H6 | 2.0790 | 3.1860 | 1.0089 | 2.3144 | 1.7405 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.183 |
|
C1 |
N3 |
H6 |
121.812 |
| O2 |
C1 |
N3 |
125.002 |
|
O2 |
C1 |
H4 |
123.071 |
| N3 |
C1 |
H4 |
111.927 |
|
H5 |
N3 |
H6 |
119.005 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.363 |
|
|
|
| 2 |
O |
-0.448 |
|
|
|
| 3 |
N |
-0.698 |
|
|
|
| 4 |
H |
0.103 |
|
|
|
| 5 |
H |
0.344 |
|
|
|
| 6 |
H |
0.337 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.761 |
0.737 |
-0.001 |
3.832 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.245 |
1.080 |
0.002 |
| y |
1.080 |
-15.048 |
0.000 |
| z |
0.002 |
0.000 |
-18.404 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.519 |
1.080 |
0.002 |
| y |
1.080 |
2.777 |
0.000 |
| z |
0.002 |
0.000 |
-2.258 |
|
| Polar |
|---|
| 3z2-r2 | -4.515 |
|---|
| x2-y2 | -2.197 |
|---|
| xy | 1.080 |
|---|
| xz | 0.002 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.214 |
-0.115 |
0.000 |
| y |
-0.115 |
3.199 |
0.000 |
| z |
0.000 |
0.000 |
1.432 |
<r2> (average value of r
2) Å
2
| <r2> |
40.908 |
| (<r2>)1/2 |
6.396 |
