return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHBrCH3 (i-propyl bromide)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-2688.135233
Energy at 298.15K-2688.145768
HF Energy-2687.576687
Nuclear repulsion energy249.917790
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3172 3020 24.03      
2 A' 3148 2998 30.48      
3 A' 3133 2984 1.38      
4 A' 3068 2921 25.63      
5 A' 1553 1479 7.96      
6 A' 1543 1470 7.24      
7 A' 1473 1403 4.87      
8 A' 1297 1235 48.79      
9 A' 1222 1163 24.52      
10 A' 1086 1035 6.49      
11 A' 921 877 5.67      
12 A' 558 531 16.50      
13 A' 414 394 1.61      
14 A' 308 294 1.71      
15 A' 276 263 0.74      
16 A" 3167 3016 10.74      
17 A" 3140 2990 2.31      
18 A" 3063 2917 10.84      
19 A" 1535 1461 0.47      
20 A" 1532 1459 2.73      
21 A" 1462 1392 10.43      
22 A" 1403 1336 0.33      
23 A" 1184 1128 0.55      
24 A" 986 939 0.02      
25 A" 968 921 1.39      
26 A" 297 282 0.62      
27 A" 257 245 0.04      

Unscaled Zero Point Vibrational Energy (zpe) 21082.1 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 20076.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.26681 0.09587 0.07563

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.456 -0.911 0.000
Br2 -0.066 1.013 0.000
H3 1.549 -0.877 0.000
C4 -0.066 -1.560 1.273
C5 -0.066 -1.560 -1.273
H6 -1.161 -1.526 1.301
H7 0.251 -2.612 1.301
H8 0.320 -1.055 2.164
H9 -1.161 -1.526 -1.301
H10 0.251 -2.612 -1.301
H11 0.320 -1.055 -2.164

Atom - Atom Distances (Å)
  C1 Br2 H3 C4 C5 H6 H7 H8 H9 H10 H11
C11.99341.09351.52081.52082.16422.15122.17272.16422.15122.1727
Br21.99342.48542.87032.87033.05543.86423.01763.05543.86423.0176
H31.09352.48542.16662.16663.07522.52762.49463.07522.52762.4946
C41.52082.87032.16662.54521.09601.09901.09472.79672.79843.4946
C51.52082.87032.16662.54522.79672.79843.49461.09601.09901.0947
H62.16423.05543.07521.09602.79671.78151.77792.60113.15323.7970
H72.15123.86422.52761.09902.79841.78151.78143.15322.60233.7993
H82.17273.01762.49461.09473.49461.77791.78143.79703.79934.3274
H92.16423.05543.07522.79671.09602.60113.15323.79701.78151.7779
H102.15123.86422.52762.79841.09903.15322.60233.79931.78151.7814
H112.17273.01762.49463.49461.09473.79703.79934.32741.77791.7814

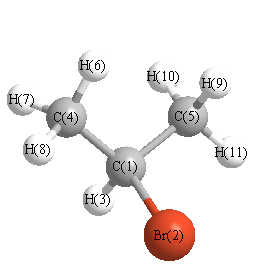
picture of i-propyl bromide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H6 110.543 C1 C4 H7 109.341
C1 C4 H8 111.299 C1 C5 H9 110.543
C1 C5 H10 109.341 C1 C5 H11 111.299
Br2 C1 H3 103.365 Br2 C1 C4 108.782
Br2 C1 C5 108.782 H3 C1 C4 110.882
H3 C1 C5 110.882 C4 C1 C5 113.604
H6 C4 H7 108.503 H6 C4 H8 108.498
H7 C4 H8 108.591 H9 C5 H10 108.503
H9 C5 H11 108.498 H10 C5 H11 108.591
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability