return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHClCH3 (Propane, 2-chloro-)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-577.661923
Energy at 298.15K-577.669807
HF Energy-577.169152
Nuclear repulsion energy166.479835
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3258 3009 27.47      
2 A' 3240 2993 38.02      
3 A' 3216 2970 1.73      
4 A' 3160 2919 17.71      
5 A' 1595 1474 8.34      
6 A' 1583 1462 6.46      
7 A' 1517 1401 6.77      
8 A' 1389 1283 40.95      
9 A' 1261 1165 12.82      
10 A' 1154 1066 15.14      
11 A' 955 882 9.77      
12 A' 682 630 29.08      
13 A' 445 411 1.77      
14 A' 358 330 1.25      
15 A' 288 266 0.10      
16 A" 3254 3005 12.65      
17 A" 3232 2986 1.99      
18 A" 3155 2914 12.21      
19 A" 1575 1455 0.79      
20 A" 1571 1451 2.10      
21 A" 1505 1390 10.91      
22 A" 1442 1332 0.53      
23 A" 1217 1124 1.85      
24 A" 1012 934 0.04      
25 A" 993 917 1.49      
26 A" 342 316 1.72      
27 A" 258 239 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 21827.2 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 20161.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.27087 0.15269 0.10729

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Cl1 -0.879 1.019 0.000
C2 0.562 -0.066 0.000
C3 0.562 -0.903 1.265
C4 0.562 -0.903 -1.265
H5 1.416 0.608 0.000
H6 1.455 -1.529 1.286
H7 1.455 -1.529 -1.286
H8 0.560 -0.274 2.153
H9 0.560 -0.274 -2.153
H10 -0.314 -1.548 1.297
H11 -0.314 -1.548 -1.297

Atom - Atom Distances (Å)
  Cl1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11
Cl11.80312.71432.71432.33093.68663.68662.89392.89392.93112.9311
C21.80311.51691.51691.08722.14332.14332.16282.16282.15642.1564
C32.71431.51692.53022.14711.09092.77471.08793.47531.08922.7841
C42.71431.51692.53022.14712.77471.09093.47531.08792.78411.0892
H52.33091.08722.14712.14712.49442.49442.47852.47853.05373.0537
H63.68662.14331.09092.77472.49442.57241.76853.76871.76933.1313
H73.68662.14332.77471.09092.49442.57243.76871.76853.13131.7693
H82.89392.16281.08793.47532.47851.76853.76874.30561.76693.7806
H92.89392.16283.47531.08792.47853.76871.76854.30563.78061.7669
H102.93112.15641.08922.78413.05371.76933.13131.76693.78062.5946
H112.93112.15642.78411.08923.05373.13131.76933.78061.76692.5946

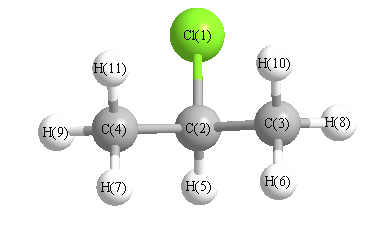
picture of Propane, 2-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Cl1 C2 C3 109.377 Cl1 C2 C4 109.377
Cl1 C3 H5 55.844 C2 C3 H6 109.458
C2 C3 H8 111.193 C2 C3 H10 110.602
C2 C4 H7 109.458 C2 C4 H9 111.193
C2 C4 H11 110.602 C3 C2 C4 113.022
C3 C2 H5 109.980 C4 C2 H5 109.980
H6 C3 H8 108.523 H6 C3 H10 108.497
H7 C4 H9 108.523 H7 C4 H11 108.497
H8 C3 H10 108.500 H9 C4 H11 108.500
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability