return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHClCH3 (Propane, 2-chloro-)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-577.746664
Energy at 298.15K-577.754462
HF Energy-577.168604
Nuclear repulsion energy165.734255
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3169 3018 27.87      
2 A' 3150 3000 38.81      
3 A' 3122 2973 3.51      
4 A' 3073 2926 18.41      
5 A' 1555 1481 8.21      
6 A' 1542 1469 6.15      
7 A' 1474 1404 6.31      
8 A' 1351 1286 37.07      
9 A' 1229 1170 11.70      
10 A' 1122 1069 13.47      
11 A' 931 887 8.31      
12 A' 659 628 24.87      
13 A' 435 414 1.66      
14 A' 349 332 1.19      
15 A' 283 269 0.11      
16 A" 3165 3014 13.06      
17 A" 3144 2994 2.23      
18 A" 3068 2922 12.39      
19 A" 1536 1462 0.65      
20 A" 1531 1458 2.14      
21 A" 1462 1392 10.74      
22 A" 1403 1337 0.66      
23 A" 1188 1131 1.86      
24 A" 986 939 0.01      
25 A" 967 921 1.39      
26 A" 333 317 1.63      
27 A" 255 242 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 21240.1 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 20226.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.26868 0.15139 0.10643

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Cl1 -0.887 1.019 0.000
C2 0.568 -0.065 0.000
C3 0.568 -0.904 1.269
C4 0.568 -0.904 -1.269
H5 1.423 0.618 0.000
H6 1.469 -1.532 1.291
H7 1.469 -1.532 -1.291
H8 0.563 -0.269 2.162
H9 0.563 -0.269 -2.162
H10 -0.313 -1.556 1.299
H11 -0.313 -1.556 -1.299

Atom - Atom Distances (Å)
  Cl1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11
Cl11.81442.72532.72532.34503.70503.70502.90462.90462.94082.9408
C21.81441.52181.52181.09472.15222.15222.17182.17182.16482.1648
C32.72531.52182.53902.15881.09842.78641.09533.48981.09642.7923
C42.72531.52182.53902.15882.78641.09843.48981.09532.79231.0964
H52.34501.09472.15882.15882.50852.50852.49062.49063.07063.0706
H63.70502.15221.09842.78642.50852.58271.78153.78711.78213.1439
H73.70502.15222.78641.09842.50852.58273.78711.78153.14391.7821
H82.90462.17181.09533.48982.49061.78153.78714.32431.77983.7948
H92.90462.17183.48981.09532.49063.78711.78154.32433.79481.7798
H102.94082.16481.09642.79233.07061.78213.14391.77983.79482.5975
H112.94082.16482.79231.09643.07063.14391.78213.79481.77982.5975

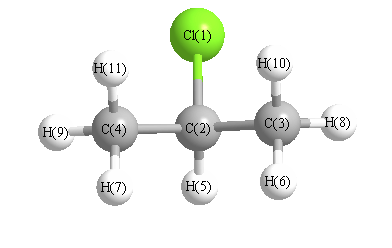
picture of Propane, 2-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Cl1 C2 C3 109.240 Cl1 C2 C4 109.240
Cl1 C3 H5 55.949 C2 C3 H6 109.383
C2 C3 H8 111.120 C2 C3 H10 110.500
C2 C4 H7 109.383 C2 C4 H9 111.120
C2 C4 H11 110.500 C3 C2 C4 113.066
C3 C2 H5 110.120 C4 C2 H5 110.120
H6 C3 H8 108.602 H6 C3 H10 108.575
H7 C4 H9 108.602 H7 C4 H11 108.575
H8 C3 H10 108.600 H9 C4 H11 108.600
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability