return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH(NH2)CH3 (2-Propanamine)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-173.897623
Energy at 298.15K-173.908227
HF Energy-173.284143
Nuclear repulsion energy135.385128
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3464 3299 2.61      
2 A' 3152 3002 53.64      
3 A' 3132 2983 48.09      
4 A' 3062 2916 4.17      
5 A' 2967 2825 73.37      
6 A' 1725 1643 35.98      
7 A' 1559 1485 6.08      
8 A' 1547 1473 2.85      
9 A' 1470 1400 15.04      
10 A' 1426 1358 16.23      
11 A' 1247 1188 4.19      
12 A' 1201 1144 23.07      
13 A' 1035 986 8.79      
14 A' 928 884 122.82      
15 A' 846 806 1.22      
16 A' 487 463 11.76      
17 A' 376 358 0.13      
18 A' 280 266 0.29      
19 A" 3551 3382 1.46      
20 A" 3146 2996 0.89      
21 A" 3129 2980 20.27      
22 A" 3058 2912 37.74      
23 A" 1539 1466 0.14      
24 A" 1538 1464 0.52      
25 A" 1476 1405 16.01      
26 A" 1435 1366 0.15      
27 A" 1303 1241 0.04      
28 A" 1076 1025 1.03      
29 A" 994 946 0.40      
30 A" 956 911 0.20      
31 A" 414 394 6.70      
32 A" 311 297 44.03      
33 A" 232 221 6.72      

Unscaled Zero Point Vibrational Energy (zpe) 27029.8 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 25740.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.27830 0.26558 0.15551

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.294 0.221 0.000
N2 -0.918 1.051 0.000
H3 1.223 0.824 0.000
C4 0.294 -0.640 1.261
C5 0.294 -0.640 -1.261
H6 -0.923 1.660 -0.818
H7 -0.923 1.660 0.818
H8 -0.604 -1.269 1.280
H9 -0.604 -1.269 -1.280
H10 1.182 -1.285 1.294
H11 1.182 -1.285 -1.294
H12 0.296 -0.016 2.165
H13 0.296 -0.016 -2.165

Atom - Atom Distances (Å)
  C1 N2 H3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.46891.10751.52671.52672.05502.05502.16052.16052.17492.17492.17742.1774
N21.46892.15262.43262.43261.02011.02012.66842.66843.39693.39692.70082.7008
H31.10752.15262.14412.14412.44432.44433.05973.05972.47502.47502.50022.5002
C41.52672.43262.14412.52103.33102.64001.09702.76741.09762.78001.09893.4816
C51.52672.43262.14412.52102.64003.33102.76741.09702.78001.09763.48161.0989
H62.05501.02012.44433.33102.64001.63653.61752.98264.19123.65123.63182.4710
H72.05501.02012.44432.64003.33101.63652.98263.61753.65124.19122.47103.6318
H82.16052.66843.05971.09702.76743.61752.98262.56051.78653.13331.77863.7747
H92.16052.66843.05972.76741.09702.98263.61752.56053.13331.78653.77471.7786
H102.17493.39692.47501.09762.78004.19123.65121.78653.13332.58781.77633.7892
H112.17493.39692.47502.78001.09763.65124.19123.13331.78652.58783.78921.7763
H122.17742.70082.50021.09893.48163.63182.47101.77863.77471.77633.78924.3291
H132.17742.70082.50023.48161.09892.47103.63183.77471.77863.78921.77634.3291

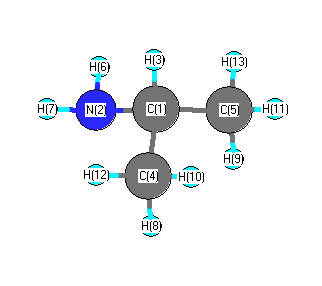
picture of 2-Propanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 H6 109.994 C1 N2 H7 109.994
C1 C4 H8 109.777 C1 C4 H10 110.881
C1 C4 H12 111.006 C1 C5 H9 109.777
C1 C5 H11 110.881 C1 C5 H13 111.006
N2 C1 H3 112.589 N2 C1 C4 108.583
N2 C1 C5 108.583 H3 C1 C4 107.906
H3 C1 C5 107.906 C4 C1 C5 111.306
H6 N2 H7 106.667 H8 C4 H10 108.980
H8 C4 H12 108.186 H9 C5 H11 108.980
H9 C5 H13 108.186 H10 C4 H12 107.935
H11 C5 H13 107.935
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability