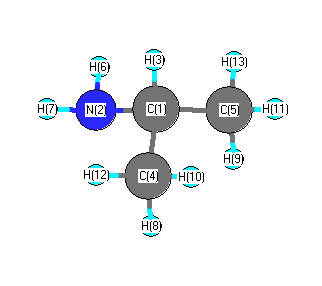
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3481 |
3304 |
2.52 |
|
|
|
| 2 |
A' |
3167 |
3006 |
55.31 |
|
|
|
| 3 |
A' |
3146 |
2986 |
49.46 |
|
|
|
| 4 |
A' |
3077 |
2921 |
3.70 |
|
|
|
| 5 |
A' |
2971 |
2820 |
83.29 |
|
|
|
| 6 |
A' |
1715 |
1628 |
32.62 |
|
|
|
| 7 |
A' |
1560 |
1481 |
4.99 |
|
|
|
| 8 |
A' |
1546 |
1468 |
3.39 |
|
|
|
| 9 |
A' |
1463 |
1389 |
11.98 |
|
|
|
| 10 |
A' |
1415 |
1343 |
21.32 |
|
|
|
| 11 |
A' |
1237 |
1174 |
4.90 |
|
|
|
| 12 |
A' |
1190 |
1129 |
20.92 |
|
|
|
| 13 |
A' |
1024 |
972 |
6.84 |
|
|
|
| 14 |
A' |
903 |
857 |
128.33 |
|
|
|
| 15 |
A' |
837 |
795 |
1.78 |
|
|
|
| 16 |
A' |
482 |
458 |
11.54 |
|
|
|
| 17 |
A' |
369 |
351 |
0.12 |
|
|
|
| 18 |
A' |
274 |
260 |
0.24 |
|
|
|
| 19 |
A" |
3570 |
3389 |
1.10 |
|
|
|
| 20 |
A" |
3161 |
3000 |
0.60 |
|
|
|
| 21 |
A" |
3143 |
2984 |
20.31 |
|
|
|
| 22 |
A" |
3073 |
2917 |
40.77 |
|
|
|
| 23 |
A" |
1540 |
1461 |
0.11 |
|
|
|
| 24 |
A" |
1536 |
1458 |
0.43 |
|
|
|
| 25 |
A" |
1461 |
1387 |
15.64 |
|
|
|
| 26 |
A" |
1429 |
1357 |
0.17 |
|
|
|
| 27 |
A" |
1297 |
1232 |
0.03 |
|
|
|
| 28 |
A" |
1063 |
1009 |
1.25 |
|
|
|
| 29 |
A" |
987 |
937 |
0.28 |
|
|
|
| 30 |
A" |
954 |
905 |
0.24 |
|
|
|
| 31 |
A" |
410 |
389 |
6.96 |
|
|
|
| 32 |
A" |
306 |
291 |
41.76 |
|
|
|
| 33 |
A" |
225 |
213 |
6.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27005.4 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 25633.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.029 |
|
|
|
| 2 |
N |
-0.738 |
|
|
|
| 3 |
H |
0.113 |
|
|
|
| 4 |
C |
-0.450 |
|
|
|
| 5 |
C |
-0.450 |
|
|
|
| 6 |
H |
0.300 |
|
|
|
| 7 |
H |
0.300 |
|
|
|
| 8 |
H |
0.162 |
|
|
|
| 9 |
H |
0.162 |
|
|
|
| 10 |
H |
0.145 |
|
|
|
| 11 |
H |
0.145 |
|
|
|
| 12 |
H |
0.140 |
|
|
|
| 13 |
H |
0.140 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.022 |
0.930 |
0.000 |
1.381 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.028 |
-0.512 |
0.000 |
| y |
-0.512 |
-24.475 |
0.000 |
| z |
0.000 |
0.000 |
-26.394 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.594 |
-0.512 |
0.000 |
| y |
-0.512 |
3.736 |
0.000 |
| z |
0.000 |
0.000 |
0.858 |
|
| Polar |
|---|
| 3z2-r2 | 1.715 |
|---|
| x2-y2 | -5.553 |
|---|
| xy | -0.512 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.528 |
-0.481 |
0.000 |
| y |
-0.481 |
5.875 |
0.000 |
| z |
0.000 |
0.000 |
6.250 |
<r2> (average value of r
2) Å
2
| <r2> |
93.221 |
| (<r2>)1/2 |
9.655 |