return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH(NH2)CH3 (2-Propanamine)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-173.886274
Energy at 298.15K-173.896894
HF Energy-173.284630
Nuclear repulsion energy135.583350
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3514 3298 1.23      
2 A' 3191 2995 47.98      
3 A' 3172 2978 42.90      
4 A' 3098 2908 3.70      
5 A' 3009 2824 70.66      
6 A' 1733 1627 38.90      
7 A' 1570 1473 6.82      
8 A' 1557 1461 3.29      
9 A' 1478 1388 17.05      
10 A' 1436 1348 15.86      
11 A' 1255 1178 4.36      
12 A' 1210 1136 24.29      
13 A' 1041 977 7.39      
14 A' 929 872 126.65      
15 A' 853 801 0.90      
16 A' 489 459 12.13      
17 A' 377 354 0.13      
18 A' 280 263 0.31      
19 A" 3600 3379 0.52      
20 A" 3186 2991 0.66      
21 A" 3170 2975 19.13      
22 A" 3095 2905 33.62      
23 A" 1550 1455 0.21      
24 A" 1548 1453 0.61      
25 A" 1485 1394 17.48      
26 A" 1443 1354 0.29      
27 A" 1310 1230 0.02      
28 A" 1082 1016 0.97      
29 A" 1000 938 0.47      
30 A" 961 902 0.26      
31 A" 416 391 6.85      
32 A" 310 291 44.10      
33 A" 232 218 7.05      

Unscaled Zero Point Vibrational Energy (zpe) 27290.4 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 25614.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.27906 0.26617 0.15584

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.294 0.219 0.000
N2 -0.920 1.049 0.000
H3 1.216 0.827 0.000
C4 0.294 -0.640 1.260
C5 0.294 -0.640 -1.260
H6 -0.911 1.662 -0.814
H7 -0.911 1.662 0.814
H8 -0.604 -1.264 1.281
H9 -0.604 -1.264 -1.281
H10 1.178 -1.285 1.293
H11 1.178 -1.285 -1.293
H12 0.298 -0.016 2.161
H13 0.298 -0.016 -2.161

Atom - Atom Distances (Å)
  C1 N2 H3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.47031.10451.52531.52532.04822.04822.15592.15592.17182.17182.17422.1742
N21.47032.14762.43172.43171.01901.01902.66252.66253.39413.39412.69962.6996
H31.10452.14762.14272.14272.42552.42553.05433.05432.47752.47752.49552.4955
C41.52532.43172.14272.52063.32442.63621.09432.76651.09462.77801.09593.4781
C51.52532.43172.14272.52062.63623.32442.76651.09432.77801.09463.47811.0959
H62.04821.01902.42553.32442.63621.62743.61142.97894.18183.64393.62322.4683
H72.04821.01902.42552.63623.32441.62742.97893.61143.64394.18182.46833.6232
H82.15592.66253.05431.09432.76653.61142.97892.56141.78203.13051.77383.7708
H92.15592.66253.05432.76651.09432.97893.61142.56143.13051.78203.77081.7738
H102.17183.39412.47751.09462.77804.18183.64391.78203.13052.58641.77183.7842
H112.17183.39412.47752.77801.09463.64394.18183.13051.78202.58643.78421.7718
H122.17422.69962.49551.09593.47813.62322.46831.77383.77081.77183.78424.3229
H132.17422.69962.49553.47811.09592.46833.62323.77081.77383.78421.77184.3229

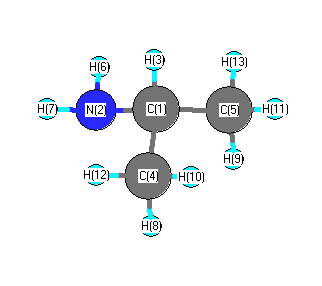
picture of 2-Propanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 H6 109.396 C1 N2 H7 109.396
C1 C4 H8 109.682 C1 C4 H10 110.918
C1 C4 H12 111.031 C1 C5 H9 109.682
C1 C5 H11 110.918 C1 C5 H13 111.031
N2 C1 H3 112.262 N2 C1 C4 108.523
N2 C1 C5 108.523 H3 C1 C4 108.065
H3 C1 C5 108.065 C4 C1 C5 111.436
H6 N2 H7 105.986 H8 C4 H10 108.995
H8 C4 H12 108.172 H9 C5 H11 108.995
H9 C5 H13 108.172 H10 C4 H12 107.967
H11 C5 H13 107.967
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability