return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6O (Propylene oxide)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-192.428368
Energy at 298.15K-192.435402
HF Energy-191.909211
Nuclear repulsion energy125.333300
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3313 3061 30.15      
2 A 3251 3003 35.39      
3 A 3234 2988 19.87      
4 A 3233 2987 8.42      
5 A 3224 2978 21.63      
6 A 3157 2916 15.50      
7 A 1635 1510 7.22      
8 A 1587 1466 5.69      
9 A 1573 1453 3.94      
10 A 1542 1424 17.59      
11 A 1494 1380 4.11      
12 A 1369 1265 4.50      
13 A 1267 1170 4.27      
14 A 1237 1143 3.23      
15 A 1230 1136 1.50      
16 A 1197 1105 8.02      
17 A 1102 1018 9.19      
18 A 1051 971 15.58      
19 A 960 887 2.48      
20 A 925 855 38.70      
21 A 840 775 5.76      
22 A 429 396 5.11      
23 A 384 355 4.50      
24 A 216 200 0.61      

Unscaled Zero Point Vibrational Energy (zpe) 19724.9 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 18219.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.61334 0.22429 0.20029

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.817 -0.781 -0.246
C2 -1.501 0.102 -0.147
H3 -1.398 0.334 -1.206
H4 -2.065 -0.826 -0.054
H5 -2.071 0.898 0.334
C6 -0.149 -0.041 0.488
H7 -0.150 -0.266 1.550
C8 1.039 0.610 -0.057
H9 0.952 1.216 -0.951
H10 1.868 0.863 0.592

Atom - Atom Distances (Å)
  O1 C2 H3 H4 H5 C6 H7 C8 H9 H10
O12.48272.65942.88823.39021.42052.10311.42132.12292.1239
C22.48271.08871.08961.09041.50102.20032.59202.81173.5325
H32.65941.08871.76541.77242.13763.08402.70822.52293.7657
H42.88821.08961.76541.76662.14052.55983.41973.75164.3286
H53.39021.09041.77241.76662.14462.55393.14763.30003.9476
C61.42051.50102.13762.14052.14461.08581.45952.20472.2123
H72.10312.20033.08402.55982.55391.08582.18243.10892.5027
C81.42132.59202.70823.41973.14761.45952.18241.08371.0833
H92.12292.81172.52293.75163.30002.20473.10891.08371.8293
H102.12393.53253.76574.32863.94762.21232.50271.08331.8293

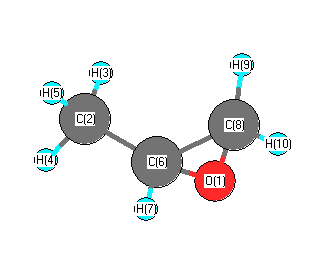
picture of Propylene oxide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C6 C2 116.347 O1 C6 H7 113.421
O1 C6 C8 59.125 O1 C8 C6 59.071
O1 C8 H9 115.210 O1 C8 H10 115.330
C2 C6 H7 115.612 C2 C6 C8 122.208
H3 C2 H4 108.276 H3 C2 H5 108.854
H3 C2 C6 110.247 H4 C2 H5 108.268
H4 C2 C6 110.420 H5 C2 C6 110.708
C6 O1 C8 61.804 C6 C8 H9 119.466
C6 C8 H10 120.205 H7 C6 C8 117.300
H9 C8 H10 115.171
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability