return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(CH3)3NH2 (2-Propanamine, 2-methyl-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-213.522385
Energy at 298.15K-213.535166
HF Energy-213.293321
Nuclear repulsion energy199.622837
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3464 3288 3.19      
2 A' 3171 3010 47.92      
3 A' 3146 2986 77.11      
4 A' 3136 2976 4.13      
5 A' 3076 2920 2.81      
6 A' 3060 2905 24.17      
7 A' 1711 1624 28.65      
8 A' 1565 1485 6.73      
9 A' 1547 1469 2.04      
10 A' 1532 1454 0.13      
11 A' 1469 1394 3.86      
12 A' 1441 1368 9.68      
13 A' 1314 1248 17.12      
14 A' 1277 1213 34.13      
15 A' 1085 1030 14.43      
16 A' 978 929 12.81      
17 A' 938 890 15.97      
18 A' 925 878 115.39      
19 A' 757 718 1.38      
20 A' 454 431 8.67      
21 A' 422 400 0.11      
22 A' 341 323 1.22      
23 A' 273 259 0.01      
24 A" 3554 3373 1.80      
25 A" 3165 3005 0.96      
26 A" 3141 2982 35.05      
27 A" 3134 2975 16.94      
28 A" 3070 2914 41.43      
29 A" 1555 1476 4.79      
30 A" 1536 1458 0.04      
31 A" 1527 1450 0.01      
32 A" 1446 1373 10.54      
33 A" 1387 1317 1.48      
34 A" 1158 1099 2.12      
35 A" 1035 983 1.17      
36 A" 981 931 0.01      
37 A" 955 906 0.23      
38 A" 456 433 7.36      
39 A" 350 332 9.13      
40 A" 308 292 31.81      
41 A" 287 272 3.25      
42 A" 216 205 3.52      

Unscaled Zero Point Vibrational Energy (zpe) 33170.9 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 31485.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.15335 0.15256 0.14933

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.011 0.011 0.000
N2 -0.547 1.386 0.000
C3 1.527 -0.020 0.000
C4 -0.547 -0.684 1.255
C5 -0.547 -0.684 -1.255
H6 -0.185 1.884 -0.812
H7 -0.185 1.884 0.812
H8 1.913 -1.045 0.000
H9 1.924 0.488 -0.886
H10 1.924 0.488 0.886
H11 -0.186 -0.185 2.161
H12 -0.186 -0.185 -2.161
H13 -1.639 -0.656 1.265
H14 -0.220 -1.727 1.296
H15 -1.639 -0.656 -1.265
H16 -0.220 -1.727 -1.296

Atom - Atom Distances (Å)
  C1 N2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.47581.53821.53151.53152.04962.04962.19432.18062.18062.17682.17682.16702.17792.16702.1779
N21.47582.50562.42062.42061.01991.01993.45822.77412.77412.69552.69552.63813.38772.63813.3877
C31.53822.50562.51372.51372.68632.68631.09521.09561.09562.76242.76243.46852.76523.46852.7652
C41.53152.42062.51372.50993.31712.63152.78503.47322.75951.09593.47111.09261.09402.74642.7751
C51.53152.42062.51372.50992.63153.31712.78502.75953.47323.47111.09592.74642.77511.09261.0940
H62.04961.01992.68633.31712.63151.62503.69332.52983.04623.62242.46963.58934.18212.96193.6440
H72.04961.01992.68632.63153.31711.62503.69333.04622.52982.46963.62242.96193.64403.58934.1821
H82.19433.45821.09522.78502.78503.69333.69331.77071.77073.13273.13273.79062.58713.79062.5871
H92.18062.77411.09563.47322.75952.52983.04621.77071.77223.76652.55484.31613.77663.76123.1097
H102.18062.77411.09562.75953.47323.04622.52981.77071.77222.55483.76653.76123.10974.31613.7766
H112.17682.69552.76241.09593.47113.62242.46963.13273.76652.55484.32181.77141.76903.75103.7854
H122.17682.69552.76243.47111.09592.46963.62243.13272.55483.76654.32183.75103.78541.77141.7690
H132.16702.63813.46851.09262.74643.58932.96193.79064.31613.76121.77143.75101.77882.52953.1176
H142.17793.38772.76521.09402.77514.18213.64402.58713.77663.10971.76903.78541.77883.11762.5915
H152.16702.63813.46852.74641.09262.96193.58933.79063.76124.31613.75101.77142.52953.11761.7788
H162.17793.38772.76522.77511.09403.64404.18212.58713.10973.77663.78541.76903.11762.59151.7788

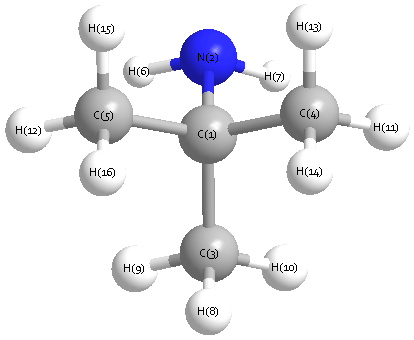
picture of 2-Propanamine, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 H6 109.047 C1 N2 H7 109.047
C1 C3 H8 111.758 C1 C3 H9 110.646
C1 C3 H10 110.646 C1 C4 H11 110.797
C1 C4 H13 110.217 C1 C4 H14 111.004
C1 C5 H12 110.797 C1 C5 H15 110.217
N2 C1 C3 112.450 N2 C1 C4 107.185
N2 C1 C5 107.185 C3 C1 C4 109.944
C3 C1 C5 109.944 C4 C1 C5 110.054
H6 N2 H7 105.617 H8 C3 H9 107.846
H8 C3 H10 107.846 H9 C3 H10 107.953
H11 C4 H13 108.082 H11 C4 H14 107.769
H12 C5 H15 108.082 H13 C4 H14 108.879
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.159      
2 N -0.754      
3 C -0.453      
4 C -0.445      
5 C -0.445      
6 H 0.303      
7 H 0.303      
8 H 0.148      
9 H 0.141      
10 H 0.141      
11 H 0.139      
12 H 0.139      
13 H 0.166      
14 H 0.145      
15 H 0.166      
16 H 0.145      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 7.082 0.039 0.000
y 0.039 7.494 0.000
z 0.000 0.000 7.557


<r2> (average value of r2) Å2
<r2> 129.956
(<r2>)1/2 11.400