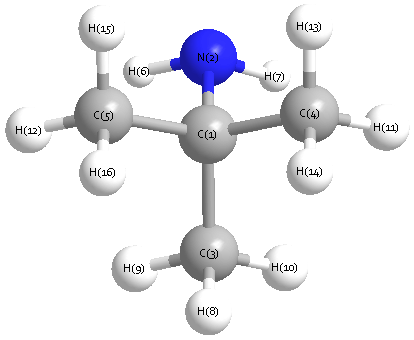
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3482 |
3312 |
3.00 |
|
|
|
| 2 |
A' |
3165 |
3010 |
45.10 |
|
|
|
| 3 |
A' |
3140 |
2986 |
71.92 |
|
|
|
| 4 |
A' |
3130 |
2977 |
5.44 |
|
|
|
| 5 |
A' |
3066 |
2915 |
2.77 |
|
|
|
| 6 |
A' |
3049 |
2900 |
26.30 |
|
|
|
| 7 |
A' |
1699 |
1616 |
30.63 |
|
|
|
| 8 |
A' |
1542 |
1467 |
7.60 |
|
|
|
| 9 |
A' |
1523 |
1448 |
2.36 |
|
|
|
| 10 |
A' |
1507 |
1433 |
0.15 |
|
|
|
| 11 |
A' |
1445 |
1375 |
6.05 |
|
|
|
| 12 |
A' |
1417 |
1348 |
14.71 |
|
|
|
| 13 |
A' |
1305 |
1241 |
17.30 |
|
|
|
| 14 |
A' |
1267 |
1205 |
37.92 |
|
|
|
| 15 |
A' |
1069 |
1016 |
13.70 |
|
|
|
| 16 |
A' |
974 |
926 |
2.52 |
|
|
|
| 17 |
A' |
933 |
887 |
30.24 |
|
|
|
| 18 |
A' |
914 |
869 |
114.82 |
|
|
|
| 19 |
A' |
763 |
726 |
0.99 |
|
|
|
| 20 |
A' |
449 |
427 |
9.02 |
|
|
|
| 21 |
A' |
413 |
393 |
0.10 |
|
|
|
| 22 |
A' |
336 |
319 |
1.15 |
|
|
|
| 23 |
A' |
264 |
251 |
0.01 |
|
|
|
| 24 |
A" |
3574 |
3399 |
1.20 |
|
|
|
| 25 |
A" |
3160 |
3005 |
0.67 |
|
|
|
| 26 |
A" |
3135 |
2982 |
31.39 |
|
|
|
| 27 |
A" |
3128 |
2975 |
18.73 |
|
|
|
| 28 |
A" |
3060 |
2910 |
43.40 |
|
|
|
| 29 |
A" |
1533 |
1458 |
5.12 |
|
|
|
| 30 |
A" |
1511 |
1437 |
0.02 |
|
|
|
| 31 |
A" |
1501 |
1427 |
0.01 |
|
|
|
| 32 |
A" |
1426 |
1357 |
15.95 |
|
|
|
| 33 |
A" |
1375 |
1307 |
0.51 |
|
|
|
| 34 |
A" |
1145 |
1089 |
2.39 |
|
|
|
| 35 |
A" |
1023 |
973 |
1.02 |
|
|
|
| 36 |
A" |
964 |
917 |
0.05 |
|
|
|
| 37 |
A" |
949 |
902 |
0.61 |
|
|
|
| 38 |
A" |
452 |
430 |
8.04 |
|
|
|
| 39 |
A" |
345 |
328 |
10.57 |
|
|
|
| 40 |
A" |
305 |
290 |
31.44 |
|
|
|
| 41 |
A" |
283 |
269 |
1.79 |
|
|
|
| 42 |
A" |
208 |
198 |
3.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32963.4 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 31348.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.124 |
|
|
|
| 2 |
N |
-0.728 |
|
|
|
| 3 |
C |
-0.501 |
|
|
|
| 4 |
C |
-0.492 |
|
|
|
| 5 |
C |
-0.492 |
|
|
|
| 6 |
H |
0.306 |
|
|
|
| 7 |
H |
0.306 |
|
|
|
| 8 |
H |
0.164 |
|
|
|
| 9 |
H |
0.157 |
|
|
|
| 10 |
H |
0.157 |
|
|
|
| 11 |
H |
0.155 |
|
|
|
| 12 |
H |
0.155 |
|
|
|
| 13 |
H |
0.182 |
|
|
|
| 14 |
H |
0.163 |
|
|
|
| 15 |
H |
0.182 |
|
|
|
| 16 |
H |
0.163 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.296 |
0.125 |
0.000 |
1.302 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.421 |
2.272 |
0.000 |
| y |
2.272 |
-32.089 |
0.000 |
| z |
0.000 |
0.000 |
-32.652 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.050 |
2.272 |
0.000 |
| y |
2.272 |
2.447 |
0.000 |
| z |
0.000 |
0.000 |
1.603 |
|
| Polar |
|---|
| 3z2-r2 | 3.206 |
|---|
| x2-y2 | -4.331 |
|---|
| xy | 2.272 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.192 |
0.041 |
0.000 |
| y |
0.041 |
7.568 |
0.000 |
| z |
0.000 |
0.000 |
7.655 |
<r2> (average value of r
2) Å
2
| <r2> |
129.252 |
| (<r2>)1/2 |
11.369 |