Jump to
S1C2
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
Geometric Data calculated at PBEPBEultrafine/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -233.353759 |
| Energy at 298.15K | -233.364769 |
| HF Energy | -233.353759 |
| Nuclear repulsion energy | 198.119950 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3611 |
3551 |
1.66 |
|
|
|
| 2 |
A |
3077 |
3026 |
42.30 |
|
|
|
| 3 |
A |
3074 |
3023 |
33.59 |
|
|
|
| 4 |
A |
3069 |
3018 |
10.57 |
|
|
|
| 5 |
A |
3067 |
3016 |
5.83 |
|
|
|
| 6 |
A |
3047 |
2996 |
57.32 |
|
|
|
| 7 |
A |
3040 |
2990 |
2.74 |
|
|
|
| 8 |
A |
2994 |
2944 |
9.79 |
|
|
|
| 9 |
A |
2975 |
2926 |
32.44 |
|
|
|
| 10 |
A |
2970 |
2921 |
18.53 |
|
|
|
| 11 |
A |
1498 |
1473 |
7.53 |
|
|
|
| 12 |
A |
1488 |
1463 |
5.01 |
|
|
|
| 13 |
A |
1479 |
1454 |
2.77 |
|
|
|
| 14 |
A |
1466 |
1442 |
0.03 |
|
|
|
| 15 |
A |
1462 |
1437 |
0.04 |
|
|
|
| 16 |
A |
1453 |
1429 |
0.03 |
|
|
|
| 17 |
A |
1399 |
1376 |
2.70 |
|
|
|
| 18 |
A |
1380 |
1357 |
32.65 |
|
|
|
| 19 |
A |
1370 |
1347 |
12.56 |
|
|
|
| 20 |
A |
1347 |
1325 |
24.05 |
|
|
|
| 21 |
A |
1234 |
1214 |
26.22 |
|
|
|
| 22 |
A |
1212 |
1192 |
48.42 |
|
|
|
| 23 |
A |
1141 |
1122 |
53.56 |
|
|
|
| 24 |
A |
1022 |
1005 |
1.26 |
|
|
|
| 25 |
A |
1006 |
989 |
8.85 |
|
|
|
| 26 |
A |
935 |
920 |
0.00 |
|
|
|
| 27 |
A |
915 |
900 |
25.93 |
|
|
|
| 28 |
A |
904 |
889 |
10.03 |
|
|
|
| 29 |
A |
890 |
875 |
0.09 |
|
|
|
| 30 |
A |
739 |
727 |
1.90 |
|
|
|
| 31 |
A |
451 |
444 |
12.61 |
|
|
|
| 32 |
A |
445 |
438 |
8.54 |
|
|
|
| 33 |
A |
400 |
393 |
0.54 |
|
|
|
| 34 |
A |
332 |
327 |
36.46 |
|
|
|
| 35 |
A |
328 |
322 |
1.18 |
|
|
|
| 36 |
A |
312 |
307 |
63.59 |
|
|
|
| 37 |
A |
268 |
263 |
0.04 |
|
|
|
| 38 |
A |
256 |
252 |
3.08 |
|
|
|
| 39 |
A |
203 |
200 |
2.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29126.5 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 28645.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.004 |
-0.000 |
0.022 |
| H2 |
-1.759 |
-1.284 |
-0.188 |
| H3 |
-0.208 |
-2.168 |
-0.101 |
| H4 |
-0.679 |
-1.320 |
-1.606 |
| C5 |
-0.699 |
-1.267 |
-0.504 |
| H6 |
1.989 |
-0.893 |
0.027 |
| H7 |
1.989 |
0.892 |
0.026 |
| H8 |
1.596 |
-0.000 |
-1.476 |
| C9 |
1.486 |
-0.000 |
-0.379 |
| H10 |
-0.207 |
2.168 |
-0.101 |
| H11 |
-1.758 |
1.284 |
-0.189 |
| H12 |
-0.679 |
1.321 |
-1.606 |
| C13 |
-0.699 |
1.268 |
-0.504 |
| H14 |
-0.920 |
0.000 |
1.748 |
| O15 |
0.011 |
0.000 |
1.457 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
H4 |
C5 |
H6 |
H7 |
H8 |
C9 |
H10 |
H11 |
H12 |
C13 |
H14 |
O15 |
| C1 | | 2.1911 | 2.1820 | 2.2047 | 1.5421 | 2.1763 | 2.1763 | 2.1855 | 1.5344 | 2.1820 | 2.1911 | 2.2047 | 1.5421 | 1.9583 | 1.4356 |
H2 | 2.1911 | | 1.7878 | 1.7823 | 1.1054 | 3.7742 | 4.3394 | 3.8159 | 3.4942 | 3.7856 | 2.5677 | 3.1558 | 2.7807 | 2.4697 | 2.7362 | H3 | 2.1820 | 1.7878 | | 1.7910 | 1.1025 | 2.5431 | 3.7698 | 3.1377 | 2.7649 | 4.3364 | 3.7856 | 3.8290 | 3.4942 | 2.9370 | 2.6788 | H4 | 2.2047 | 1.7823 | 1.7910 | | 1.1038 | 3.1574 | 3.8320 | 2.6341 | 2.8170 | 3.8289 | 3.1555 | 2.6412 | 2.8132 | 3.6125 | 3.4064 | C5 | 1.5421 | 1.1054 | 1.1025 | 1.1038 | | 2.7657 | 3.4895 | 2.7966 | 2.5290 | 3.4942 | 2.7806 | 2.8133 | 2.5353 | 2.5933 | 2.4406 | H6 | 2.1763 | 3.7742 | 2.5431 | 3.1574 | 2.7657 | | 1.7857 | 1.7913 | 1.1024 | 3.7697 | 4.3394 | 3.8320 | 3.4895 | 3.4964 | 2.5996 | H7 | 2.1763 | 4.3394 | 3.7698 | 3.8320 | 3.4895 | 1.7857 | | 1.7913 | 1.1024 | 2.5431 | 3.7742 | 3.1571 | 2.7657 | 3.4965 | 2.5997 | H8 | 2.1855 | 3.8159 | 3.1377 | 2.6341 | 2.7966 | 1.7913 | 1.7913 | | 1.1023 | 3.1379 | 3.8159 | 2.6341 | 2.7967 | 4.0897 | 3.3342 | C9 | 1.5344 | 3.4942 | 2.7649 | 2.8170 | 2.5290 | 1.1024 | 1.1024 | 1.1023 | | 2.7650 | 3.4942 | 2.8169 | 2.5290 | 3.2113 | 2.3553 | H10 | 2.1820 | 3.7856 | 4.3364 | 3.8289 | 3.4942 | 3.7697 | 2.5431 | 3.1379 | 2.7650 | | 1.7878 | 1.7910 | 1.1025 | 2.9369 | 2.6786 | H11 | 2.1911 | 2.5677 | 3.7856 | 3.1555 | 2.7806 | 4.3394 | 3.7742 | 3.8159 | 3.4942 | 1.7878 | | 1.7823 | 1.1054 | 2.4698 | 2.7363 | H12 | 2.2047 | 3.1558 | 3.8290 | 2.6412 | 2.8133 | 3.8320 | 3.1571 | 2.6341 | 2.8169 | 1.7910 | 1.7823 | | 1.1038 | 3.6126 | 3.4064 | C13 | 1.5421 | 2.7807 | 3.4942 | 2.8132 | 2.5353 | 3.4895 | 2.7657 | 2.7967 | 2.5290 | 1.1025 | 1.1054 | 1.1038 | | 2.5933 | 2.4406 | H14 | 1.9583 | 2.4697 | 2.9370 | 3.6125 | 2.5933 | 3.4964 | 3.4965 | 4.0897 | 3.2113 | 2.9369 | 2.4698 | 3.6126 | 2.5933 | | 0.9754 | O15 | 1.4356 | 2.7362 | 2.6788 | 3.4064 | 2.4406 | 2.5996 | 2.5997 | 3.3342 | 2.3553 | 2.6786 | 2.7363 | 3.4064 | 2.4406 | 0.9754 | |
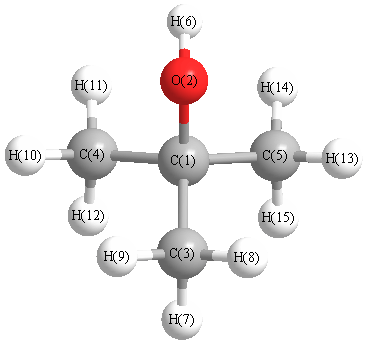 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C5 |
H2 |
110.621 |
|
C1 |
C5 |
H3 |
110.079 |
| C1 |
C5 |
H4 |
111.798 |
|
C1 |
C9 |
H6 |
110.172 |
| C1 |
C9 |
H7 |
110.173 |
|
C1 |
C9 |
H8 |
110.903 |
| C1 |
C13 |
H10 |
110.077 |
|
C1 |
C13 |
H11 |
110.621 |
| C1 |
C13 |
H12 |
111.798 |
|
C1 |
O15 |
H14 |
107.078 |
| H2 |
C5 |
H3 |
108.134 |
|
H2 |
C5 |
H4 |
107.556 |
| H3 |
C5 |
H4 |
108.536 |
|
C5 |
C1 |
C9 |
110.580 |
| C5 |
C1 |
C13 |
110.575 |
|
C5 |
C1 |
O15 |
110.040 |
| H6 |
C9 |
H7 |
108.171 |
|
H6 |
C9 |
H8 |
108.677 |
| H7 |
C9 |
H8 |
108.677 |
|
C9 |
C1 |
C13 |
110.579 |
| C9 |
C1 |
O15 |
104.889 |
|
H10 |
C13 |
H11 |
108.134 |
| H10 |
C13 |
H12 |
108.537 |
|
H11 |
C13 |
H12 |
107.557 |
| C13 |
C1 |
O15 |
110.039 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.257 |
|
|
|
| 2 |
H |
0.141 |
|
|
|
| 3 |
H |
0.167 |
|
|
|
| 4 |
H |
0.153 |
|
|
|
| 5 |
C |
-0.485 |
|
|
|
| 6 |
H |
0.165 |
|
|
|
| 7 |
H |
0.165 |
|
|
|
| 8 |
H |
0.151 |
|
|
|
| 9 |
C |
-0.473 |
|
|
|
| 10 |
H |
0.167 |
|
|
|
| 11 |
H |
0.141 |
|
|
|
| 12 |
H |
0.153 |
|
|
|
| 13 |
C |
-0.485 |
|
|
|
| 14 |
H |
0.375 |
|
|
|
| 15 |
O |
-0.590 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.174 |
0.000 |
-0.852 |
1.450 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.379 |
0.004 |
2.775 |
| y |
0.004 |
-33.148 |
0.004 |
| z |
2.775 |
0.004 |
-33.907 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.149 |
0.004 |
2.775 |
| y |
0.004 |
-0.505 |
0.004 |
| z |
2.775 |
0.004 |
-1.643 |
|
| Polar |
|---|
| 3z2-r2 | -3.287 |
|---|
| x2-y2 | 1.769 |
|---|
| xy | 0.004 |
|---|
| xz | 2.775 |
|---|
| yz | 0.004 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.477 |
0.000 |
0.164 |
| y |
0.000 |
7.117 |
0.000 |
| z |
0.164 |
0.000 |
7.047 |
<r2> (average value of r
2) Å
2
| <r2> |
126.547 |
| (<r2>)1/2 |
11.249 |
