return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(CH3)3SH (2-Propanethiol, 2-methyl-)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-555.398175
Energy at 298.15K-555.409263
HF Energy-554.805945
Nuclear repulsion energy245.571130
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3252 3004 27.11      
2 A' 3237 2990 30.99      
3 A' 3228 2981 38.26      
4 A' 3162 2921 20.23      
5 A' 3154 2913 28.09      
6 A' 2830 2614 21.51      
7 A' 1610 1487 8.58      
8 A' 1596 1475 9.44      
9 A' 1583 1462 0.31      
10 A' 1531 1414 1.39      
11 A' 1503 1389 10.46      
12 A' 1348 1245 0.06      
13 A' 1290 1191 39.96      
14 A' 1128 1042 2.58      
15 A' 1007 930 1.17      
16 A' 937 865 6.27      
17 A' 889 821 1.47      
18 A' 637 588 4.67      
19 A' 417 385 1.18      
20 A' 390 360 0.19      
21 A' 320 295 0.22      
22 A' 301 278 0.96      
23 A" 3251 3003 27.24      
24 A" 3246 2998 4.26      
25 A" 3222 2976 0.72      
26 A" 3150 2910 15.89      
27 A" 1598 1476 7.73      
28 A" 1580 1459 0.41      
29 A" 1573 1453 0.03      
30 A" 1504 1389 9.80      
31 A" 1332 1230 2.46      
32 A" 1114 1029 0.07      
33 A" 1028 949 0.01      
34 A" 999 923 0.55      
35 A" 423 391 0.50      
36 A" 322 298 2.36      
37 A" 301 278 0.01      
38 A" 270 250 6.74      
39 A" 219 203 14.06      

Unscaled Zero Point Vibrational Energy (zpe) 30240.4 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 27933.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.15074 0.10034 0.09942

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.341 -0.007 0.000
S2 -1.495 0.088 0.000
C3 0.831 1.440 0.000
C4 0.831 -0.727 1.253
C5 0.831 -0.727 -1.253
H6 -1.728 -1.228 0.000
H7 1.921 1.457 0.000
H8 0.483 1.974 -0.883
H9 0.483 1.974 0.883
H10 1.922 -0.749 1.266
H11 1.922 -0.749 -1.266
H12 0.484 -0.224 2.153
H13 0.476 -1.757 1.280
H14 0.484 -0.224 -2.153
H15 0.476 -1.757 -1.280

Atom - Atom Distances (Å)
  C1 S2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.83871.52761.52611.52612.40252.15362.17322.17322.15722.15722.16902.17282.16902.1728
S21.83872.69052.76512.76511.33693.68012.87242.87243.73913.73912.94162.98832.94162.9883
C31.52762.69052.50362.50363.69701.09011.08891.08892.75422.75422.74373.46242.74373.4624
C41.52612.76512.50362.50592.89262.74373.46122.74821.09132.74551.08791.08963.46062.7576
C51.52612.76512.50362.50592.89262.74372.74823.46122.74551.09133.46062.75761.08791.0896
H62.40251.33693.69702.89262.89264.53013.99033.99033.89263.89263.24612.60283.24612.6028
H72.15363.68011.09012.74372.74374.53011.76471.76472.54332.54333.08653.74913.08653.7491
H82.17322.87241.08893.46122.74823.99031.76471.76673.75543.10303.74874.31292.53863.7519
H92.17322.87241.08892.74823.46123.99031.76471.76673.10303.75542.53863.75193.74874.3129
H102.15723.73912.75421.09132.74553.89262.54333.75543.10302.53261.76871.76283.74643.0972
H112.15723.73912.75422.74551.09133.89262.54333.10303.75542.53263.74643.09721.76871.7628
H122.16902.94162.74371.08793.46063.24613.08653.74872.53861.76873.74641.76384.30663.7602
H132.17282.98833.46241.08962.75762.60283.74914.31293.75191.76283.09721.76383.76022.5607
H142.16902.94162.74373.46061.08793.24613.08652.53863.74873.74641.76874.30663.76021.7638
H152.17282.98833.46242.75761.08962.60283.74913.75194.31293.09721.76283.76022.56071.7638

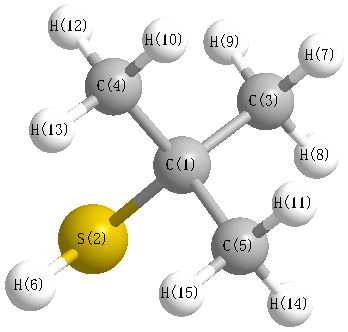
picture of 2-Propanethiol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 H6 97.053 C1 C3 H7 109.571
C1 C3 H8 111.206 C1 C3 H9 111.206
C1 C4 H10 109.892 C1 C4 H12 111.037
C1 C4 H13 111.241 C1 C5 H11 109.892
C1 C5 H14 111.037 C1 C5 H15 111.241
S2 C1 C3 105.745 S2 C1 C4 110.176
S2 C1 C5 110.176 C3 C1 C4 110.138
C3 C1 C5 110.138 C4 C1 C5 110.373
H7 C3 H8 108.161 H7 C3 H9 108.161
H8 C3 H9 108.434 H10 C4 H12 108.511
H10 C4 H13 107.859 H11 C5 H14 108.511
H11 C5 H15 107.859 H12 C4 H13 108.198
H14 C5 H15 108.198
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability