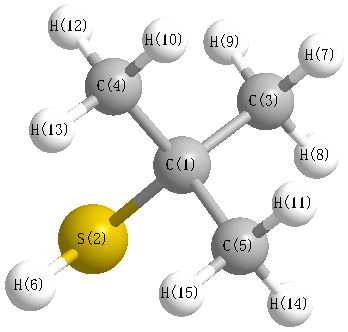
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3150 |
3013 |
24.81 |
|
|
|
| 2 |
A' |
3133 |
2997 |
25.14 |
|
|
|
| 3 |
A' |
3121 |
2986 |
43.90 |
|
|
|
| 4 |
A' |
3058 |
2926 |
21.64 |
|
|
|
| 5 |
A' |
3049 |
2917 |
34.81 |
|
|
|
| 6 |
A' |
2702 |
2585 |
24.35 |
|
|
|
| 7 |
A' |
1538 |
1471 |
8.18 |
|
|
|
| 8 |
A' |
1524 |
1458 |
9.29 |
|
|
|
| 9 |
A' |
1509 |
1444 |
0.62 |
|
|
|
| 10 |
A' |
1450 |
1388 |
1.09 |
|
|
|
| 11 |
A' |
1421 |
1360 |
11.35 |
|
|
|
| 12 |
A' |
1272 |
1217 |
0.74 |
|
|
|
| 13 |
A' |
1209 |
1156 |
46.87 |
|
|
|
| 14 |
A' |
1068 |
1022 |
2.23 |
|
|
|
| 15 |
A' |
952 |
911 |
0.98 |
|
|
|
| 16 |
A' |
888 |
849 |
4.85 |
|
|
|
| 17 |
A' |
836 |
800 |
1.63 |
|
|
|
| 18 |
A' |
592 |
567 |
5.72 |
|
|
|
| 19 |
A' |
389 |
372 |
1.09 |
|
|
|
| 20 |
A' |
365 |
349 |
0.47 |
|
|
|
| 21 |
A' |
295 |
283 |
0.28 |
|
|
|
| 22 |
A' |
280 |
268 |
0.85 |
|
|
|
| 23 |
A" |
3148 |
3012 |
25.16 |
|
|
|
| 24 |
A" |
3143 |
3007 |
5.87 |
|
|
|
| 25 |
A" |
3116 |
2981 |
0.83 |
|
|
|
| 26 |
A" |
3045 |
2913 |
18.31 |
|
|
|
| 27 |
A" |
1526 |
1460 |
7.99 |
|
|
|
| 28 |
A" |
1506 |
1441 |
0.51 |
|
|
|
| 29 |
A" |
1498 |
1434 |
0.01 |
|
|
|
| 30 |
A" |
1422 |
1360 |
10.06 |
|
|
|
| 31 |
A" |
1259 |
1205 |
4.63 |
|
|
|
| 32 |
A" |
1057 |
1012 |
0.09 |
|
|
|
| 33 |
A" |
974 |
932 |
0.01 |
|
|
|
| 34 |
A" |
945 |
904 |
0.49 |
|
|
|
| 35 |
A" |
397 |
380 |
0.54 |
|
|
|
| 36 |
A" |
300 |
287 |
4.00 |
|
|
|
| 37 |
A" |
282 |
270 |
0.00 |
|
|
|
| 38 |
A" |
257 |
246 |
9.46 |
|
|
|
| 39 |
A" |
214 |
205 |
9.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28945.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 27691.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.120 |
|
|
|
| 2 |
S |
-0.089 |
|
|
|
| 3 |
C |
-0.507 |
|
|
|
| 4 |
C |
-0.484 |
|
|
|
| 5 |
C |
-0.484 |
|
|
|
| 6 |
H |
0.098 |
|
|
|
| 7 |
H |
0.174 |
|
|
|
| 8 |
H |
0.185 |
|
|
|
| 9 |
H |
0.185 |
|
|
|
| 10 |
H |
0.167 |
|
|
|
| 11 |
H |
0.167 |
|
|
|
| 12 |
H |
0.186 |
|
|
|
| 13 |
H |
0.167 |
|
|
|
| 14 |
H |
0.186 |
|
|
|
| 15 |
H |
0.167 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.661 |
-0.864 |
0.000 |
1.872 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.956 |
2.368 |
0.000 |
| y |
2.368 |
-38.653 |
0.000 |
| z |
0.000 |
0.000 |
-41.568 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.846 |
2.368 |
0.000 |
| y |
2.368 |
3.109 |
0.000 |
| z |
0.000 |
0.000 |
-1.263 |
|
| Polar |
|---|
| 3z2-r2 | -2.526 |
|---|
| x2-y2 | -3.304 |
|---|
| xy | 2.368 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.940 |
0.123 |
0.000 |
| y |
0.123 |
8.500 |
0.000 |
| z |
0.000 |
0.000 |
7.852 |
<r2> (average value of r
2) Å
2
| <r2> |
165.137 |
| (<r2>)1/2 |
12.851 |