return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(CH3)3SH (2-Propanethiol, 2-methyl-)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-555.526601
Energy at 298.15K-555.537521
HF Energy-554.804969
Nuclear repulsion energy244.472173
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3155 3004 27.06      
2 A' 3140 2990 29.96      
3 A' 3130 2981 39.77      
4 A' 3066 2919 18.86      
5 A' 3058 2912 29.07      
6 A' 2704 2575 26.00      
7 A' 1563 1489 8.39      
8 A' 1550 1476 9.08      
9 A' 1537 1464 0.28      
10 A' 1482 1411 1.29      
11 A' 1455 1385 10.50      
12 A' 1309 1246 0.11      
13 A' 1250 1190 35.78      
14 A' 1092 1040 2.23      
15 A' 978 931 0.86      
16 A' 906 863 5.57      
17 A' 862 821 1.16      
18 A' 616 587 3.89      
19 A' 407 387 1.13      
20 A' 380 362 0.15      
21 A' 311 296 0.22      
22 A' 294 280 0.88      
23 A" 3153 3003 27.56      
24 A" 3148 2998 4.42      
25 A" 3125 2976 0.72      
26 A" 3054 2909 15.91      
27 A" 1551 1477 7.43      
28 A" 1534 1461 0.38      
29 A" 1527 1455 0.02      
30 A" 1455 1386 9.55      
31 A" 1294 1232 2.40      
32 A" 1080 1028 0.08      
33 A" 996 949 0.01      
34 A" 970 924 0.45      
35 A" 413 393 0.54      
36 A" 314 299 2.13      
37 A" 293 279 0.03      
38 A" 262 249 5.15      
39 A" 209 199 13.85      

Unscaled Zero Point Vibrational Energy (zpe) 29310.2 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 27912.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.14949 0.09955 0.09862

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.344 -0.007 0.000
S2 -1.502 0.090 0.000
C3 0.834 1.446 0.000
C4 0.834 -0.730 1.258
C5 0.834 -0.730 -1.258
H6 -1.719 -1.242 0.000
H7 1.931 1.462 0.000
H8 0.482 1.982 -0.890
H9 0.482 1.982 0.890
H10 1.933 -0.752 1.272
H11 1.933 -0.752 -1.272
H12 0.483 -0.223 2.164
H13 0.475 -1.767 1.284
H14 0.483 -0.223 -2.164
H15 0.475 -1.767 -1.284

Atom - Atom Distances (Å)
  C1 S2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.84801.53271.53141.53142.40432.16272.18302.18302.16732.16732.17892.18292.17892.1829
S21.84802.70032.77612.77611.34913.69712.88252.88253.75763.75762.95253.00052.95253.0005
C31.53272.70032.51322.51323.70641.09801.09691.09692.76692.76692.75483.47842.75483.4784
C41.53142.77612.51322.51562.89122.75553.47732.75911.09932.75821.09581.09763.47662.7685
C51.53142.77612.51322.51562.89122.75552.75913.47732.75821.09933.47662.76851.09581.0976
H62.40431.34913.70642.89122.89124.54264.00374.00373.89753.89753.25082.59573.25082.5957
H72.16273.69711.09802.75552.75554.54261.77821.77822.55352.55353.10153.76803.10153.7680
H82.18302.88251.09693.47732.75914.00371.77821.78033.77513.11803.76664.33372.54623.7695
H92.18302.88251.09692.75913.47734.00371.77821.78033.11803.77512.54623.76953.76664.3337
H102.16733.75762.76691.09932.75823.89752.55353.77513.11802.54391.78231.77633.76633.1125
H112.16733.75762.76692.75821.09933.89752.55353.11803.77512.54393.76633.11251.78231.7763
H122.17892.95252.75481.09583.47663.25083.10153.76662.54621.78233.76631.77724.32743.7777
H132.18293.00053.47841.09762.76852.59573.76804.33373.76951.77633.11251.77723.77772.5680
H142.17892.95252.75483.47661.09583.25083.10152.54623.76663.76631.78234.32743.77771.7772
H152.18293.00053.47842.76851.09762.59573.76803.76954.33373.11251.77633.77772.56801.7772

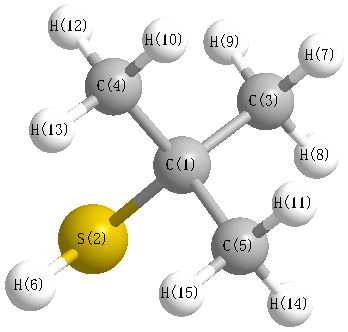
picture of 2-Propanethiol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 H6 96.280 C1 C3 H7 109.488
C1 C3 H8 111.154 C1 C3 H9 111.154
C1 C4 H10 109.857 C1 C4 H12 110.982
C1 C4 H13 111.188 C1 C5 H11 109.857
C1 C5 H14 110.982 C1 C5 H15 111.188
S2 C1 C3 105.641 S2 C1 C4 110.119
S2 C1 C5 110.119 C3 C1 C4 110.211
C3 C1 C5 110.211 C4 C1 C5 110.439
H7 C3 H8 108.226 H7 C3 H9 108.226
H8 C3 H9 108.498 H10 C4 H12 108.573
H10 C4 H13 107.909 H11 C5 H14 108.573
H11 C5 H15 107.909 H12 C4 H13 108.236
H14 C5 H15 108.236
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability