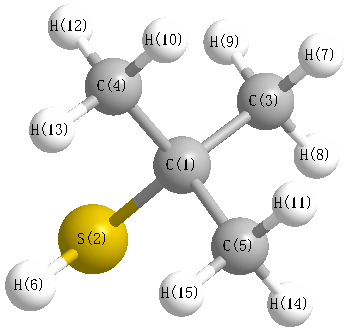
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3164 |
3009 |
23.34 |
|
|
|
| 2 |
A' |
3148 |
2994 |
23.36 |
|
|
|
| 3 |
A' |
3136 |
2982 |
42.50 |
|
|
|
| 4 |
A' |
3070 |
2920 |
20.00 |
|
|
|
| 5 |
A' |
3061 |
2911 |
33.03 |
|
|
|
| 6 |
A' |
2723 |
2590 |
22.48 |
|
|
|
| 7 |
A' |
1540 |
1464 |
8.45 |
|
|
|
| 8 |
A' |
1526 |
1451 |
9.56 |
|
|
|
| 9 |
A' |
1510 |
1436 |
0.64 |
|
|
|
| 10 |
A' |
1453 |
1381 |
1.42 |
|
|
|
| 11 |
A' |
1424 |
1354 |
12.59 |
|
|
|
| 12 |
A' |
1282 |
1219 |
0.45 |
|
|
|
| 13 |
A' |
1216 |
1156 |
45.82 |
|
|
|
| 14 |
A' |
1073 |
1020 |
2.40 |
|
|
|
| 15 |
A' |
958 |
911 |
0.91 |
|
|
|
| 16 |
A' |
896 |
852 |
5.08 |
|
|
|
| 17 |
A' |
847 |
806 |
1.41 |
|
|
|
| 18 |
A' |
602 |
573 |
5.41 |
|
|
|
| 19 |
A' |
391 |
372 |
1.13 |
|
|
|
| 20 |
A' |
368 |
350 |
0.40 |
|
|
|
| 21 |
A' |
299 |
284 |
0.24 |
|
|
|
| 22 |
A' |
283 |
269 |
0.89 |
|
|
|
| 23 |
A" |
3163 |
3008 |
22.35 |
|
|
|
| 24 |
A" |
3158 |
3003 |
6.45 |
|
|
|
| 25 |
A" |
3131 |
2978 |
0.66 |
|
|
|
| 26 |
A" |
3057 |
2907 |
17.26 |
|
|
|
| 27 |
A" |
1529 |
1454 |
8.13 |
|
|
|
| 28 |
A" |
1507 |
1433 |
0.52 |
|
|
|
| 29 |
A" |
1500 |
1426 |
0.01 |
|
|
|
| 30 |
A" |
1424 |
1354 |
11.42 |
|
|
|
| 31 |
A" |
1268 |
1206 |
4.01 |
|
|
|
| 32 |
A" |
1062 |
1010 |
0.13 |
|
|
|
| 33 |
A" |
977 |
929 |
0.02 |
|
|
|
| 34 |
A" |
952 |
905 |
0.57 |
|
|
|
| 35 |
A" |
400 |
380 |
0.55 |
|
|
|
| 36 |
A" |
304 |
289 |
3.83 |
|
|
|
| 37 |
A" |
286 |
272 |
0.00 |
|
|
|
| 38 |
A" |
258 |
245 |
8.74 |
|
|
|
| 39 |
A" |
214 |
204 |
10.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29077.6 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 27652.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.133 |
|
|
|
| 2 |
S |
-0.086 |
|
|
|
| 3 |
C |
-0.510 |
|
|
|
| 4 |
C |
-0.487 |
|
|
|
| 5 |
C |
-0.487 |
|
|
|
| 6 |
H |
0.100 |
|
|
|
| 7 |
H |
0.176 |
|
|
|
| 8 |
H |
0.187 |
|
|
|
| 9 |
H |
0.187 |
|
|
|
| 10 |
H |
0.169 |
|
|
|
| 11 |
H |
0.169 |
|
|
|
| 12 |
H |
0.188 |
|
|
|
| 13 |
H |
0.169 |
|
|
|
| 14 |
H |
0.188 |
|
|
|
| 15 |
H |
0.169 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.649 |
-0.867 |
0.000 |
1.863 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.059 |
2.379 |
0.000 |
| y |
2.379 |
-38.769 |
0.000 |
| z |
0.000 |
0.000 |
-41.698 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.826 |
2.379 |
0.000 |
| y |
2.379 |
3.110 |
0.000 |
| z |
0.000 |
0.000 |
-1.284 |
|
| Polar |
|---|
| 3z2-r2 | -2.568 |
|---|
| x2-y2 | -3.290 |
|---|
| xy | 2.379 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.879 |
0.123 |
0.000 |
| y |
0.123 |
8.466 |
0.000 |
| z |
0.000 |
0.000 |
7.825 |
<r2> (average value of r
2) Å
2
| <r2> |
164.476 |
| (<r2>)1/2 |
12.825 |