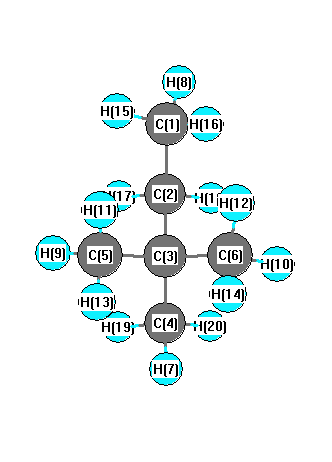
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3133 |
2997 |
52.43 |
|
|
|
| 2 |
A' |
3127 |
2992 |
62.68 |
|
|
|
| 3 |
A' |
3122 |
2987 |
66.85 |
|
|
|
| 4 |
A' |
3116 |
2981 |
0.20 |
|
|
|
| 5 |
A' |
3061 |
2928 |
21.37 |
|
|
|
| 6 |
A' |
3051 |
2919 |
12.17 |
|
|
|
| 7 |
A' |
3041 |
2910 |
37.23 |
|
|
|
| 8 |
A' |
3030 |
2899 |
30.79 |
|
|
|
| 9 |
A' |
1546 |
1479 |
11.80 |
|
|
|
| 10 |
A' |
1537 |
1470 |
7.04 |
|
|
|
| 11 |
A' |
1533 |
1466 |
5.79 |
|
|
|
| 12 |
A' |
1514 |
1448 |
0.04 |
|
|
|
| 13 |
A' |
1504 |
1438 |
0.05 |
|
|
|
| 14 |
A' |
1451 |
1388 |
4.10 |
|
|
|
| 15 |
A' |
1436 |
1373 |
6.99 |
|
|
|
| 16 |
A' |
1421 |
1360 |
6.36 |
|
|
|
| 17 |
A' |
1387 |
1327 |
0.01 |
|
|
|
| 18 |
A' |
1292 |
1236 |
3.83 |
|
|
|
| 19 |
A' |
1253 |
1199 |
8.19 |
|
|
|
| 20 |
A' |
1108 |
1060 |
2.12 |
|
|
|
| 21 |
A' |
1046 |
1001 |
2.42 |
|
|
|
| 22 |
A' |
1020 |
976 |
3.63 |
|
|
|
| 23 |
A' |
946 |
905 |
1.13 |
|
|
|
| 24 |
A' |
889 |
850 |
0.22 |
|
|
|
| 25 |
A' |
719 |
688 |
0.40 |
|
|
|
| 26 |
A' |
483 |
462 |
0.50 |
|
|
|
| 27 |
A' |
405 |
388 |
0.02 |
|
|
|
| 28 |
A' |
353 |
338 |
0.01 |
|
|
|
| 29 |
A' |
283 |
271 |
0.06 |
|
|
|
| 30 |
A' |
257 |
246 |
0.02 |
|
|
|
| 31 |
A" |
3139 |
3004 |
35.73 |
|
|
|
| 32 |
A" |
3126 |
2991 |
25.97 |
|
|
|
| 33 |
A" |
3119 |
2984 |
34.86 |
|
|
|
| 34 |
A" |
3116 |
2981 |
0.19 |
|
|
|
| 35 |
A" |
3068 |
2935 |
20.55 |
|
|
|
| 36 |
A" |
3045 |
2913 |
37.93 |
|
|
|
| 37 |
A" |
1540 |
1474 |
0.02 |
|
|
|
| 38 |
A" |
1528 |
1462 |
11.36 |
|
|
|
| 39 |
A" |
1512 |
1446 |
0.20 |
|
|
|
| 40 |
A" |
1505 |
1440 |
0.22 |
|
|
|
| 41 |
A" |
1420 |
1359 |
9.11 |
|
|
|
| 42 |
A" |
1350 |
1291 |
0.67 |
|
|
|
| 43 |
A" |
1260 |
1206 |
1.68 |
|
|
|
| 44 |
A" |
1114 |
1065 |
0.41 |
|
|
|
| 45 |
A" |
1009 |
965 |
0.58 |
|
|
|
| 46 |
A" |
966 |
924 |
0.02 |
|
|
|
| 47 |
A" |
950 |
909 |
0.35 |
|
|
|
| 48 |
A" |
790 |
756 |
2.93 |
|
|
|
| 49 |
A" |
417 |
399 |
0.02 |
|
|
|
| 50 |
A" |
335 |
320 |
0.01 |
|
|
|
| 51 |
A" |
293 |
281 |
0.00 |
|
|
|
| 52 |
A" |
255 |
244 |
0.00 |
|
|
|
| 53 |
A" |
218 |
208 |
0.01 |
|
|
|
| 54 |
A" |
94 |
90 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41614.8 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 39812.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.519 |
|
|
-0.280 |
| 2 |
C |
-0.289 |
|
|
-0.006 |
| 3 |
C |
0.055 |
|
|
0.607 |
| 4 |
C |
-0.498 |
|
|
-0.584 |
| 5 |
C |
-0.502 |
|
|
-0.468 |
| 6 |
C |
-0.502 |
|
|
-0.468 |
| 7 |
H |
0.160 |
|
|
0.143 |
| 8 |
H |
0.167 |
|
|
0.080 |
| 9 |
H |
0.160 |
|
|
0.102 |
| 10 |
H |
0.160 |
|
|
0.102 |
| 11 |
H |
0.164 |
|
|
0.102 |
| 12 |
H |
0.164 |
|
|
0.102 |
| 13 |
H |
0.161 |
|
|
0.100 |
| 14 |
H |
0.161 |
|
|
0.100 |
| 15 |
H |
0.168 |
|
|
0.063 |
| 16 |
H |
0.168 |
|
|
0.063 |
| 17 |
H |
0.152 |
|
|
0.002 |
| 18 |
H |
0.152 |
|
|
0.002 |
| 19 |
H |
0.160 |
|
|
0.119 |
| 20 |
H |
0.160 |
|
|
0.119 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.022 |
-0.035 |
0.000 |
0.042 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.001 |
-0.025 |
0.000 |
0.025 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.307 |
-0.288 |
0.000 |
| y |
-0.288 |
-41.382 |
0.000 |
| z |
0.000 |
0.000 |
-41.224 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.004 |
-0.288 |
0.000 |
| y |
-0.288 |
-0.117 |
0.000 |
| z |
0.000 |
0.000 |
0.121 |
|
| Polar |
|---|
| 3z2-r2 | 0.242 |
|---|
| x2-y2 | 0.075 |
|---|
| xy | -0.288 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
196.324 |
| (<r2>)1/2 |
14.012 |