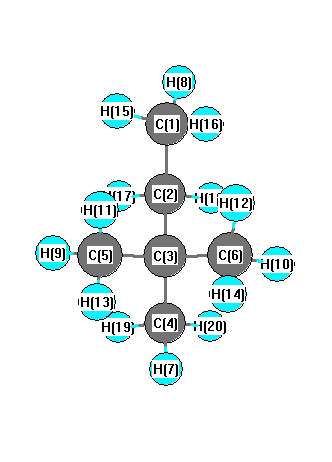
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3147 |
2993 |
42.55 |
|
|
|
| 2 |
A' |
3142 |
2988 |
64.20 |
|
|
|
| 3 |
A' |
3137 |
2983 |
65.51 |
|
|
|
| 4 |
A' |
3131 |
2977 |
1.06 |
|
|
|
| 5 |
A' |
3073 |
2922 |
20.75 |
|
|
|
| 6 |
A' |
3063 |
2913 |
10.81 |
|
|
|
| 7 |
A' |
3053 |
2903 |
33.52 |
|
|
|
| 8 |
A' |
3041 |
2892 |
31.69 |
|
|
|
| 9 |
A' |
1545 |
1470 |
12.28 |
|
|
|
| 10 |
A' |
1537 |
1462 |
7.05 |
|
|
|
| 11 |
A' |
1534 |
1459 |
6.04 |
|
|
|
| 12 |
A' |
1514 |
1440 |
0.08 |
|
|
|
| 13 |
A' |
1503 |
1430 |
0.13 |
|
|
|
| 14 |
A' |
1453 |
1381 |
4.65 |
|
|
|
| 15 |
A' |
1441 |
1371 |
7.19 |
|
|
|
| 16 |
A' |
1421 |
1352 |
8.02 |
|
|
|
| 17 |
A' |
1391 |
1323 |
0.11 |
|
|
|
| 18 |
A' |
1301 |
1237 |
3.52 |
|
|
|
| 19 |
A' |
1261 |
1200 |
7.42 |
|
|
|
| 20 |
A' |
1111 |
1057 |
2.03 |
|
|
|
| 21 |
A' |
1053 |
1002 |
2.84 |
|
|
|
| 22 |
A' |
1021 |
971 |
3.56 |
|
|
|
| 23 |
A' |
952 |
905 |
1.22 |
|
|
|
| 24 |
A' |
896 |
852 |
0.23 |
|
|
|
| 25 |
A' |
727 |
692 |
0.42 |
|
|
|
| 26 |
A' |
486 |
462 |
0.52 |
|
|
|
| 27 |
A' |
405 |
386 |
0.02 |
|
|
|
| 28 |
A' |
354 |
337 |
0.02 |
|
|
|
| 29 |
A' |
269 |
256 |
0.04 |
|
|
|
| 30 |
A' |
255 |
242 |
0.05 |
|
|
|
| 31 |
A" |
3151 |
2997 |
33.27 |
|
|
|
| 32 |
A" |
3139 |
2985 |
29.35 |
|
|
|
| 33 |
A" |
3133 |
2979 |
28.75 |
|
|
|
| 34 |
A" |
3130 |
2977 |
0.27 |
|
|
|
| 35 |
A" |
3081 |
2930 |
20.31 |
|
|
|
| 36 |
A" |
3056 |
2906 |
35.69 |
|
|
|
| 37 |
A" |
1541 |
1466 |
0.11 |
|
|
|
| 38 |
A" |
1530 |
1455 |
11.65 |
|
|
|
| 39 |
A" |
1510 |
1436 |
0.16 |
|
|
|
| 40 |
A" |
1505 |
1432 |
0.11 |
|
|
|
| 41 |
A" |
1423 |
1353 |
10.63 |
|
|
|
| 42 |
A" |
1355 |
1289 |
0.45 |
|
|
|
| 43 |
A" |
1267 |
1205 |
1.35 |
|
|
|
| 44 |
A" |
1116 |
1061 |
0.42 |
|
|
|
| 45 |
A" |
1011 |
962 |
0.50 |
|
|
|
| 46 |
A" |
964 |
917 |
0.04 |
|
|
|
| 47 |
A" |
955 |
908 |
0.44 |
|
|
|
| 48 |
A" |
795 |
756 |
2.92 |
|
|
|
| 49 |
A" |
419 |
399 |
0.02 |
|
|
|
| 50 |
A" |
337 |
320 |
0.02 |
|
|
|
| 51 |
A" |
291 |
277 |
0.00 |
|
|
|
| 52 |
A" |
259 |
247 |
0.00 |
|
|
|
| 53 |
A" |
209 |
199 |
0.01 |
|
|
|
| 54 |
A" |
91 |
86 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41742.7 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 39697.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.524 |
|
|
|
| 2 |
C |
-0.292 |
|
|
|
| 3 |
C |
0.043 |
|
|
|
| 4 |
C |
-0.501 |
|
|
|
| 5 |
C |
-0.505 |
|
|
|
| 6 |
C |
-0.505 |
|
|
|
| 7 |
H |
0.162 |
|
|
|
| 8 |
H |
0.169 |
|
|
|
| 9 |
H |
0.162 |
|
|
|
| 10 |
H |
0.162 |
|
|
|
| 11 |
H |
0.166 |
|
|
|
| 12 |
H |
0.166 |
|
|
|
| 13 |
H |
0.163 |
|
|
|
| 14 |
H |
0.163 |
|
|
|
| 15 |
H |
0.169 |
|
|
|
| 16 |
H |
0.169 |
|
|
|
| 17 |
H |
0.154 |
|
|
|
| 18 |
H |
0.154 |
|
|
|
| 19 |
H |
0.162 |
|
|
|
| 20 |
H |
0.162 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.041 |
0.000 |
0.000 |
0.041 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.225 |
0.173 |
0.000 |
| y |
0.173 |
-41.701 |
0.000 |
| z |
0.000 |
0.000 |
-41.345 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.298 |
0.173 |
0.000 |
| y |
0.173 |
-0.416 |
0.000 |
| z |
0.000 |
0.000 |
0.118 |
|
| Polar |
|---|
| 3z2-r2 | 0.236 |
|---|
| x2-y2 | 0.476 |
|---|
| xy | 0.173 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.934 |
-0.389 |
0.000 |
| y |
-0.389 |
9.398 |
0.000 |
| z |
0.000 |
0.000 |
9.208 |
<r2> (average value of r
2) Å
2
| <r2> |
195.403 |
| (<r2>)1/2 |
13.979 |