return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (3-methylenepentane)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-235.533056
Energy at 298.15K-235.545523
HF Energy-235.274563
Nuclear repulsion energy239.841082
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3182 3021 11.41      
2 A 3161 3000 40.99      
3 A 3155 2995 5.27      
4 A 3111 2953 14.94      
5 A 3082 2926 22.61      
6 A 3067 2911 0.70      
7 A 1732 1644 10.46      
8 A 1564 1484 0.29      
9 A 1552 1473 3.92      
10 A 1534 1456 4.61      
11 A 1493 1418 1.13      
12 A 1453 1379 0.66      
13 A 1383 1313 4.87      
14 A 1316 1250 0.66      
15 A 1148 1090 2.06      
16 A 1101 1045 0.70      
17 A 1026 974 0.00      
18 A 825 783 3.30      
19 A 773 733 0.02      
20 A 731 693 0.09      
21 A 457 434 0.59      
22 A 268 254 0.04      
23 A 238 226 0.00      
24 A 39 37 0.01      
25 B 3262 3096 21.56      
26 B 3158 2998 30.60      
27 B 3156 2995 70.85      
28 B 3112 2954 19.62      
29 B 3083 2926 36.56      
30 B 3066 2910 38.59      
31 B 1554 1475 10.34      
32 B 1552 1473 1.23      
33 B 1535 1457 5.43      
34 B 1453 1379 3.52      
35 B 1394 1323 1.30      
36 B 1347 1279 1.87      
37 B 1299 1233 1.33      
38 B 1135 1078 2.60      
39 B 1071 1017 1.29      
40 B 1047 993 1.66      
41 B 924 877 38.53      
42 B 923 876 5.84      
43 B 803 762 0.87      
44 B 600 569 6.27      
45 B 435 413 1.73      
46 B 292 277 0.12      
47 B 206 196 0.35      
48 B 108 102 0.07      

Unscaled Zero Point Vibrational Energy (zpe) 36953.9 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 35076.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.13072 0.09484 0.06502

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 2.008
C2 0.000 0.000 0.669
C3 0.000 1.297 -0.111
C4 0.000 -1.297 -0.111
C5 1.230 1.474 -1.013
C6 -1.230 -1.474 -1.013
H7 -0.003 0.923 2.580
H8 0.003 -0.923 2.580
H9 -0.055 2.133 0.593
H10 0.055 -2.133 0.593
H11 -0.904 1.357 -0.731
H12 0.904 -1.357 -0.731
H13 1.205 2.447 -1.512
H14 -1.205 -2.447 -1.512
H15 2.152 1.417 -0.427
H16 -2.152 -1.417 -0.427
H17 1.278 0.706 -1.790
H18 -1.278 -0.706 -1.790

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.33922.48472.48473.57953.57951.08591.08592.56022.56023.18743.18744.45334.45333.54483.54484.06864.0686
C21.33921.51341.51342.55252.55252.12202.12202.13502.13502.14872.14873.49253.49252.79962.79962.85922.8592
C32.48471.51342.59341.53543.16292.71663.48851.09483.50171.09732.87072.17664.17502.17803.47742.19032.9086
C42.48471.51342.59343.16291.53543.48852.71663.50171.09482.87071.09734.17502.17663.47742.17802.90862.1903
C53.57952.55251.53543.16293.84013.83784.49002.15974.11972.15552.86371.09384.64301.09384.48761.09353.4126
C63.57952.55253.16291.53543.84014.49003.83784.11972.15972.86372.15554.64301.09384.48761.09383.41261.0935
H71.08592.12202.71663.48853.83784.49001.84672.32623.64553.45814.12064.53005.43563.73134.37404.55804.8342
H81.08592.12203.48852.71664.49003.83781.84673.64552.32624.12063.45815.43564.53004.37403.73134.83424.5580
H92.56022.13501.09483.50172.15974.11972.32623.64554.26721.75383.85362.47345.17032.53434.24703.08043.9029
H102.56022.13503.50171.09484.11972.15973.64552.32624.26723.85361.75385.17032.47344.24702.53433.90293.0804
H113.18742.14871.09732.87072.15552.86373.45814.12061.75383.85363.26032.49923.89503.07103.05662.51042.3486
H123.18742.14872.87071.09732.86372.15554.12063.45813.85361.75383.26033.89502.49923.05663.07102.34862.5104
H134.45333.49252.17664.17501.09384.64304.53005.43562.47345.17032.49923.89505.45581.77075.23231.76464.0230
H144.45333.49254.17502.17664.64301.09385.43564.53005.17032.47343.89502.49925.45585.23231.77074.02301.7646
H153.54482.79962.17803.47741.09384.48763.73134.37402.53434.24703.07103.05661.77075.23235.15261.76824.2573
H163.54482.79963.47742.17804.48761.09384.37403.73134.24702.53433.05663.07105.23231.77075.15264.25731.7682
H174.06862.85922.19032.90861.09353.41264.55804.83423.08043.90292.51042.34861.76464.02301.76824.25732.9195
H184.06862.85922.90862.19033.41261.09354.83424.55803.90293.08042.34862.51044.02301.76464.25731.76822.9195

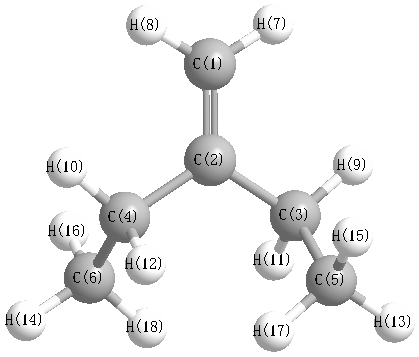
picture of 3-methylenepentane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.038 C1 C2 C4 121.038
C2 C1 H7 121.750 C2 C1 H8 121.750
C2 C3 C5 113.694 C2 C3 H9 108.823
C2 C3 H11 109.756 C2 C4 C6 113.694
C2 C4 H10 108.823 C2 C4 H12 109.756
C3 C2 C4 117.924 C3 C5 H13 110.639
C3 C5 H15 110.752 C3 C5 H17 111.756
C4 C6 H14 110.639 C4 C6 H16 110.752
C4 C6 H18 111.756 C5 C3 H9 109.253
C5 C3 H11 108.779 C6 C4 H10 109.253
C6 C4 H12 108.779 H7 C1 H8 116.501
H9 C3 H11 106.270 H10 C4 H12 106.270
H13 C5 H15 108.089 H13 C5 H17 107.568
H14 C6 H16 108.089 H14 C6 H18 107.568
H15 C5 H17 107.887 H16 C6 H18 107.887
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.427      
2 C 0.194      
3 C -0.337      
4 C -0.337      
5 C -0.465      
6 C -0.465      
7 H 0.146      
8 H 0.146      
9 H 0.153      
10 H 0.153      
11 H 0.154      
12 H 0.154      
13 H 0.154      
14 H 0.154      
15 H 0.158      
16 H 0.158      
17 H 0.153      
18 H 0.153      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 7.279 0.533 0.000
y 0.533 9.426 0.000
z 0.000 0.000 11.047


<r2> (average value of r2) Å2
<r2> 203.662
(<r2>)1/2 14.271