return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHClCHCH2CH3 ((E)-1-Chloro-1-butene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-616.451161
Energy at 298.15K-616.458254
HF Energy-616.237087
Nuclear repulsion energy200.446145
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3257 3091 10.01      
2 A 3204 3042 7.62      
3 A 3164 3003 32.10      
4 A 3157 2996 34.64      
5 A 3111 2953 12.19      
6 A 3083 2926 28.11      
7 A 3066 2911 26.41      
8 A 1715 1628 25.92      
9 A 1556 1477 4.44      
10 A 1548 1469 6.04      
11 A 1532 1455 2.92      
12 A 1458 1384 1.16      
13 A 1392 1322 2.60      
14 A 1346 1278 12.42      
15 A 1329 1261 7.01      
16 A 1285 1220 8.02      
17 A 1167 1108 1.46      
18 A 1116 1060 1.55      
19 A 1057 1004 9.17      
20 A 982 932 40.93      
21 A 930 883 3.05      
22 A 851 808 48.12      
23 A 813 771 8.96      
24 A 774 734 7.79      
25 A 447 424 1.87      
26 A 382 362 3.32      
27 A 298 283 0.24      
28 A 206 195 0.21      
29 A 168 159 0.31      
30 A 101 96 0.11      

Unscaled Zero Point Vibrational Energy (zpe) 22247.1 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 21117.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.59158 0.04655 0.04549

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 0.370 -1.196 0.012
C2 0.418 -0.106 0.020
H3 -0.821 1.637 0.018
Cl4 -2.306 -0.223 -0.006
C5 -0.734 0.557 0.010
H6 1.870 1.217 -0.860
H7 1.873 1.183 0.887
C8 1.778 0.547 -0.018
H9 3.886 0.032 -0.048
H10 2.869 -1.100 0.885
H11 2.853 -1.134 -0.857
C12 2.920 -0.473 -0.000

Atom - Atom Distances (Å)
  H1 C2 H3 Cl4 C5 H6 H7 C8 H9 H10 H11 C12
H11.09103.07232.84712.07142.97142.94662.24053.72442.64892.63132.6498
C21.09102.13782.72651.32932.15192.12801.50893.47112.78272.78482.5283
H33.07232.13782.37991.08322.86122.86622.81824.97314.67474.68384.2944
Cl42.84712.72652.37991.75444.49874.49824.15576.19705.32365.30725.2312
C52.07141.32931.08321.75442.82382.82102.51254.65024.06134.05953.7963
H62.97142.15192.86124.49872.82381.74711.07982.47533.06742.54812.1676
H72.94662.12802.86624.49822.82101.74711.11022.50052.49093.06142.1509
C82.24051.50892.81824.15572.51251.07981.11022.17012.17192.16461.5312
H93.72443.47114.97316.19704.65022.47532.50052.17011.78461.75521.0915
H102.64892.78274.67475.32364.06133.06742.49092.17191.78461.74241.0851
H112.63132.78484.68385.30724.05952.54813.06142.16461.75521.74241.0843
C122.64982.52834.29445.23123.79632.16762.15091.53121.09151.08511.0843

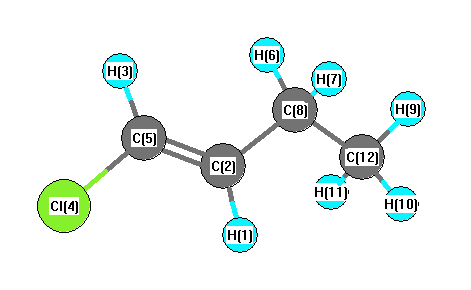
picture of (E)-1-Chloro-1-butene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C5 117.359 H1 C2 C8 118.142
C2 C5 H3 124.464 C2 C5 Cl4 123.714
C2 C8 H6 111.381 C2 C8 H7 107.715
C2 C8 C12 112.540 H3 C5 Cl4 111.820
C5 C2 C8 124.444 H6 C8 H7 105.827
H6 C8 C12 111.052 H7 C8 C12 107.981
C8 C12 H9 110.546 C8 C12 H10 111.077
C8 C12 H11 110.542 H9 C12 H10 110.144
H9 C12 H11 107.543 H10 C12 H11 106.863
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 H 0.176      
2 C -0.067      
3 H 0.201      
4 Cl -0.041      
5 C -0.285      
6 H 0.159      
7 H 0.165      
8 C -0.324      
9 H 0.158      
10 H 0.156      
11 H 0.163      
12 C -0.461      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 11.409 0.245 0.667
y 0.245 6.009 -0.778
z 0.667 -0.778 5.795


<r2> (average value of r2) Å2
<r2> 237.944
(<r2>)1/2 15.425