Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -637.490962 |
| Energy at 298.15K | -637.495962 |
| HF Energy | -636.976320 |
| Nuclear repulsion energy | 157.628325 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3229 |
2983 |
15.27 |
|
|
|
| 2 |
A' |
3196 |
2952 |
19.63 |
|
|
|
| 3 |
A' |
1619 |
1496 |
0.59 |
|
|
|
| 4 |
A' |
1587 |
1466 |
7.18 |
|
|
|
| 5 |
A' |
1516 |
1400 |
5.44 |
|
|
|
| 6 |
A' |
1374 |
1269 |
17.27 |
|
|
|
| 7 |
A' |
1152 |
1064 |
93.74 |
|
|
|
| 8 |
A' |
1136 |
1049 |
19.67 |
|
|
|
| 9 |
A' |
842 |
778 |
48.80 |
|
|
|
| 10 |
A' |
403 |
372 |
2.51 |
|
|
|
| 11 |
A' |
257 |
238 |
12.97 |
|
|
|
| 12 |
A" |
3297 |
3045 |
10.55 |
|
|
|
| 13 |
A" |
3253 |
3005 |
20.08 |
|
|
|
| 14 |
A" |
1372 |
1268 |
|
|
|
|
| 15 |
A" |
1290 |
1192 |
1.54 |
|
|
|
| 16 |
A" |
1140 |
1053 |
2.97 |
|
|
|
| 17 |
A" |
836 |
772 |
0.39 |
|
|
|
| 18 |
A" |
141 |
130 |
10.91 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13820.5 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 12766.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.603 |
0.000 |
| C2 |
0.988 |
-0.541 |
0.000 |
| Cl3 |
-1.662 |
-0.033 |
0.000 |
| F4 |
2.258 |
0.008 |
0.000 |
| H5 |
0.129 |
1.218 |
0.885 |
| H6 |
0.129 |
1.218 |
-0.885 |
| H7 |
0.875 |
-1.159 |
0.888 |
| H8 |
0.875 |
-1.159 |
-0.888 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5120 | 1.7799 | 2.3347 | 1.0850 | 1.0850 | 2.1588 | 2.1588 |
C2 | 1.5120 | | 2.6989 | 1.3829 | 2.1483 | 2.1483 | 1.0878 | 1.0878 | Cl3 | 1.7799 | 2.6989 | | 3.9201 | 2.3570 | 2.3570 | 2.9148 | 2.9148 | F4 | 2.3347 | 1.3829 | 3.9201 | | 2.6036 | 2.6036 | 2.0152 | 2.0152 | H5 | 1.0850 | 2.1483 | 2.3570 | 2.6036 | | 1.7695 | 2.4913 | 3.0577 | H6 | 1.0850 | 2.1483 | 2.3570 | 2.6036 | 1.7695 | | 3.0577 | 2.4913 | H7 | 2.1588 | 1.0878 | 2.9148 | 2.0152 | 2.4913 | 3.0577 | | 1.7761 | H8 | 2.1588 | 1.0878 | 2.9148 | 2.0152 | 3.0577 | 2.4913 | 1.7761 | |
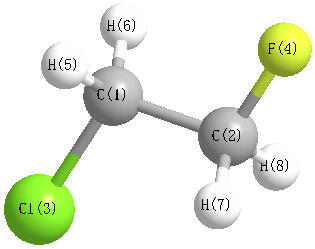 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.429 |
|
C1 |
C2 |
H7 |
111.224 |
| C1 |
C2 |
H8 |
111.224 |
|
C2 |
C1 |
Cl3 |
109.874 |
| C2 |
C1 |
H5 |
110.558 |
|
C2 |
C1 |
H6 |
110.558 |
| Cl3 |
C1 |
H5 |
108.259 |
|
Cl3 |
C1 |
H6 |
108.259 |
| F4 |
C2 |
H7 |
108.716 |
|
F4 |
C2 |
H8 |
108.716 |
| H5 |
C1 |
H6 |
109.270 |
|
H7 |
C2 |
H8 |
109.448 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -637.490024 |
| Energy at 298.15K | -637.495134 |
| HF Energy | -636.975059 |
| Nuclear repulsion energy | 161.948668 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3283 |
3032 |
7.22 |
|
|
|
| 2 |
A |
3240 |
2993 |
26.43 |
|
|
|
| 3 |
A |
3216 |
2971 |
14.00 |
|
|
|
| 4 |
A |
3174 |
2932 |
26.43 |
|
|
|
| 5 |
A |
1605 |
1482 |
1.22 |
|
|
|
| 6 |
A |
1564 |
1444 |
10.49 |
|
|
|
| 7 |
A |
1517 |
1401 |
19.39 |
|
|
|
| 8 |
A |
1432 |
1323 |
31.29 |
|
|
|
| 9 |
A |
1346 |
1243 |
0.52 |
|
|
|
| 10 |
A |
1295 |
1196 |
2.85 |
|
|
|
| 11 |
A |
1189 |
1098 |
62.28 |
|
|
|
| 12 |
A |
1140 |
1053 |
21.32 |
|
|
|
| 13 |
A |
1040 |
961 |
4.60 |
|
|
|
| 14 |
A |
913 |
843 |
9.14 |
|
|
|
| 15 |
A |
742 |
685 |
29.61 |
|
|
|
| 16 |
A |
490 |
453 |
13.89 |
|
|
|
| 17 |
A |
299 |
276 |
0.88 |
|
|
|
| 18 |
A |
143 |
132 |
2.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13812.9 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 12759.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.088 |
0.844 |
-0.293 |
| C2 |
1.201 |
0.415 |
0.358 |
| Cl3 |
-1.418 |
-0.283 |
0.066 |
| F4 |
1.638 |
-0.766 |
-0.202 |
| H5 |
-0.380 |
1.827 |
0.068 |
| H6 |
0.031 |
0.878 |
-1.371 |
| H7 |
1.962 |
1.177 |
0.186 |
| H8 |
1.074 |
0.264 |
1.428 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5067 | 1.7796 | 2.3621 | 1.0866 | 1.0855 | 2.1316 | 2.1559 |
C2 | 1.5067 | | 2.7259 | 1.3777 | 2.1394 | 2.1385 | 1.0907 | 1.0880 | Cl3 | 1.7796 | 2.7259 | | 3.1053 | 2.3509 | 2.3483 | 3.6837 | 2.8919 | F4 | 2.3621 | 1.3777 | 3.1053 | | 3.2963 | 2.5790 | 2.0075 | 2.0091 | H5 | 1.0866 | 2.1394 | 2.3509 | 3.2963 | | 1.7721 | 2.4334 | 2.5305 | H6 | 1.0855 | 2.1385 | 2.3483 | 2.5790 | 1.7721 | | 2.4984 | 3.0494 | H7 | 2.1316 | 1.0907 | 3.6837 | 2.0075 | 2.4334 | 2.4984 | | 1.7788 | H8 | 2.1559 | 1.0880 | 2.8919 | 2.0091 | 2.5305 | 3.0494 | 1.7788 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
109.873 |
|
C1 |
C2 |
H7 |
109.260 |
| C1 |
C2 |
H8 |
111.354 |
|
C2 |
C1 |
Cl3 |
111.826 |
| C2 |
C1 |
H5 |
110.118 |
|
C2 |
C1 |
H6 |
110.111 |
| Cl3 |
C1 |
H5 |
107.753 |
|
Cl3 |
C1 |
H6 |
107.615 |
| F4 |
C2 |
H7 |
108.272 |
|
F4 |
C2 |
H8 |
108.565 |
| H5 |
C1 |
H6 |
109.337 |
|
H7 |
C2 |
H8 |
109.459 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
