Jump to
S1C2
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -637.573497 |
| Energy at 298.15K | -637.578430 |
| HF Energy | -636.975452 |
| Nuclear repulsion energy | 156.743604 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3137 |
2987 |
15.79 |
|
|
|
| 2 |
A' |
3099 |
2951 |
20.31 |
|
|
|
| 3 |
A' |
1576 |
1501 |
0.46 |
|
|
|
| 4 |
A' |
1545 |
1472 |
5.85 |
|
|
|
| 5 |
A' |
1469 |
1399 |
4.90 |
|
|
|
| 6 |
A' |
1333 |
1269 |
16.38 |
|
|
|
| 7 |
A' |
1116 |
1063 |
27.31 |
|
|
|
| 8 |
A' |
1097 |
1045 |
72.81 |
|
|
|
| 9 |
A' |
813 |
774 |
42.38 |
|
|
|
| 10 |
A' |
391 |
372 |
2.33 |
|
|
|
| 11 |
A' |
249 |
237 |
12.37 |
|
|
|
| 12 |
A" |
3202 |
3049 |
11.70 |
|
|
|
| 13 |
A" |
3154 |
3004 |
21.20 |
|
|
|
| 14 |
A" |
1332 |
1269 |
0.02 |
|
|
|
| 15 |
A" |
1251 |
1191 |
1.06 |
|
|
|
| 16 |
A" |
1110 |
1057 |
2.89 |
|
|
|
| 17 |
A" |
817 |
778 |
0.39 |
|
|
|
| 18 |
A" |
138 |
131 |
10.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13413.0 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 12773.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.606 |
0.000 |
| C2 |
0.991 |
-0.543 |
0.000 |
| Cl3 |
-1.670 |
-0.036 |
0.000 |
| F4 |
2.271 |
0.012 |
0.000 |
| H5 |
0.130 |
1.224 |
0.891 |
| H6 |
0.130 |
1.224 |
-0.891 |
| H7 |
0.877 |
-1.165 |
0.895 |
| H8 |
0.877 |
-1.165 |
-0.895 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5172 | 1.7892 | 2.3471 | 1.0924 | 1.0924 | 2.1695 | 2.1695 |
C2 | 1.5172 | | 2.7090 | 1.3955 | 2.1581 | 2.1581 | 1.0957 | 1.0957 | Cl3 | 1.7892 | 2.7090 | | 3.9414 | 2.3713 | 2.3713 | 2.9265 | 2.9265 | F4 | 2.3471 | 1.3955 | 3.9414 | | 2.6162 | 2.6162 | 2.0324 | 2.0324 | H5 | 1.0924 | 2.1581 | 2.3713 | 2.6162 | | 1.7822 | 2.5034 | 3.0750 | H6 | 1.0924 | 2.1581 | 2.3713 | 2.6162 | 1.7822 | | 3.0750 | 2.5034 | H7 | 2.1695 | 1.0957 | 2.9265 | 2.0324 | 2.5034 | 3.0750 | | 1.7892 | H8 | 2.1695 | 1.0957 | 2.9265 | 2.0324 | 3.0750 | 2.5034 | 1.7892 | |
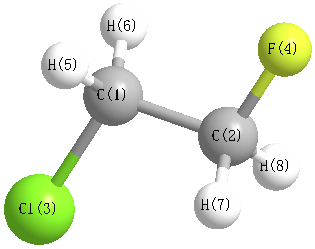 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.311 |
|
C1 |
C2 |
H7 |
111.239 |
| C1 |
C2 |
H8 |
111.239 |
|
C2 |
C1 |
Cl3 |
109.760 |
| C2 |
C1 |
H5 |
110.533 |
|
C2 |
C1 |
H6 |
110.533 |
| Cl3 |
C1 |
H5 |
108.314 |
|
Cl3 |
C1 |
H6 |
108.314 |
| F4 |
C2 |
H7 |
108.750 |
|
F4 |
C2 |
H8 |
108.750 |
| H5 |
C1 |
H6 |
109.328 |
|
H7 |
C2 |
H8 |
109.467 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -637.572684 |
| Energy at 298.15K | -637.577727 |
| HF Energy | -636.974185 |
| Nuclear repulsion energy | 161.129572 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3187 |
3035 |
8.54 |
|
|
|
| 2 |
A |
3141 |
2991 |
28.37 |
|
|
|
| 3 |
A |
3122 |
2973 |
12.99 |
|
|
|
| 4 |
A |
3077 |
2931 |
28.33 |
|
|
|
| 5 |
A |
1561 |
1486 |
1.12 |
|
|
|
| 6 |
A |
1522 |
1449 |
9.23 |
|
|
|
| 7 |
A |
1470 |
1399 |
17.85 |
|
|
|
| 8 |
A |
1390 |
1324 |
28.24 |
|
|
|
| 9 |
A |
1304 |
1242 |
0.56 |
|
|
|
| 10 |
A |
1256 |
1196 |
1.67 |
|
|
|
| 11 |
A |
1148 |
1093 |
45.61 |
|
|
|
| 12 |
A |
1106 |
1054 |
27.96 |
|
|
|
| 13 |
A |
1012 |
964 |
4.57 |
|
|
|
| 14 |
A |
887 |
844 |
8.92 |
|
|
|
| 15 |
A |
717 |
683 |
25.23 |
|
|
|
| 16 |
A |
476 |
453 |
13.00 |
|
|
|
| 17 |
A |
292 |
278 |
0.85 |
|
|
|
| 18 |
A |
140 |
133 |
2.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13404.0 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 12764.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.088 |
0.851 |
-0.295 |
| C2 |
1.205 |
0.420 |
0.360 |
| Cl3 |
-1.422 |
-0.285 |
0.066 |
| F4 |
1.641 |
-0.774 |
-0.202 |
| H5 |
-0.383 |
1.840 |
0.068 |
| H6 |
0.034 |
0.884 |
-1.381 |
| H7 |
1.974 |
1.185 |
0.184 |
| H8 |
1.075 |
0.272 |
1.438 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5118 | 1.7885 | 2.3745 | 1.0943 | 1.0929 | 2.1431 | 2.1660 |
C2 | 1.5118 | | 2.7350 | 1.3901 | 2.1500 | 2.1482 | 1.0987 | 1.0959 | Cl3 | 1.7885 | 2.7350 | | 3.1129 | 2.3652 | 2.3620 | 3.7017 | 2.9023 | F4 | 2.3745 | 1.3901 | 3.1129 | | 3.3172 | 2.5924 | 2.0245 | 2.0255 | H5 | 1.0943 | 2.1500 | 2.3652 | 3.3172 | | 1.7850 | 2.4490 | 2.5425 | H6 | 1.0929 | 2.1482 | 2.3620 | 2.5924 | 1.7850 | | 2.5107 | 3.0664 | H7 | 2.1431 | 1.0987 | 3.7017 | 2.0245 | 2.4490 | 2.5107 | | 1.7922 | H8 | 2.1660 | 1.0959 | 2.9023 | 2.0255 | 2.5425 | 3.0664 | 1.7922 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
109.755 |
|
C1 |
C2 |
H7 |
109.345 |
| C1 |
C2 |
H8 |
111.330 |
|
C2 |
C1 |
Cl3 |
111.659 |
| C2 |
C1 |
H5 |
110.146 |
|
C2 |
C1 |
H6 |
110.085 |
| Cl3 |
C1 |
H5 |
107.820 |
|
Cl3 |
C1 |
H6 |
107.662 |
| F4 |
C2 |
H7 |
108.301 |
|
F4 |
C2 |
H8 |
108.549 |
| H5 |
C1 |
H6 |
109.393 |
|
H7 |
C2 |
H8 |
109.504 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
