Jump to
S1C2
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -638.238229 |
| Energy at 298.15K | -638.243029 |
| HF Energy | -638.238229 |
| Nuclear repulsion energy | 155.813230 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3041 |
2991 |
16.49 |
|
|
|
| 2 |
A' |
2987 |
2937 |
25.35 |
|
|
|
| 3 |
A' |
1500 |
1475 |
0.75 |
|
|
|
| 4 |
A' |
1473 |
1448 |
4.20 |
|
|
|
| 5 |
A' |
1394 |
1371 |
4.76 |
|
|
|
| 6 |
A' |
1259 |
1238 |
14.72 |
|
|
|
| 7 |
A' |
1056 |
1038 |
16.87 |
|
|
|
| 8 |
A' |
1053 |
1035 |
86.39 |
|
|
|
| 9 |
A' |
753 |
740 |
44.57 |
|
|
|
| 10 |
A' |
371 |
365 |
1.71 |
|
|
|
| 11 |
A' |
232 |
228 |
11.06 |
|
|
|
| 12 |
A" |
3106 |
3054 |
10.44 |
|
|
|
| 13 |
A" |
3037 |
2987 |
26.06 |
|
|
|
| 14 |
A" |
1273 |
1252 |
0.03 |
|
|
|
| 15 |
A" |
1191 |
1171 |
0.54 |
|
|
|
| 16 |
A" |
1047 |
1030 |
3.00 |
|
|
|
| 17 |
A" |
783 |
770 |
1.23 |
|
|
|
| 18 |
A" |
131 |
129 |
8.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12843.2 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 12631.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.610 |
0.000 |
| C2 |
1.006 |
-0.534 |
0.000 |
| Cl3 |
-1.682 |
-0.049 |
0.000 |
| F4 |
2.284 |
0.026 |
0.000 |
| H5 |
0.117 |
1.236 |
0.897 |
| H6 |
0.117 |
1.236 |
-0.897 |
| H7 |
0.883 |
-1.164 |
0.900 |
| H8 |
0.883 |
-1.164 |
-0.900 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5236 | 1.8063 | 2.3572 | 1.1002 | 1.1002 | 2.1764 | 2.1764 |
C2 | 1.5236 | | 2.7310 | 1.3948 | 2.1746 | 2.1746 | 1.1050 | 1.1050 | Cl3 | 1.8063 | 2.7310 | | 3.9659 | 2.3858 | 2.3858 | 2.9376 | 2.9376 | F4 | 2.3572 | 1.3948 | 3.9659 | | 2.6389 | 2.6389 | 2.0459 | 2.0459 | H5 | 1.1002 | 2.1746 | 2.3858 | 2.6389 | | 1.7946 | 2.5193 | 3.0944 | H6 | 1.1002 | 2.1746 | 2.3858 | 2.6389 | 1.7946 | | 3.0944 | 2.5193 | H7 | 2.1764 | 1.1050 | 2.9376 | 2.0459 | 2.5193 | 3.0944 | | 1.7993 | H8 | 2.1764 | 1.1050 | 2.9376 | 2.0459 | 3.0944 | 2.5193 | 1.7993 | |
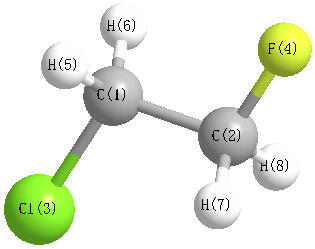 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.657 |
|
C1 |
C2 |
H7 |
110.769 |
| C1 |
C2 |
H8 |
110.769 |
|
C2 |
C1 |
Cl3 |
109.905 |
| C2 |
C1 |
H5 |
110.916 |
|
C2 |
C1 |
H6 |
110.916 |
| Cl3 |
C1 |
H5 |
107.860 |
|
Cl3 |
C1 |
H6 |
107.860 |
| F4 |
C2 |
H7 |
109.306 |
|
F4 |
C2 |
H8 |
109.306 |
| H5 |
C1 |
H6 |
109.282 |
|
H7 |
C2 |
H8 |
109.002 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.443 |
|
|
|
| 2 |
C |
0.010 |
|
|
|
| 3 |
Cl |
-0.062 |
|
|
|
| 4 |
F |
-0.273 |
|
|
|
| 5 |
H |
0.219 |
|
|
|
| 6 |
H |
0.219 |
|
|
|
| 7 |
H |
0.165 |
|
|
|
| 8 |
H |
0.165 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.441 |
0.129 |
0.000 |
0.459 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.480 |
-0.508 |
0.000 |
| y |
-0.508 |
-28.774 |
0.000 |
| z |
0.000 |
0.000 |
-29.410 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.388 |
-0.508 |
0.000 |
| y |
-0.508 |
4.172 |
0.000 |
| z |
0.000 |
0.000 |
3.217 |
|
| Polar |
|---|
| 3z2-r2 | 6.434 |
|---|
| x2-y2 | -7.707 |
|---|
| xy | -0.508 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.211 |
0.339 |
0.000 |
| y |
0.339 |
4.311 |
0.000 |
| z |
0.000 |
0.000 |
3.962 |
<r2> (average value of r
2) Å
2
| <r2> |
135.358 |
| (<r2>)1/2 |
11.634 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -638.237833 |
| Energy at 298.15K | -638.242742 |
| HF Energy | -638.237833 |
| Nuclear repulsion energy | 159.875840 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3093 |
3042 |
7.63 |
|
|
|
| 2 |
A |
3031 |
2981 |
25.13 |
|
|
|
| 3 |
A |
3016 |
2967 |
18.02 |
|
|
|
| 4 |
A |
2954 |
2905 |
34.90 |
|
|
|
| 5 |
A |
1480 |
1456 |
3.02 |
|
|
|
| 6 |
A |
1444 |
1420 |
9.63 |
|
|
|
| 7 |
A |
1397 |
1374 |
17.08 |
|
|
|
| 8 |
A |
1309 |
1287 |
31.79 |
|
|
|
| 9 |
A |
1245 |
1225 |
0.71 |
|
|
|
| 10 |
A |
1199 |
1179 |
2.72 |
|
|
|
| 11 |
A |
1094 |
1076 |
62.52 |
|
|
|
| 12 |
A |
1049 |
1031 |
16.66 |
|
|
|
| 13 |
A |
951 |
935 |
6.72 |
|
|
|
| 14 |
A |
843 |
829 |
5.74 |
|
|
|
| 15 |
A |
664 |
653 |
29.58 |
|
|
|
| 16 |
A |
454 |
446 |
11.58 |
|
|
|
| 17 |
A |
280 |
276 |
0.96 |
|
|
|
| 18 |
A |
131 |
129 |
1.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12817.3 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 12605.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.084 |
0.852 |
-0.292 |
| C2 |
1.216 |
0.415 |
0.355 |
| Cl3 |
-1.442 |
-0.286 |
0.065 |
| F4 |
1.667 |
-0.774 |
-0.200 |
| H5 |
-0.378 |
1.846 |
0.080 |
| H6 |
0.026 |
0.891 |
-1.387 |
| H7 |
1.974 |
1.205 |
0.179 |
| H8 |
1.087 |
0.279 |
1.445 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5167 | 1.8073 | 2.3909 | 1.1018 | 1.1007 | 2.1406 | 2.1719 |
C2 | 1.5167 | | 2.7642 | 1.3870 | 2.1601 | 2.1633 | 1.1094 | 1.1055 | Cl3 | 1.8073 | 2.7642 | | 3.1586 | 2.3825 | 2.3761 | 3.7291 | 2.9361 | F4 | 2.3909 | 1.3870 | 3.1586 | | 3.3352 | 2.6218 | 2.0379 | 2.0376 | H5 | 1.1018 | 2.1601 | 2.3825 | 3.3352 | | 1.7965 | 2.4399 | 2.5424 | H6 | 1.1007 | 2.1633 | 2.3761 | 2.6218 | 1.7965 | | 2.5196 | 3.0856 | H7 | 2.1406 | 1.1094 | 3.7291 | 2.0379 | 2.4399 | 2.5196 | | 1.8017 | H8 | 2.1719 | 1.1055 | 2.9361 | 2.0376 | 2.5424 | 3.0856 | 1.8017 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.771 |
|
C1 |
C2 |
H7 |
108.202 |
| C1 |
C2 |
H8 |
110.867 |
|
C2 |
C1 |
Cl3 |
112.229 |
| C2 |
C1 |
H5 |
110.156 |
|
C2 |
C1 |
H6 |
110.478 |
| Cl3 |
C1 |
H5 |
107.479 |
|
Cl3 |
C1 |
H6 |
107.074 |
| F4 |
C2 |
H7 |
108.939 |
|
F4 |
C2 |
H8 |
109.145 |
| H5 |
C1 |
H6 |
109.309 |
|
H7 |
C2 |
H8 |
108.870 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.437 |
|
|
|
| 2 |
C |
0.012 |
|
|
|
| 3 |
Cl |
-0.057 |
|
|
|
| 4 |
F |
-0.264 |
|
|
|
| 5 |
H |
0.205 |
|
|
|
| 6 |
H |
0.220 |
|
|
|
| 7 |
H |
0.154 |
|
|
|
| 8 |
H |
0.167 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.026 |
2.461 |
0.189 |
2.673 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.762 |
1.966 |
0.824 |
| y |
1.966 |
-29.108 |
-0.397 |
| z |
0.824 |
-0.397 |
-29.274 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.571 |
1.966 |
0.824 |
| y |
1.966 |
1.910 |
-0.397 |
| z |
0.824 |
-0.397 |
1.661 |
|
| Polar |
|---|
| 3z2-r2 | 3.323 |
|---|
| x2-y2 | -3.654 |
|---|
| xy | 1.966 |
|---|
| xz | 0.824 |
|---|
| yz | -0.397 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.775 |
0.890 |
-0.084 |
| y |
0.890 |
4.722 |
-0.139 |
| z |
-0.084 |
-0.139 |
4.081 |
<r2> (average value of r
2) Å
2
| <r2> |
117.127 |
| (<r2>)1/2 |
10.823 |
