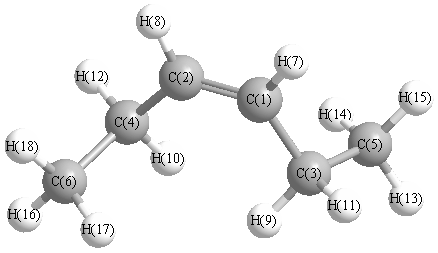
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3157 |
3020 |
62.66 |
|
|
|
| 2 |
A |
3138 |
3002 |
0.00 |
|
|
|
| 3 |
A |
3131 |
2996 |
72.60 |
|
|
|
| 4 |
A |
3105 |
2970 |
3.56 |
|
|
|
| 5 |
A |
3055 |
2923 |
0.33 |
|
|
|
| 6 |
A |
3035 |
2904 |
5.83 |
|
|
|
| 7 |
A |
1754 |
1678 |
0.89 |
|
|
|
| 8 |
A |
1534 |
1467 |
9.41 |
|
|
|
| 9 |
A |
1523 |
1457 |
3.64 |
|
|
|
| 10 |
A |
1513 |
1448 |
0.44 |
|
|
|
| 11 |
A |
1432 |
1370 |
0.18 |
|
|
|
| 12 |
A |
1361 |
1302 |
0.40 |
|
|
|
| 13 |
A |
1318 |
1261 |
0.03 |
|
|
|
| 14 |
A |
1287 |
1232 |
0.19 |
|
|
|
| 15 |
A |
1099 |
1052 |
0.86 |
|
|
|
| 16 |
A |
1082 |
1035 |
1.53 |
|
|
|
| 17 |
A |
1034 |
989 |
0.05 |
|
|
|
| 18 |
A |
1011 |
967 |
1.08 |
|
|
|
| 19 |
A |
858 |
821 |
1.94 |
|
|
|
| 20 |
A |
786 |
752 |
0.76 |
|
|
|
| 21 |
A |
515 |
492 |
0.38 |
|
|
|
| 22 |
A |
306 |
293 |
0.00 |
|
|
|
| 23 |
A |
207 |
198 |
0.00 |
|
|
|
| 24 |
A |
197 |
188 |
0.04 |
|
|
|
| 25 |
A |
62 |
60 |
0.01 |
|
|
|
| 26 |
B |
3138 |
3003 |
80.64 |
|
|
|
| 27 |
B |
3134 |
2998 |
2.28 |
|
|
|
| 28 |
B |
3128 |
2993 |
2.48 |
|
|
|
| 29 |
B |
3094 |
2960 |
7.36 |
|
|
|
| 30 |
B |
3055 |
2922 |
73.00 |
|
|
|
| 31 |
B |
3034 |
2903 |
50.64 |
|
|
|
| 32 |
B |
1532 |
1465 |
2.74 |
|
|
|
| 33 |
B |
1523 |
1457 |
8.72 |
|
|
|
| 34 |
B |
1503 |
1438 |
1.22 |
|
|
|
| 35 |
B |
1456 |
1393 |
4.62 |
|
|
|
| 36 |
B |
1430 |
1368 |
2.52 |
|
|
|
| 37 |
B |
1341 |
1283 |
12.92 |
|
|
|
| 38 |
B |
1298 |
1241 |
0.44 |
|
|
|
| 39 |
B |
1188 |
1136 |
0.20 |
|
|
|
| 40 |
B |
1105 |
1058 |
6.11 |
|
|
|
| 41 |
B |
1052 |
1007 |
3.74 |
|
|
|
| 42 |
B |
923 |
883 |
8.11 |
|
|
|
| 43 |
B |
818 |
783 |
5.50 |
|
|
|
| 44 |
B |
760 |
727 |
30.04 |
|
|
|
| 45 |
B |
567 |
543 |
6.83 |
|
|
|
| 46 |
B |
359 |
343 |
0.95 |
|
|
|
| 47 |
B |
241 |
230 |
0.04 |
|
|
|
| 48 |
B |
54 |
52 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36615.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 35029.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.125 |
|
|
|
| 2 |
C |
-0.125 |
|
|
|
| 3 |
C |
-0.356 |
|
|
|
| 4 |
C |
-0.356 |
|
|
|
| 5 |
C |
-0.506 |
|
|
|
| 6 |
C |
-0.506 |
|
|
|
| 7 |
H |
0.145 |
|
|
|
| 8 |
H |
0.145 |
|
|
|
| 9 |
H |
0.166 |
|
|
|
| 10 |
H |
0.166 |
|
|
|
| 11 |
H |
0.169 |
|
|
|
| 12 |
H |
0.169 |
|
|
|
| 13 |
H |
0.166 |
|
|
|
| 14 |
H |
0.174 |
|
|
|
| 15 |
H |
0.166 |
|
|
|
| 16 |
H |
0.166 |
|
|
|
| 17 |
H |
0.174 |
|
|
|
| 18 |
H |
0.166 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.201 |
0.201 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.854 |
0.446 |
0.000 |
| y |
0.446 |
-37.819 |
0.000 |
| z |
0.000 |
0.000 |
-38.440 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.724 |
0.446 |
0.000 |
| y |
0.446 |
1.828 |
0.000 |
| z |
0.000 |
0.000 |
0.896 |
|
| Polar |
|---|
| 3z2-r2 | 1.793 |
|---|
| x2-y2 | -3.035 |
|---|
| xy | 0.446 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.549 |
-0.886 |
0.000 |
| y |
-0.886 |
12.824 |
0.000 |
| z |
0.000 |
0.000 |
8.715 |
<r2> (average value of r
2) Å
2
| <r2> |
258.475 |
| (<r2>)1/2 |
16.077 |