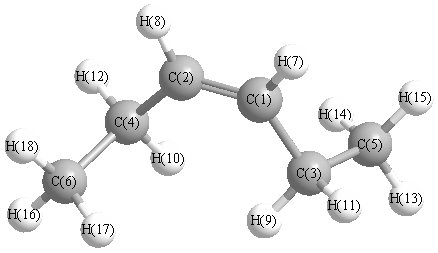
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3178 |
3017 |
62.04 |
|
|
|
| 2 |
A |
3159 |
2999 |
1.14 |
|
|
|
| 3 |
A |
3153 |
2993 |
72.67 |
|
|
|
| 4 |
A |
3131 |
2972 |
1.55 |
|
|
|
| 5 |
A |
3079 |
2922 |
0.43 |
|
|
|
| 6 |
A |
3062 |
2907 |
5.01 |
|
|
|
| 7 |
A |
1747 |
1658 |
0.46 |
|
|
|
| 8 |
A |
1559 |
1480 |
7.43 |
|
|
|
| 9 |
A |
1549 |
1470 |
3.10 |
|
|
|
| 10 |
A |
1541 |
1462 |
0.08 |
|
|
|
| 11 |
A |
1456 |
1382 |
0.16 |
|
|
|
| 12 |
A |
1384 |
1314 |
0.47 |
|
|
|
| 13 |
A |
1334 |
1266 |
0.01 |
|
|
|
| 14 |
A |
1306 |
1239 |
0.26 |
|
|
|
| 15 |
A |
1114 |
1057 |
0.64 |
|
|
|
| 16 |
A |
1098 |
1043 |
1.03 |
|
|
|
| 17 |
A |
1037 |
985 |
0.10 |
|
|
|
| 18 |
A |
1011 |
959 |
0.89 |
|
|
|
| 19 |
A |
863 |
819 |
1.35 |
|
|
|
| 20 |
A |
796 |
755 |
0.57 |
|
|
|
| 21 |
A |
518 |
491 |
0.31 |
|
|
|
| 22 |
A |
314 |
298 |
0.00 |
|
|
|
| 23 |
A |
212 |
201 |
0.00 |
|
|
|
| 24 |
A |
199 |
189 |
0.03 |
|
|
|
| 25 |
A |
56 |
53 |
0.01 |
|
|
|
| 26 |
B |
3160 |
2999 |
83.58 |
|
|
|
| 27 |
B |
3155 |
2995 |
2.81 |
|
|
|
| 28 |
B |
3150 |
2990 |
2.19 |
|
|
|
| 29 |
B |
3117 |
2958 |
8.55 |
|
|
|
| 30 |
B |
3078 |
2922 |
66.90 |
|
|
|
| 31 |
B |
3060 |
2905 |
50.59 |
|
|
|
| 32 |
B |
1556 |
1477 |
2.17 |
|
|
|
| 33 |
B |
1549 |
1470 |
7.05 |
|
|
|
| 34 |
B |
1530 |
1452 |
0.90 |
|
|
|
| 35 |
B |
1474 |
1399 |
3.54 |
|
|
|
| 36 |
B |
1453 |
1380 |
1.69 |
|
|
|
| 37 |
B |
1364 |
1295 |
15.01 |
|
|
|
| 38 |
B |
1317 |
1250 |
0.22 |
|
|
|
| 39 |
B |
1204 |
1143 |
0.25 |
|
|
|
| 40 |
B |
1122 |
1065 |
5.75 |
|
|
|
| 41 |
B |
1058 |
1004 |
3.03 |
|
|
|
| 42 |
B |
931 |
883 |
6.28 |
|
|
|
| 43 |
B |
828 |
786 |
4.27 |
|
|
|
| 44 |
B |
767 |
728 |
27.77 |
|
|
|
| 45 |
B |
571 |
542 |
5.80 |
|
|
|
| 46 |
B |
365 |
346 |
0.82 |
|
|
|
| 47 |
B |
245 |
232 |
0.03 |
|
|
|
| 48 |
B |
50 |
47 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36977.8 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 35099.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.114 |
|
|
|
| 2 |
C |
-0.114 |
|
|
|
| 3 |
C |
-0.314 |
|
|
|
| 4 |
C |
-0.314 |
|
|
|
| 5 |
C |
-0.453 |
|
|
|
| 6 |
C |
-0.453 |
|
|
|
| 7 |
H |
0.130 |
|
|
|
| 8 |
H |
0.130 |
|
|
|
| 9 |
H |
0.149 |
|
|
|
| 10 |
H |
0.149 |
|
|
|
| 11 |
H |
0.150 |
|
|
|
| 12 |
H |
0.150 |
|
|
|
| 13 |
H |
0.148 |
|
|
|
| 14 |
H |
0.155 |
|
|
|
| 15 |
H |
0.149 |
|
|
|
| 16 |
H |
0.148 |
|
|
|
| 17 |
H |
0.155 |
|
|
|
| 18 |
H |
0.149 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.184 |
0.184 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.360 |
0.483 |
0.000 |
| y |
0.483 |
-38.538 |
0.000 |
| z |
0.000 |
0.000 |
-39.047 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.567 |
0.483 |
0.000 |
| y |
0.483 |
1.665 |
0.000 |
| z |
0.000 |
0.000 |
0.902 |
|
| Polar |
|---|
| 3z2-r2 | 1.804 |
|---|
| x2-y2 | -2.822 |
|---|
| xy | 0.483 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.427 |
-0.833 |
0.000 |
| y |
-0.833 |
12.460 |
0.000 |
| z |
0.000 |
0.000 |
8.585 |
<r2> (average value of r
2) Å
2
| <r2> |
257.673 |
| (<r2>)1/2 |
16.052 |