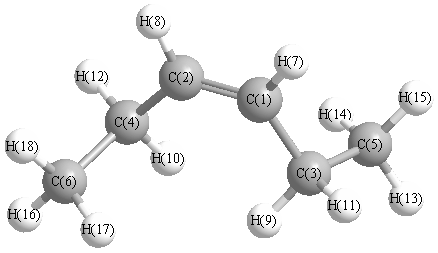
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3075 |
3024 |
65.07 |
|
|
|
| 2 |
A |
3064 |
3014 |
1.97 |
|
|
|
| 3 |
A |
3057 |
3006 |
60.67 |
|
|
|
| 4 |
A |
3023 |
2973 |
6.75 |
|
|
|
| 5 |
A |
2982 |
2933 |
0.29 |
|
|
|
| 6 |
A |
2956 |
2907 |
6.18 |
|
|
|
| 7 |
A |
1694 |
1666 |
0.95 |
|
|
|
| 8 |
A |
1491 |
1466 |
8.40 |
|
|
|
| 9 |
A |
1480 |
1455 |
3.54 |
|
|
|
| 10 |
A |
1468 |
1444 |
0.56 |
|
|
|
| 11 |
A |
1384 |
1361 |
0.22 |
|
|
|
| 12 |
A |
1315 |
1294 |
0.52 |
|
|
|
| 13 |
A |
1279 |
1257 |
0.05 |
|
|
|
| 14 |
A |
1249 |
1228 |
0.17 |
|
|
|
| 15 |
A |
1064 |
1047 |
0.84 |
|
|
|
| 16 |
A |
1050 |
1032 |
1.56 |
|
|
|
| 17 |
A |
1006 |
989 |
0.13 |
|
|
|
| 18 |
A |
969 |
953 |
0.98 |
|
|
|
| 19 |
A |
836 |
822 |
2.14 |
|
|
|
| 20 |
A |
765 |
752 |
0.72 |
|
|
|
| 21 |
A |
500 |
491 |
0.45 |
|
|
|
| 22 |
A |
301 |
296 |
0.00 |
|
|
|
| 23 |
A |
201 |
198 |
0.00 |
|
|
|
| 24 |
A |
190 |
187 |
0.05 |
|
|
|
| 25 |
A |
58 |
57 |
0.01 |
|
|
|
| 26 |
B |
3065 |
3014 |
70.47 |
|
|
|
| 27 |
B |
3057 |
3006 |
1.74 |
|
|
|
| 28 |
B |
3050 |
2999 |
7.88 |
|
|
|
| 29 |
B |
3015 |
2965 |
8.17 |
|
|
|
| 30 |
B |
2982 |
2933 |
72.66 |
|
|
|
| 31 |
B |
2954 |
2906 |
48.46 |
|
|
|
| 32 |
B |
1489 |
1464 |
2.46 |
|
|
|
| 33 |
B |
1480 |
1455 |
8.67 |
|
|
|
| 34 |
B |
1456 |
1432 |
1.03 |
|
|
|
| 35 |
B |
1411 |
1388 |
5.16 |
|
|
|
| 36 |
B |
1382 |
1359 |
1.88 |
|
|
|
| 37 |
B |
1296 |
1275 |
14.01 |
|
|
|
| 38 |
B |
1259 |
1238 |
0.67 |
|
|
|
| 39 |
B |
1154 |
1135 |
0.25 |
|
|
|
| 40 |
B |
1071 |
1053 |
5.18 |
|
|
|
| 41 |
B |
1026 |
1009 |
4.12 |
|
|
|
| 42 |
B |
898 |
883 |
8.67 |
|
|
|
| 43 |
B |
795 |
781 |
5.54 |
|
|
|
| 44 |
B |
736 |
724 |
27.86 |
|
|
|
| 45 |
B |
555 |
546 |
7.09 |
|
|
|
| 46 |
B |
348 |
342 |
0.89 |
|
|
|
| 47 |
B |
235 |
232 |
0.03 |
|
|
|
| 48 |
B |
49 |
48 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35607.8 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 35020.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.101 |
|
|
|
| 2 |
C |
-0.101 |
|
|
|
| 3 |
C |
-0.333 |
|
|
|
| 4 |
C |
-0.333 |
|
|
|
| 5 |
C |
-0.480 |
|
|
|
| 6 |
C |
-0.480 |
|
|
|
| 7 |
H |
0.125 |
|
|
|
| 8 |
H |
0.125 |
|
|
|
| 9 |
H |
0.154 |
|
|
|
| 10 |
H |
0.154 |
|
|
|
| 11 |
H |
0.158 |
|
|
|
| 12 |
H |
0.158 |
|
|
|
| 13 |
H |
0.156 |
|
|
|
| 14 |
H |
0.165 |
|
|
|
| 15 |
H |
0.158 |
|
|
|
| 16 |
H |
0.156 |
|
|
|
| 17 |
H |
0.165 |
|
|
|
| 18 |
H |
0.158 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.214 |
0.214 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.976 |
0.422 |
0.000 |
| y |
0.422 |
-37.993 |
0.000 |
| z |
0.000 |
0.000 |
-38.714 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.622 |
0.422 |
0.000 |
| y |
0.422 |
1.851 |
0.000 |
| z |
0.000 |
0.000 |
0.771 |
|
| Polar |
|---|
| 3z2-r2 | 1.542 |
|---|
| x2-y2 | -2.982 |
|---|
| xy | 0.422 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.737 |
-0.993 |
0.000 |
| y |
-0.993 |
13.276 |
0.000 |
| z |
0.000 |
0.000 |
8.959 |
<r2> (average value of r
2) Å
2
| <r2> |
260.934 |
| (<r2>)1/2 |
16.153 |